Chapter 4: The d- and f-Block Elements
On a small island near Stockholm lies a quarry called Ytterby. In the late eighteenth century, workers digging there for feldspar came across an unusually heavy black mineral. A sample reached the Finnish chemist Johan Gadolin, who analysed it and reported that it contained a new “earth” — an oxide of an element nobody had yet seen. That single rock turned out to be one of the richest chemical puzzles in history.
Over the next hundred years, chemist after chemist tried to break Gadolin’s earth apart. Every time they thought they had a pure substance, careful work showed it was still a mixture. The elements hidden inside behaved so alike — same charge, almost the same size, nearly the same reactions — that ordinary chemical separation simply failed. Crystallisation had to be repeated thousands of times to tease them apart. Four of the elements finally isolated were named after that one village: yttrium, ytterbium, terbium and erbium.
Why should a whole family of elements be so stubbornly identical? And why, on the other side of the periodic table, does a single metal such as manganese behave so differently in each of its many compounds — green in one, deep purple in another, a mild ion here and a fierce oxidising agent there? Both questions lead to the same place: the quiet filling of inner d and f orbitals.
Fig. 4.0 – Aqueous solutions of transition metals, gadolinite ore, and the d- and f-blocks.
Keep these questions in mind as you read. This chapter will give you everything you need to reason your way to the answers.
4.1 Position in the Periodic Table
Two Blocks Built on Inner Orbitals
Look at a long-form periodic table and you will see a wide middle region that seems to hold the table together — the elements of groups 3 to 12. These make up the d-block. In the four long periods of the table, it is the d orbitals of these atoms that are being progressively filled with electrons.
Below the main body of the table sits a separate panel of two rows. These are the elements of the f-block, in which the 4f and 5f orbitals are progressively filled. They are printed separately only to keep the table a manageable width; chemically, they belong inside it.
The elements of the d-block are usually called transition metals, and those of the f-block are called inner transition metals. Both names will be used constantly through this Chapter, so it is worth fixing them now.
The Four Series of Transition Metals
There are mainly four series of the transition metals. The 3d series runs from scandium (Sc) to zinc (Zn). The 4d series runs from yttrium (Y) to cadmium (Cd). The 5d series contains lanthanum (La) together with the elements from hafnium (Hf) to mercury (Hg). The 6d series contains actinium (Ac) along with the elements from rutherfordium (Rf) to copernicium (Cn).
The inner transition metals form two series of their own. The 4f series runs from cerium (Ce) to lutetium (Lu) and its members are called lanthanoids. The 5f series runs from thorium (Th) to lawrencium (Lr), and these are the actinoids.
What Counts as a Transition Element
The name “transition metal” was coined for a historical reason: the chemical properties of these metals were seen as transitional — a bridge between the very reactive, electropositive s-block metals on the left and the p-block elements on the right. The modern definition, however, is based squarely on electronic structure.
Notice the two-part condition carefully, because exam questions are built on it. It is enough for the ion to have a partly filled d subshell; the neutral atom need not.
This definition has an immediate consequence. Zinc, cadmium and mercury of group 12 have a full \(d^{10}\) configuration both in the ground state of the atom and in their common oxidation states. Neither the atom nor the usual ion has a partly filled d subshell, so strictly they are not regarded as transition metals. All the same, they are the end members of the 3d, 4d and 5d series respectively, and their chemistry is studied along with that of the transition metals.
Why the d and f Elements Deserve a Separate Study
It is the presence of partly filled d or f orbitals that makes transition elements behave so differently from the non-transition elements. That single structural fact is behind their variable oxidation states, their coloured ions, their magnetic behaviour, their catalytic power and their complexes — all of which are taken up later in this Chapter. For this reason, transition elements and their compounds are studied separately. This does not mean a new theory is needed: the usual theory of valence that works for the non-transition elements can be applied successfully to the transition elements too.
The Plan of This Chapter
The route ahead is worth knowing in advance. We shall first take up the electronic configuration, occurrence and general characteristics of the transition elements, with special emphasis on the trends in the properties of the first row (3d) transition metals. We shall then look at the preparation and properties of two important compounds of this block. After that we turn to certain general aspects of the inner transition metals — their electronic configurations, oxidation states and chemical reactivity.
Where the d-Block Sits
The d-block occupies the large middle section of the periodic table, flanked between the s-block and the p-block, as shown in Fig. 4.1. The d orbitals filled here belong to the penultimate energy level of the atom — that is, the shell just inside the outermost one. In an element of the 4th period, for instance, the outermost shell is the 4th, and the d orbitals receiving electrons are the 3d orbitals. Filling these penultimate d orbitals gives rise to the four rows of the transition metals: 3d, 4d, 5d and 6d.
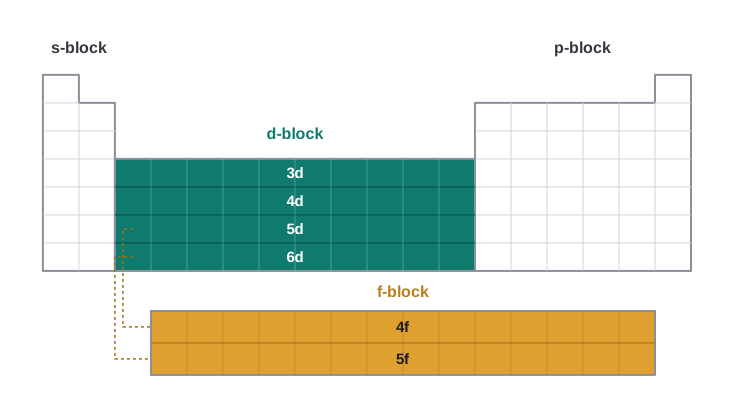
Fig. 4.1 – Position of the d- and f-blocks in the periodic table.
All these series of transition elements are set out in Table 4.1 in the next section, where their electronic configurations are given.
4.2 Electronic Configurations of the d-Block Elements
The General Pattern
In general, the electronic configuration of the outer orbitals of these elements is
\[(n-1)d^{1-10}\,ns^{1-2}\]
except for palladium, whose configuration is \(4d^{10}5s^{0}\).
Read the formula slowly. The \((n-1)\) stands for the inner d orbitals, which may hold anywhere from one to ten electrons. The \(ns\) part is the outermost s orbital, which may hold one or two electrons. So in an element of the 4th period such as iron, \(n = 4\): the 4s orbital is the outermost one, and the d orbitals being filled are the 3d orbitals, one shell inside it.
You may wonder why the outer 4s orbital gets filled before the inner 3d orbital at all. The reason, which you met while learning the Aufbau principle, is that the 4s orbital happens to lie slightly lower in energy than the 3d orbital when they are empty, so electrons enter 4s first. The gap between them, however, is very small — and that small gap is the source of everything that follows.
Why Chromium and Copper Break the Rule
The general formula has several exceptions, precisely because the energy difference between the \((n-1)d\) and \(ns\) orbitals is so little. On top of that, half-filled and completely filled sets of orbitals are relatively more stable than partly filled ones.
A consequence of this factor shows up in the configurations of chromium and copper in the 3d series. Consider chromium, which has a \(3d^5 4s^1\) configuration instead of \(3d^4 4s^2\); the energy gap between the two sets (3d and 4s) of orbitals is small enough to prevent the electron entering the 3d orbitals — that is, the electron is not held back in 4s but moves so that the 3d set becomes exactly half-filled. Similarly in the case of copper, the configuration is \(3d^{10}4s^1\) and not \(3d^9 4s^2\), giving a completely filled 3d set.
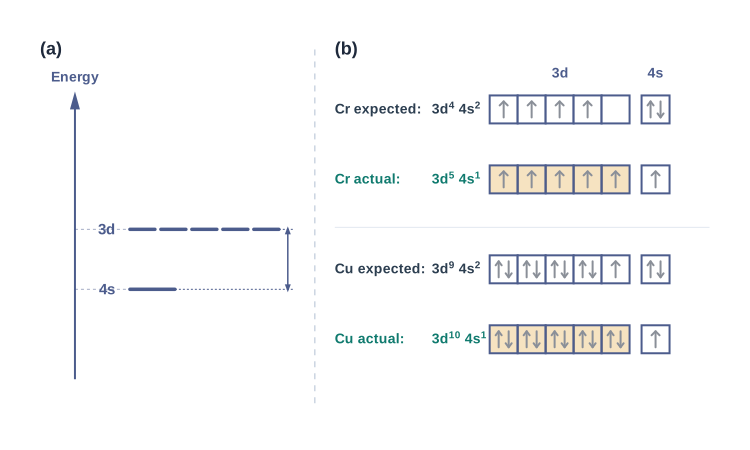
Fig. 4.2 – (a) 3d and 4s energy levels and (b) anomalous configurations of Cr and Cu.
The ground state electronic configurations of the outer orbitals of the transition elements are given in Table 4.1. Only the outermost s electrons and the \((n-1)d\) electrons are listed; the inner noble-gas core is understood.
Table 4.1: Electronic Configurations of outer orbitals of the Transition Elements (ground state)
1st Series
| Sc | Ti | V | Cr | Mn | Fe | Co | Ni | Cu | Zn | |
|---|---|---|---|---|---|---|---|---|---|---|
| \(Z\) | 21 | 22 | 23 | 24 | 25 | 26 | 27 | 28 | 29 | 30 |
| 4s | 2 | 2 | 2 | 1 | 2 | 2 | 2 | 2 | 1 | 2 |
| 3d | 1 | 2 | 3 | 5 | 5 | 6 | 7 | 8 | 10 | 10 |
2nd Series
| Y | Zr | Nb | Mo | Tc | Ru | Rh | Pd | Ag | Cd | |
|---|---|---|---|---|---|---|---|---|---|---|
| \(Z\) | 39 | 40 | 41 | 42 | 43 | 44 | 45 | 46 | 47 | 48 |
| 5s | 2 | 2 | 1 | 1 | 1 | 1 | 1 | 0 | 1 | 2 |
| 4d | 1 | 2 | 4 | 5 | 6 | 7 | 8 | 10 | 10 | 10 |
3rd Series
| La | Hf | Ta | W | Re | Os | Ir | Pt | Au | Hg | |
|---|---|---|---|---|---|---|---|---|---|---|
| \(Z\) | 57 | 72 | 73 | 74 | 75 | 76 | 77 | 78 | 79 | 80 |
| 6s | 2 | 2 | 2 | 2 | 2 | 2 | 2 | 1 | 1 | 2 |
| 5d | 1 | 2 | 3 | 4 | 5 | 6 | 7 | 9 | 10 | 10 |
4th Series
| Ac | Rf | Db | Sg | Bh | Hs | Mt | Ds | Rg | Cn | |
|---|---|---|---|---|---|---|---|---|---|---|
| \(Z\) | 89 | 104 | 105 | 106 | 107 | 108 | 109 | 110 | 111 | 112 |
| 7s | 2 | 2 | 2 | 2 | 2 | 2 | 2 | 2 | 1 | 2 |
| 6d | 1 | 2 | 3 | 4 | 5 | 6 | 7 | 8 | 10 | 10 |
The End Members: Zn, Cd, Hg and Cn
The electronic configurations of the outer orbitals of Zn, Cd, Hg and Cn are represented by the general formula \((n-1)d^{10}ns^{2}\). The orbitals in these elements are completely filled in the ground state as well as in their common oxidation states. Therefore, as we noted while defining the term, they are not regarded as transition elements.
What Partly Filled d Orbitals Make Possible
The d orbitals of the transition elements protrude to the periphery of an atom more than the other orbitals (that is, more than the s and p orbitals). They therefore lie exposed. This has two consequences: these orbitals are more influenced by the surroundings, and they in turn affect the atoms or molecules surrounding them.
In some respects, ions of a given \(d^n\) configuration (\(n = 1 - 9\)) have similar magnetic and electronic properties, whichever metal they came from. With partly filled d orbitals, these elements exhibit certain characteristic properties: display of a variety of oxidation states, formation of coloured ions, and entering into complex formation with a variety of ligands.
The transition metals and their compounds also exhibit catalytic property and paramagnetic behaviour. All these characteristics are discussed in detail later in this Chapter; for now, simply note that every one of them can be traced back to the partly filled d set.
Reading the Block Across, Not Down
There are greater similarities in the properties of the transition elements of a horizontal row, in contrast to the non-transition elements, where a group (a vertical column) is usually the family that behaves alike. However, some group similarities also exist among the transition metals. Accordingly, we shall first study the general characteristics and their trends in the horizontal rows — particularly the 3d row — and then consider some group similarities.
Writing the Configuration of an Ion
One point that trips students up constantly: when a d-block element forms an ion, the \(ns\) electrons are removed first, before the \((n-1)d\) electrons — even though the \(ns\) orbital was filled first. So the ion, unlike the atom, has no outer s electrons left to speak of.
4.3 General Properties of the Transition Elements (d-Block)
We will discuss the properties of elements of the first transition series only in the following sections. The 3d row is the row for which reliable data are easiest to compare, and once you understand it, the other rows fall into place.
4.3.1 Physical Properties
Nearly all the transition elements display typical metallic properties: high tensile strength, ductility, malleability, high thermal and electrical conductivity, and metallic lustre. With the exceptions of Zn, Cd, Hg and Mn, they have one or more typical metallic structures at normal temperatures — that is, their atoms pack in the close-packed or body-centred arrangements you met in the solid state.
Lattice Structures of Transition Metals
| Sc | Ti | V | Cr | Mn | Fe | Co | Ni | Cu | Zn |
|---|---|---|---|---|---|---|---|---|---|
| hcp (bcc) | hcp (bcc) | bcc | bcc | X (bcc, ccp) | bcc (hcp) | ccp (hcp) | ccp | ccp | X (hcp) |
| Y | Zr | Nb | Mo | Tc | Ru | Rh | Pd | Ag | Cd |
|---|---|---|---|---|---|---|---|---|---|
| hcp (bcc) | hcp (bcc) | bcc | bcc | hcp | hcp | ccp | ccp | ccp | X (hcp) |
| La | Hf | Ta | W | Re | Os | Ir | Pt | Au | Hg |
|---|---|---|---|---|---|---|---|---|---|
| hcp (ccp, bcc) | hcp (bcc) | bcc | bcc | hcp | hcp | ccp | ccp | ccp | X |
(bcc = body centred cubic; hcp = hexagonal close packed; ccp = cubic close packed; X = a typical metal structure).
Hardness, Melting Points and the d Electrons
The transition metals (with the exception of Zn, Cd and Hg) are very hard and have low volatility. Their melting and boiling points are high. Fig. 4.3 depicts the melting points of transition metals belonging to the 3d, 4d and 5d series; notice how each curve rises to a peak and then falls away.
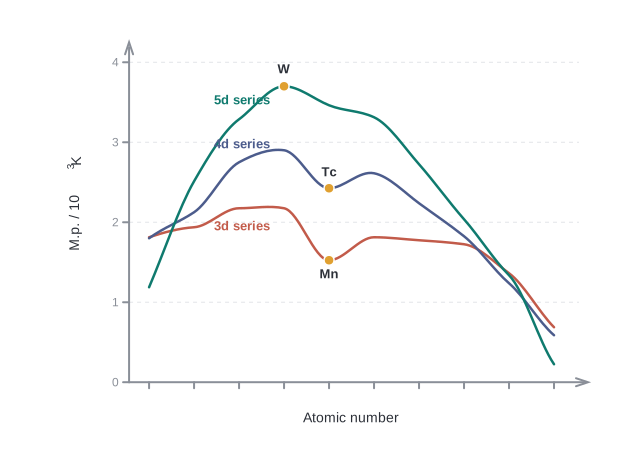
Fig. 4.3 – Trends in melting points of transition elements.
The high melting points of these metals are attributed to the involvement of a greater number of electrons from the \((n-1)d\) orbitals, in addition to the \(ns\) electrons, in the interatomic metallic bonding. In an s-block metal only the s electrons are available to glue the lattice together; in a transition metal the d electrons join in as well, and the metallic bond becomes far stronger.
In any row, the melting points of these metals rise to a maximum at \(d^5\), except for the anomalous values of Mn and Tc, and then fall regularly as the atomic number increases.
They also have high enthalpies of atomisation, which are shown in Fig. 4.4. The enthalpy of atomisation is the enthalpy needed to break one mole of the solid metal into free gaseous atoms — in effect, a direct measure of how strong the metallic bonding is.
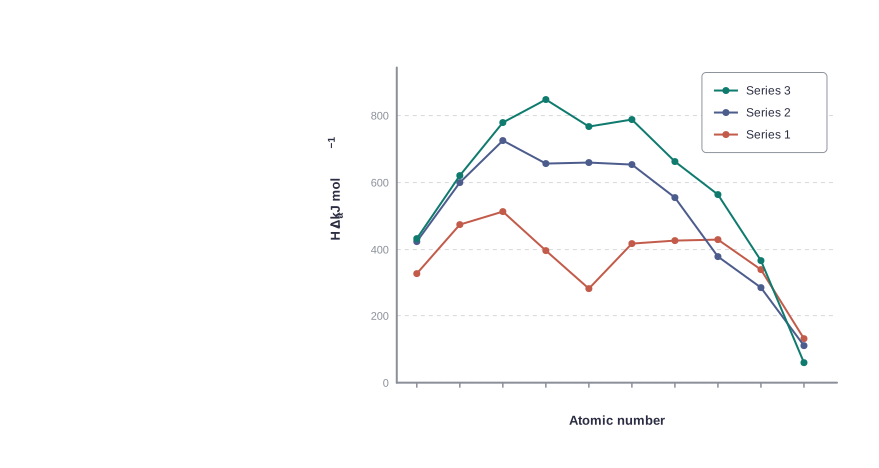
Fig. 4.4 – Trends in enthalpies of atomisation of transition elements.
The maxima at about the middle of each series indicate that one unpaired electron per d orbital is particularly favourable for strong interatomic interaction. In general, the greater the number of valence electrons available, the stronger is the resultant bonding. Since the enthalpy of atomisation is an important factor in determining the standard electrode potential of a metal, metals with a very high enthalpy of atomisation (that is, a very high boiling point) tend to be noble in their reactions — they resist being converted into ions. We shall return to this point when we take up electrode potentials.
Another generalisation that may be drawn from Fig. 4.4 is that the metals of the second and third series have greater enthalpies of atomisation than the corresponding elements of the first series. This is an important factor in accounting for the occurrence of much more frequent metal–metal bonding in compounds of the heavy transition metals.
4.3.2 Variation in Atomic and Ionic Sizes of Transition Metals
In general, ions of the same charge in a given series show a progressive decrease in radius with increasing atomic number. The reason is a tug-of-war between two effects. Each time the nuclear charge increases by unity, the new electron enters a d orbital. It may be recalled that the shielding effect of a d electron is not very effective — a d electron is poor at screening other electrons from the nucleus. Hence the net electrostatic attraction between the nuclear charge and the outermost electron increases, and the ionic radius decreases.
The same trend is observed in the atomic radii of a given series. However, the variation within a series is quite small: the added d electron does shield to some extent, so the contraction is gentle rather than dramatic (look along the metallic radii row of Table 4.2 and you will see the values crawl from 164 pm down to about 125 pm and then rise slightly again).
The Lanthanoid Contraction
An interesting point emerges when the atomic sizes of one series are compared with those of the corresponding elements in the other series. The curves in Fig. 4.5 show an increase from the first (3d) to the second (4d) series of elements, but the radii of the third (5d) series are virtually the same as those of the corresponding members of the second series.
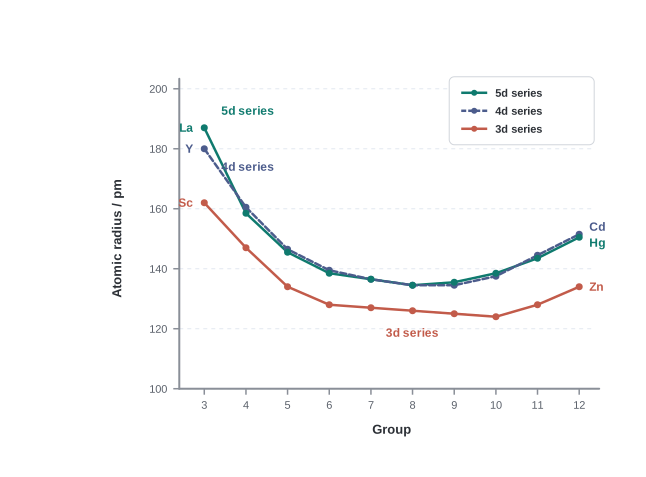
Fig. 4.5 – Trends in atomic radii of transition elements.
This phenomenon is associated with the intervention of the 4f orbitals, which must be filled before the 5d series of elements begins. The filling of 4f before the 5d orbital results in a regular decrease in atomic radii called lanthanoid contraction, which essentially compensates for the expected increase in atomic size with increasing atomic number.
The net result of the lanthanoid contraction is that the second and the third d series exhibit similar radii (for example, Zr 160 pm, Hf 159 pm) and have very similar physical and chemical properties — much more similar than one would expect on the basis of the usual family relationship between an element and the one below it.
The factor responsible for the lanthanoid contraction is somewhat similar to that observed in an ordinary transition series and is attributed to a similar cause: the imperfect shielding of one electron by another in the same set of orbitals. However, the shielding of one 4f electron by another is even less than that of one d electron by another, and as the nuclear charge increases along the series, there is a fairly regular decrease in the size of the entire \(4f^{\,n}\) orbitals.
Density Across the Series
The decrease in metallic radius, coupled with an increase in atomic mass, results in a general increase in the density of these elements. Density is mass per unit volume, so heavier atoms packed into smaller volumes must give a denser metal. Thus, from titanium (\(Z = 22\)) to copper (\(Z = 29\)), the significant increase in the density may be noted in Table 4.2 — from \(4.1\ \text{g cm}^{-3}\) to \(8.9\ \text{g cm}^{-3}\).
Table 4.2: Electronic Configurations and some other Properties of the First Series of Transition Elements
| Element | Sc | Ti | V | Cr | Mn | Fe | Co | Ni | Cu | Zn |
|---|---|---|---|---|---|---|---|---|---|---|
| Atomic number | 21 | 22 | 23 | 24 | 25 | 26 | 27 | 28 | 29 | 30 |
| Configuration, M | \(3d^14s^2\) | \(3d^24s^2\) | \(3d^34s^2\) | \(3d^54s^1\) | \(3d^54s^2\) | \(3d^64s^2\) | \(3d^74s^2\) | \(3d^84s^2\) | \(3d^{10}4s^1\) | \(3d^{10}4s^2\) |
| Configuration, M⁺ | \(3d^14s^1\) | \(3d^24s^1\) | \(3d^34s^1\) | \(3d^5\) | \(3d^54s^1\) | \(3d^64s^1\) | \(3d^74s^1\) | \(3d^84s^1\) | \(3d^{10}\) | \(3d^{10}4s^1\) |
| Configuration, M²⁺ | \(3d^1\) | \(3d^2\) | \(3d^3\) | \(3d^4\) | \(3d^5\) | \(3d^6\) | \(3d^7\) | \(3d^8\) | \(3d^9\) | \(3d^{10}\) |
| Configuration, M³⁺ | [Ar] | \(3d^1\) | \(3d^2\) | \(3d^3\) | \(3d^4\) | \(3d^5\) | \(3d^6\) | \(3d^7\) | – | – |
| Enthalpy of atomisation, \(\Delta_a H^\circ\)/kJ mol⁻¹ | 326 | 473 | 515 | 397 | 281 | 416 | 425 | 430 | 339 | 126 |
| Ionisation enthalpy \(\Delta_i H^\circ\) I /kJ mol⁻¹ | 631 | 656 | 650 | 653 | 717 | 762 | 758 | 736 | 745 | 906 |
| \(\Delta_i H^\circ\) II /kJ mol⁻¹ | 1235 | 1309 | 1414 | 1592 | 1509 | 1561 | 1644 | 1752 | 1958 | 1734 |
| \(\Delta_i H^\circ\) III /kJ mol⁻¹ | 2393 | 2657 | 2833 | 2990 | 3260 | 2962 | 3243 | 3402 | 3556 | 3837 |
| Metallic radius M /pm | 164 | 147 | 135 | 129 | 137 | 126 | 125 | 125 | 128 | 137 |
| Ionic radius M²⁺ /pm | – | – | 79 | 82 | 82 | 77 | 74 | 70 | 73 | 75 |
| Ionic radius M³⁺ /pm | 73 | 67 | 64 | 62 | 65 | 65 | 61 | 60 | – | – |
| Standard electrode potential \(E^\circ\)(M²⁺/M) /V | – | –1.63 | –1.18 | –0.90 | –1.18 | –0.44 | –0.28 | –0.25 | +0.34 | –0.76 |
| Standard electrode potential \(E^\circ\)(M³⁺/M²⁺) /V | – | –0.37 | –0.26 | –0.41 | +1.57 | +0.77 | +1.97 | – | – | – |
| Density /g cm⁻³ | 3.43 | 4.1 | 6.07 | 7.19 | 7.21 | 7.8 | 8.7 | 8.9 | 8.9 | 7.1 |
4.3.3 Ionisation Enthalpies
There is an increase in ionisation enthalpy along each series of the transition elements from left to right, due to an increase in nuclear charge which accompanies the filling of the inner d orbitals. Table 4.2 gives the values of the first three ionisation enthalpies of the first series of transition elements.
These values show that the successive enthalpies of these elements do not increase as steeply as in the case of non-transition elements. Also, the variation in ionisation enthalpy along a series of transition elements is much less in comparison to the variation along a period of non-transition elements. The first ionisation enthalpy, in general, increases; but the magnitude of the increase in the second and third ionisation enthalpies for the successive elements is much higher along a series.
Why the First Ionisation Enthalpy Wanders
The irregular trend in the first ionisation enthalpy of the metals of the 3d series, though of little chemical significance, can be accounted for by considering that the removal of one electron alters the relative energies of the 4s and 3d orbitals.
You have learnt that when d-block elements form ions, the \(ns\) electrons are lost before the \((n-1)d\) electrons. As we move along the period in the 3d series, we see that the nuclear charge increases from scandium to zinc, but electrons are added to the orbital of an inner subshell, i.e., the 3d orbitals. These 3d electrons shield the 4s electrons from the increasing nuclear charge somewhat more effectively than the outer shell electrons can shield one another. Therefore, the atomic radii decrease less rapidly, and ionisation energies increase only slightly along the 3d series.
The doubly or more highly charged ions have \(d^n\) configurations with no 4s electrons. A general trend of increasing values of the second ionisation enthalpy is expected, as the effective nuclear charge increases — one d electron does not shield another electron from the influence of the nuclear charge, because d orbitals differ in direction and so the electrons in them do not get in each other’s way.
However, the trend of steady increase in second and third ionisation enthalpy breaks for the formation of \(\mathrm{Mn^{2+}}\) and \(\mathrm{Fe^{3+}}\) respectively. In both cases, the ion produced has the \(d^5\) configuration. Similar breaks occur at corresponding elements in the later transition series.
Forming the M²⁺ Ion
The lowest common oxidation state of these metals is +2. To form the \(\mathrm{M^{2+}}\) ions from the gaseous atoms, the sum of the first and second ionisation enthalpy is required, in addition to the enthalpy of atomisation (the metal must first be converted to free atoms).
The dominant term is the second ionisation enthalpy, which shows unusually high values for Cr and Cu, where the \(\mathrm{M^{+}}\) ions have the \(d^5\) and \(d^{10}\) configurations respectively — stable arrangements that resist losing a further electron. The value for Zn is correspondingly low, as the ionisation causes the removal of one 4s electron, which results in the formation of the stable \(d^{10}\) configuration.
The trend in the third ionisation enthalpies is not complicated by the 4s orbital factor, and shows the greater difficulty of removing an electron from the \(d^5\) (\(\mathrm{Mn^{2+}}\)) and \(d^{10}\) (\(\mathrm{Zn^{2+}}\)) ions. In general, the third ionisation enthalpies are quite high. Also, the high values for the third ionisation enthalpies of copper, nickel and zinc indicate why it is difficult to obtain an oxidation state greater than two for these elements.
Although ionisation enthalpies give some guidance concerning the relative stabilities of oxidation states, this problem is very complex and not amenable to ready generalisation. Hydration enthalpy, lattice enthalpy and the nature of the ligand all have a say, as we shall see shortly.
4.3.4 Oxidation States
One of the most notable features of the transition elements is the great variety of oxidation states these may show in their compounds. Table 4.3 lists the common oxidation states of the first row transition elements; the most common ones are printed in bold.
Table 4.3: Oxidation States of the first row Transition Metals (the most common ones are in bold types)
| Sc | Ti | V | Cr | Mn | Fe | Co | Ni | Cu | Zn |
|---|---|---|---|---|---|---|---|---|---|
| +3 | +2 | +2 | +2 | +2 | +2 | +2 | +2 | +1 | +2 |
| +3 | +3 | +3 | +3 | +3 | +3 | +3 | +2 | ||
| +4 | +4 | +4 | +4 | +4 | +4 | +4 | |||
| +5 | +5 | +5 | |||||||
| +6 | +6 | +6 | |||||||
| +7 |
Why the Middle of the Series is Richest
The elements which give the greatest number of oxidation states occur in or near the middle of the series. Manganese, for example, exhibits all the oxidation states from +2 to +7.
The lesser number of oxidation states at the extreme ends stems from one of two opposite causes: either too few electrons to lose or share (Sc, Ti), or too many d electrons — hence fewer orbitals available in which to share electrons with others — for higher valence (Cu, Zn).
Thus, early in the series, scandium(II) is virtually unknown, and titanium(IV) is more stable than Ti(III) or Ti(II). At the other end, the only oxidation state of zinc is +2, and no d electrons are involved in it at all.
The maximum oxidation states of reasonable stability correspond in value to the sum of the s and d electrons up to manganese: \(\mathrm{Ti^{IV}O_2}\), \(\mathrm{V^{V}O_2^{+}}\), \(\mathrm{Cr^{VI}O_4^{2-}}\), \(\mathrm{Mn^{VII}O_4^{-}}\). After manganese, there is a rather abrupt decrease in the stability of higher oxidation states, so the typical species to follow are \(\mathrm{Fe^{II,III}}\), \(\mathrm{Co^{II,III}}\), \(\mathrm{Ni^{II}}\), \(\mathrm{Cu^{I,II}}\), \(\mathrm{Zn^{II}}\).
Steps of One, Not Two
The variability of oxidation states, a characteristic of transition elements, arises out of the incomplete filling of d orbitals in such a way that their oxidation states differ from each other by unity — for example, \(\mathrm{V^{II}}\), \(\mathrm{V^{III}}\), \(\mathrm{V^{IV}}\), \(\mathrm{V^{V}}\). This is in contrast with the variability of oxidation states of non-transition elements, where oxidation states normally differ by a unit of two (think of \(\mathrm{Sn^{2+}}\) and \(\mathrm{Sn^{4+}}\), or \(\mathrm{Pb^{2+}}\) and \(\mathrm{Pb^{4+}}\)).
An interesting feature in the variability of oxidation states of the d-block elements is noticed among the groups (groups 4 through 10). Although in the p-block the lower oxidation states are favoured by the heavier members (due to the inert pair effect), the opposite is true in the groups of the d-block. For example, in group 6, Mo(VI) and W(VI) are found to be more stable than Cr(VI). Thus Cr(VI) in the form of dichromate in acidic medium is a strong oxidising agent, whereas \(\mathrm{MoO_3}\) and \(\mathrm{WO_3}\) are not.
Low oxidation states are found when a complex compound has ligands capable of \(\pi\)-acceptor character in addition to the \(\sigma\)-bonding. Such ligands can drain electron density away from the metal, allowing it to hold electrons it would otherwise lose. For example, in \(\mathrm{Ni(CO)_4}\) and \(\mathrm{Fe(CO)_5}\), the oxidation state of nickel and iron is zero.
4.3.5 Trends in the M²⁺/M Standard Electrode Potentials
Table 4.4 contains the thermochemical parameters related to the transformation of the solid metal atoms to \(\mathrm{M^{2+}}\) ions in solution, and their standard electrode potentials. Three quantities decide the outcome: the enthalpy of atomisation (turning the solid metal into gaseous atoms), the sum of the first two ionisation enthalpies (turning those atoms into \(\mathrm{M^{2+}}\) ions), and the hydration enthalpy (the energy released when the gaseous ion is surrounded by water molecules). Only the balance of all three tells us how readily the metal will go into solution as an ion.
The observed values of \(E^{\ominus}\) and those calculated using the data of Table 4.4 are compared in Fig. 4.6.
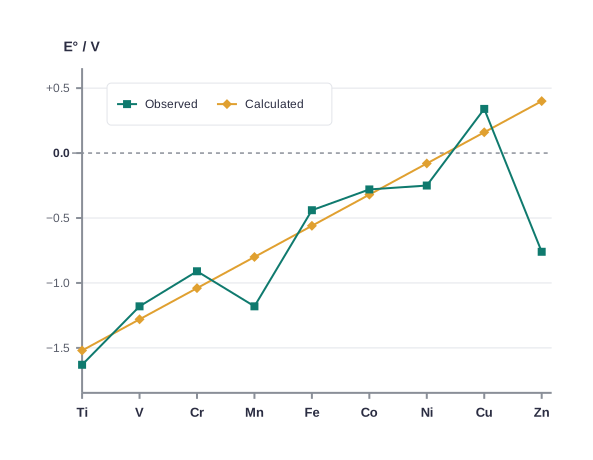
Fig. 4.6 – Observed and calculated standard electrode potentials for the 3d series.
The unique behaviour of Cu, having a positive \(E^{\ominus}\), accounts for its inability to liberate \(\mathrm{H_2}\) from acids. Only oxidising acids (nitric and hot concentrated sulphuric) react with Cu, the acids themselves being reduced. The high energy needed to transform \(\mathrm{Cu(s)}\) to \(\mathrm{Cu^{2+}(aq)}\) is not balanced by its hydration enthalpy.
The general trend towards less negative \(E^{\ominus}\) values across the series is related to the general increase in the sum of the first and second ionisation enthalpies. It is interesting to note that the values of \(E^{\ominus}\) for Mn, Ni and Zn are more negative than expected from the trend.
Table 4.4: Thermochemical data (kJ mol⁻¹) for the first row Transition Elements and the Standard Electrode Potentials for the Reduction of MII to M
| Element (M) | \(\Delta_a H^\circ\) (M) | \(\Delta_i H_1^\circ\) | \(\Delta_i H_2^\circ\) | \(\Delta_{\text{hyd}} H^\circ\)(M²⁺) | \(E^\circ\)/V |
|---|---|---|---|---|---|
| Ti | 469 | 656 | 1309 | –1866 | –1.63 |
| V | 515 | 650 | 1414 | –1895 | –1.18 |
| Cr | 398 | 653 | 1592 | –1925 | –0.90 |
| Mn | 279 | 717 | 1509 | –1862 | –1.18 |
| Fe | 418 | 762 | 1561 | –1998 | –0.44 |
| Co | 427 | 758 | 1644 | –2079 | –0.28 |
| Ni | 431 | 736 | 1752 | –2121 | –0.25 |
| Cu | 339 | 745 | 1958 | –2121 | 0.34 |
| Zn | 130 | 906 | 1734 | –2059 | –0.76 |
The stability of the half-filled d sub-shell in \(\mathrm{Mn^{2+}}\) and the completely filled \(d^{10}\) configuration in \(\mathrm{Zn^{2+}}\) are related to their \(E^\circ\) values, whereas \(E^\circ\) for Ni is related to its highest negative \(\Delta_{\text{hyd}} H^\circ\).
4.3.6 Trends in the M³⁺/M²⁺ Standard Electrode Potentials
An examination of the \(E^\circ\)(M³⁺/M²⁺) values (Table 4.2) shows the varying trends. Read each value as a measure of how badly the +3 ion wants to grab an electron and drop to +2.
The low value for Sc reflects the stability of \(\mathrm{Sc^{3+}}\), which has a noble gas configuration and has no wish to accept an electron back. The highest value for Zn is due to the removal of an electron from the stable \(d^{10}\) configuration of \(\mathrm{Zn^{2+}}\) — a change so unfavourable that \(\mathrm{Zn^{3+}}\) is effectively unobtainable.
The comparatively high value for Mn (+1.57 V) shows that \(\mathrm{Mn^{2+}}\) (\(d^5\)) is particularly stable, so \(\mathrm{Mn^{3+}}\) readily takes an electron to become it. The comparatively low value for Fe (+0.77 V) shows the extra stability of \(\mathrm{Fe^{3+}}\) (\(d^5\)), which is in no hurry to be reduced. The comparatively low value for V is related to the stability of \(\mathrm{V^{2+}}\) (half-filled \(t_{2g}\) level, Chapter 5).
4.3.7 Trends in Stability of Higher Oxidation States
Table 4.5 shows the stable halides of the 3d series of transition metals. The highest oxidation numbers are achieved in \(\mathrm{TiX_4}\) (tetrahalides), \(\mathrm{VF_5}\) and \(\mathrm{CrF_6}\).
The +7 state for Mn is not represented in simple halides, but \(\mathrm{MnO_3F}\) is known; and beyond Mn no metal has a trihalide except \(\mathrm{FeX_3}\) and \(\mathrm{CoF_3}\).
The ability of fluorine to stabilise the highest oxidation state is due to either higher lattice energy, as in the case of \(\mathrm{CoF_3}\), or higher bond enthalpy terms for the higher covalent compounds, e.g., \(\mathrm{VF_5}\) and \(\mathrm{CrF_6}\). Fluorine is small and the most electronegative of all elements, so it can crowd around a metal in large numbers and hold on to electrons tightly — exactly what a metal in a very high oxidation state requires.
Although \(\mathrm{V^{+5}}\) is represented only by \(\mathrm{VF_5}\), the other halides, however, undergo hydrolysis to give oxohalides, \(\mathrm{VOX_3}\). Another feature of the halides is their instability in the low oxidation states, e.g., \(\mathrm{VX_2}\) (X = Cl, Br or I), and the same applies to \(\mathrm{CuX}\).
Table 4.5: Formulas of Halides of 3d Metals
| Oxidation Number | |||||||||
|---|---|---|---|---|---|---|---|---|---|
| +6 | \(\mathrm{CrF_6}\) | ||||||||
| +5 | \(\mathrm{VF_5}\) | \(\mathrm{CrF_5}\) | |||||||
| +4 | \(\mathrm{TiX_4}\) | \(\mathrm{VX^{I}_4}\) | \(\mathrm{CrX_4}\) | \(\mathrm{MnF_4}\) | |||||
| +3 | \(\mathrm{TiX_3}\) | \(\mathrm{VX_3}\) | \(\mathrm{CrX_3}\) | \(\mathrm{MnF_3}\) | \(\mathrm{FeX^{I}_3}\) | \(\mathrm{CoF_3}\) | |||
| +2 | \(\mathrm{TiX_2^{III}}\) | \(\mathrm{VX_2}\) | \(\mathrm{CrX_2}\) | \(\mathrm{MnX_2}\) | \(\mathrm{FeX_2}\) | \(\mathrm{CoX_2}\) | \(\mathrm{NiX_2}\) | \(\mathrm{CuX_2^{II}}\) | \(\mathrm{ZnX_2}\) |
| +1 | \(\mathrm{CuX^{III}}\) |
Key: X = F → I; XI = F → Br; XII = F, Cl; XIII = Cl → I
Copper: Why Cu(I) Will Not Survive in Water
On the other hand, all \(\mathrm{Cu^{II}}\) halides are known except the iodide. In this case, \(\mathrm{Cu^{2+}}\) oxidises \(\mathrm{I^-}\) to \(\mathrm{I_2}\):
\[2\,\mathrm{Cu^{2+}} + 4\,\mathrm{I^{-}} \rightarrow \mathrm{Cu_2I_2}\text{(s)} + \mathrm{I_2}\]
However, many copper(I) compounds are unstable in aqueous solution and undergo disproportionation — the same species is simultaneously oxidised and reduced:
\[2\,\mathrm{Cu^{+}} \rightarrow \mathrm{Cu^{2+}} + \mathrm{Cu}\]
The stability of \(\mathrm{Cu^{2+}}\)(aq) rather than \(\mathrm{Cu^{+}}\)(aq) is due to the much more negative \(\Delta_{\text{hyd}} H^\circ\) of \(\mathrm{Cu^{2+}}\)(aq) than \(\mathrm{Cu^{+}}\), which more than compensates for the second ionisation enthalpy of Cu. The doubly charged ion is smaller and more highly charged, so water molecules bind to it far more strongly, and that release of hydration energy pays the extra ionisation bill.
Oxygen Beats Fluorine at the Top
The ability of oxygen to stabilise the highest oxidation state is demonstrated in the oxides. The highest oxidation number in the oxides (Table 4.6) coincides with the group number and is attained in \(\mathrm{Sc_2O_3}\) to \(\mathrm{Mn_2O_7}\).
Beyond Group 7, no higher oxides of Fe above \(\mathrm{Fe_2O_3}\) are known, although ferrates(VI), \(\mathrm{(FeO_4)^{2-}}\), are formed in alkaline media, but they readily decompose to \(\mathrm{Fe_2O_3}\) and \(\mathrm{O_2}\).
Besides the oxides, oxocations stabilise \(\mathrm{V^{V}}\) as \(\mathrm{VO_2^{+}}\), \(\mathrm{V^{IV}}\) as \(\mathrm{VO^{2+}}\) and \(\mathrm{Ti^{IV}}\) as \(\mathrm{TiO^{2+}}\).
The ability of oxygen to stabilise these high oxidation states exceeds that of fluorine. Thus the highest Mn fluoride is \(\mathrm{MnF_4}\), whereas the highest oxide is \(\mathrm{Mn_2O_7}\). The ability of oxygen to form multiple bonds to metals explains its superiority: one oxygen atom can use two of its electron pairs on the metal, doing the work of two fluorine atoms.
In the covalent oxide \(\mathrm{Mn_2O_7}\), each Mn is tetrahedrally surrounded by O’s, including a Mn–O–Mn bridge, as shown in Fig. 4.7. The tetrahedral \(\mathrm{[MO_4]^{n-}}\) ions are known for \(\mathrm{V^{V}}\), \(\mathrm{Cr^{VI}}\), \(\mathrm{Mn^{V}}\), \(\mathrm{Mn^{VI}}\) and \(\mathrm{Mn^{VII}}\).
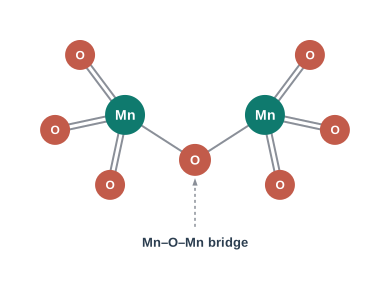
Fig. 4.7 – Structure of Mn₂O₇.
Table 4.6: Oxides of 3d Metals
| Oxidation Number | Group 3 | 4 | 5 | 6 | 7 | 8 | 9 | 10 | 11 | 12 |
|---|---|---|---|---|---|---|---|---|---|---|
| +7 | \(\mathrm{Mn_2O_7}\) | |||||||||
| +6 | \(\mathrm{CrO_3}\) | |||||||||
| +5 | \(\mathrm{V_2O_5}\) | |||||||||
| +4 | \(\mathrm{TiO_2}\) | \(\mathrm{V_2O_4}\) | \(\mathrm{CrO_2}\) | \(\mathrm{MnO_2}\) | ||||||
| +3 | \(\mathrm{Sc_2O_3}\) | \(\mathrm{Ti_2O_3}\) | \(\mathrm{V_2O_3}\) | \(\mathrm{Cr_2O_3}\) | \(\mathrm{Mn_2O_3}\), \(\mathrm{Mn_3O_4}\)* | \(\mathrm{Fe_2O_3}\), \(\mathrm{Fe_3O_4}\)* | \(\mathrm{Co_3O_4}\)* | |||
| +2 | \(\mathrm{TiO}\) | \(\mathrm{VO}\) | \(\mathrm{(CrO)}\) | \(\mathrm{MnO}\) | \(\mathrm{FeO}\) | \(\mathrm{CoO}\) | \(\mathrm{NiO}\) | \(\mathrm{CuO}\) | \(\mathrm{ZnO}\) | |
| +1 | \(\mathrm{Cu_2O}\) |
* mixed oxides
4.3.8 Chemical Reactivity and E° Values
Transition metals vary widely in their chemical reactivity. Many of them are sufficiently electropositive to dissolve in mineral acids, although a few are ‘noble’ — that is, they are unaffected by single acids.
The metals of the first series, with the exception of copper, are relatively more reactive and are oxidised by \(1\ \mathrm{M}\ \mathrm{H^{+}}\), though the actual rate at which these metals react with oxidising agents like the hydrogen ion (\(\mathrm{H^{+}}\)) is sometimes slow. For example, titanium and vanadium, in practice, are passive to dilute non-oxidising acids at room temperature — a protective oxide film on the surface keeps the acid away, even though the reaction is thermodynamically allowed.
The \(E^\circ\) values for M²⁺/M (Table 4.2) indicate a decreasing tendency to form divalent cations across the series. This general trend towards less negative \(E^\circ\) values is related to the increase in the sum of the first and second ionisation enthalpies. It is interesting to note that the \(E^\circ\) values for Mn, Ni and Zn are more negative than expected from the general trend. Whereas the stabilities of the half-filled d subshell (\(d^5\)) in \(\mathrm{Mn^{2+}}\) and the completely filled d subshell (\(d^{10}\)) in zinc are related to their \(E^\circ\) values, for nickel the \(E^\circ\) value is related to the highest negative enthalpy of hydration.
An examination of the \(E^\circ\) values for the redox couple M³⁺/M²⁺ (Table 4.2) shows that \(\mathrm{Mn^{3+}}\) and \(\mathrm{Co^{3+}}\) ions are the strongest oxidising agents in aqueous solutions. The ions \(\mathrm{Ti^{2+}}\), \(\mathrm{V^{2+}}\) and \(\mathrm{Cr^{2+}}\) are strong reducing agents and will liberate hydrogen from a dilute acid, e.g.,
\[2\,\mathrm{Cr^{2+}}\text{(aq)} + 2\,\mathrm{H^{+}}\text{(aq)} \rightarrow 2\,\mathrm{Cr^{3+}}\text{(aq)} + \mathrm{H_2}\text{(g)}\]
4.3.9 Magnetic Properties
When a magnetic field is applied to substances, mainly two types of magnetic behaviour are observed: diamagnetism and paramagnetism. Diamagnetic substances are repelled by the applied field, while paramagnetic substances are attracted to it. Substances which are attracted very strongly are said to be ferromagnetic. In fact, ferromagnetism is an extreme form of paramagnetism. Many of the transition metal ions are paramagnetic.
Paramagnetism arises from the presence of unpaired electrons, each such electron having a magnetic moment associated with its spin angular momentum and its orbital angular momentum. For the compounds of the first series of transition metals, the contribution of the orbital angular momentum is effectively quenched and hence is of no significance. For these, the magnetic moment is determined by the number of unpaired electrons and is calculated by using the ‘spin-only’ formula:
\[\mu = \sqrt{n(n+2)}\ \text{BM}\]
where \(n\) is the number of unpaired electrons and \(\mu\) is the magnetic moment in units of Bohr magneton (BM). A single unpaired electron has a magnetic moment of 1.73 Bohr magnetons (BM).
The magnetic moment increases with the increasing number of unpaired electrons. Thus, the observed magnetic moment gives a useful indication about the number of unpaired electrons present in the atom, molecule or ion — measure the magnetism, and you can work backwards to the electronic configuration. The magnetic moments calculated from the ‘spin-only’ formula and those derived experimentally for some ions of the first row transition elements are given in Table 4.7. The experimental data are mainly for hydrated ions in solution or in the solid state.
Table 4.7: Calculated and Observed Magnetic Moments (BM)
| Ion | Configuration | Unpaired electron(s) | Magnetic moment (Calculated) | Magnetic moment (Observed) |
|---|---|---|---|---|
| \(\mathrm{Sc^{3+}}\) | \(3d^0\) | 0 | 0 | 0 |
| \(\mathrm{Ti^{3+}}\) | \(3d^1\) | 1 | 1.73 | 1.75 |
| \(\mathrm{Ti^{2+}}\) | \(3d^2\) | 2 | 2.84 | 2.76 |
| \(\mathrm{V^{2+}}\) | \(3d^3\) | 3 | 3.87 | 3.86 |
| \(\mathrm{Cr^{2+}}\) | \(3d^4\) | 4 | 4.90 | 4.80 |
| \(\mathrm{Mn^{2+}}\) | \(3d^5\) | 5 | 5.92 | 5.96 |
| \(\mathrm{Fe^{2+}}\) | \(3d^6\) | 4 | 4.90 | 5.3 – 5.5 |
| \(\mathrm{Co^{2+}}\) | \(3d^7\) | 3 | 3.87 | 4.4 – 5.2 |
| \(\mathrm{Ni^{2+}}\) | \(3d^8\) | 2 | 2.84 | 2.9 – 3.4 |
| \(\mathrm{Cu^{2+}}\) | \(3d^9\) | 1 | 1.73 | 1.8 – 2.2 |
| \(\mathrm{Zn^{2+}}\) | \(3d^{10}\) | 0 | 0 | — |
4.3.10 Formation of Coloured Ions
When an electron from a lower energy d orbital is excited to a higher energy d orbital, the energy of excitation corresponds to the frequency of light absorbed (Chapter 5). This frequency generally lies in the visible region. The colour observed corresponds to the complementary colour of the light absorbed — the eye sees what is left over after the absorbed portion of white light has been removed.
The frequency of the light absorbed is determined by the nature of the ligand, that is, by the species attached to the metal ion. In aqueous solutions, where water molecules are the ligands, the colours of the ions observed are listed in Table 4.8. A few coloured solutions of d-block elements are illustrated in Fig. 4.8.
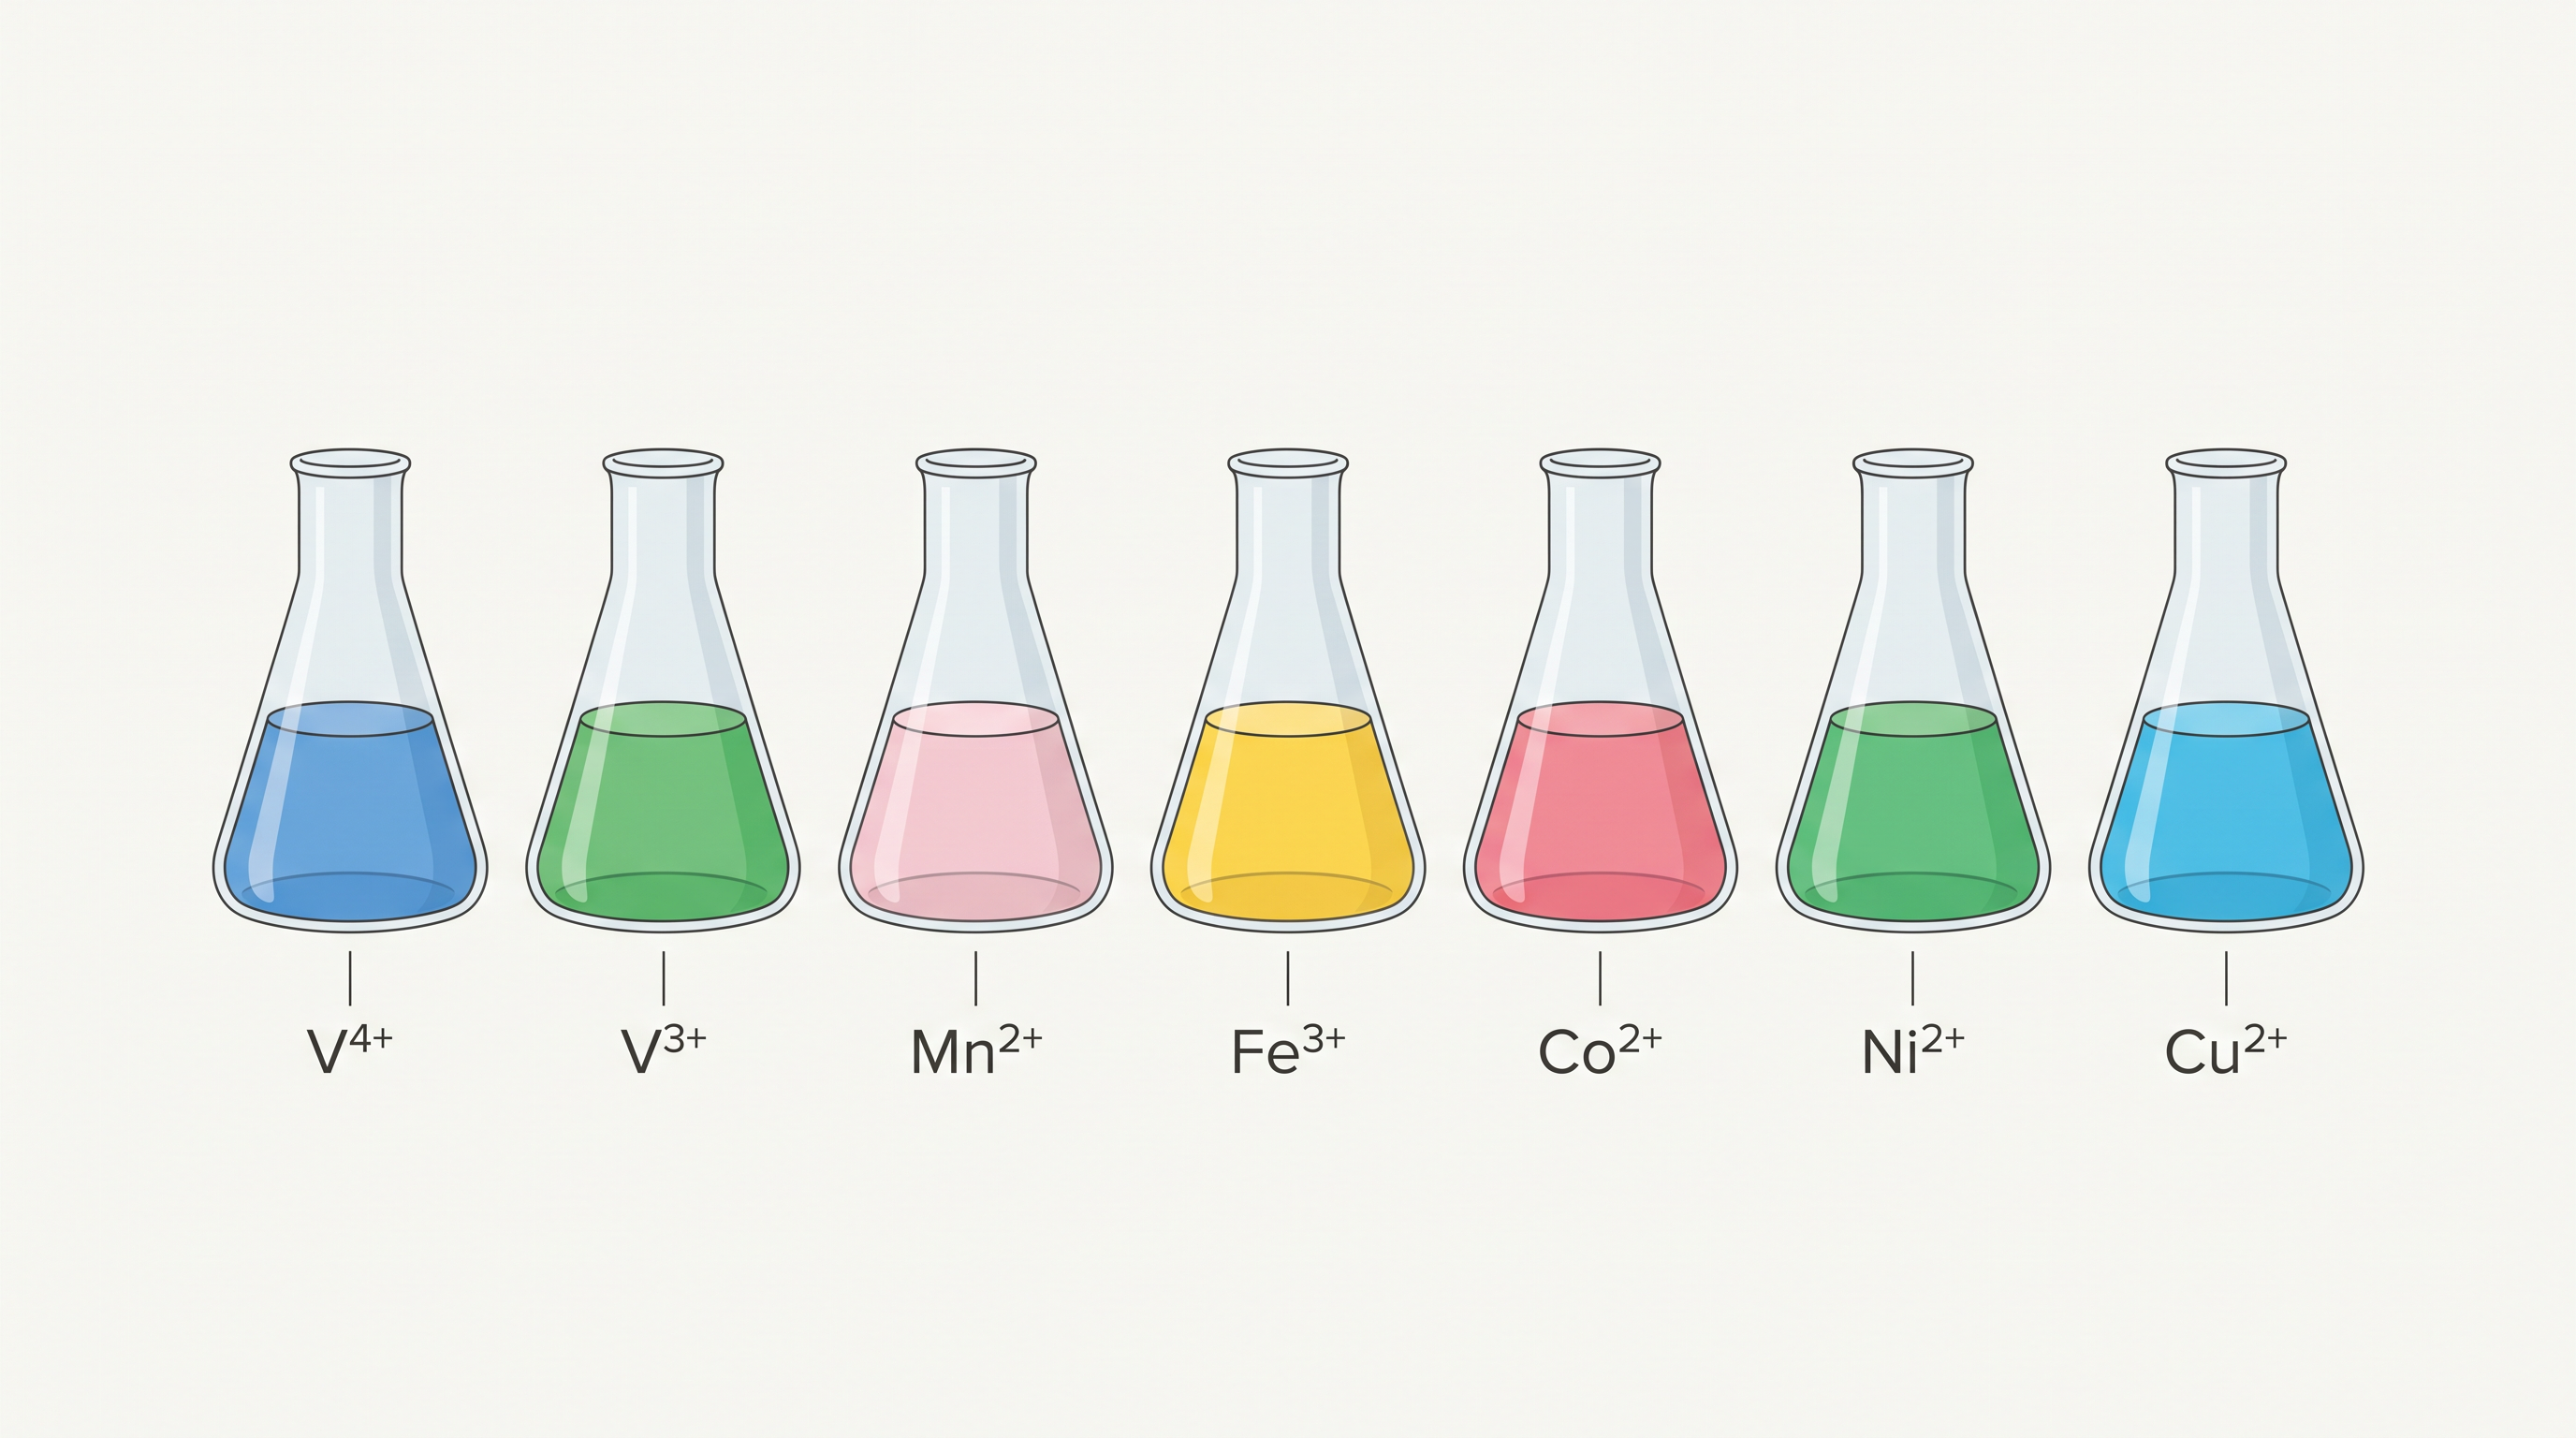
Fig. 4.8 – Colours of first-row transition metal ions in aqueous solution.
Table 4.8: Colours of Some of the First Row (aquated) Transition Metal Ions
| Configuration | Example | Colour |
|---|---|---|
| \(3d^0\) | \(\mathrm{Sc^{3+}}\) | colourless |
| \(3d^0\) | \(\mathrm{Ti^{4+}}\) | colourless |
| \(3d^1\) | \(\mathrm{Ti^{3+}}\) | purple |
| \(3d^1\) | \(\mathrm{V^{4+}}\) | blue |
| \(3d^2\) | \(\mathrm{V^{3+}}\) | green |
| \(3d^3\) | \(\mathrm{V^{2+}}\) | violet |
| \(3d^3\) | \(\mathrm{Cr^{3+}}\) | violet |
| \(3d^4\) | \(\mathrm{Mn^{3+}}\) | violet |
| \(3d^4\) | \(\mathrm{Cr^{2+}}\) | blue |
| \(3d^5\) | \(\mathrm{Mn^{2+}}\) | pink |
| \(3d^5\) | \(\mathrm{Fe^{3+}}\) | yellow |
| \(3d^6\) | \(\mathrm{Fe^{2+}}\) | green |
| \(3d^6\,3d^7\) | \(\mathrm{Co^{3+}}\), \(\mathrm{Co^{2+}}\) | blue, pink |
| \(3d^8\) | \(\mathrm{Ni^{2+}}\) | green |
| \(3d^9\) | \(\mathrm{Cu^{2+}}\) | blue |
| \(3d^{10}\) | \(\mathrm{Zn^{2+}}\) | colourless |
4.3.11 Formation of Complex Compounds
Complex compounds are those in which the metal ions bind a number of anions or neutral molecules, giving complex species with characteristic properties. A few examples are: \(\mathrm{[Fe(CN)_6]^{3-}}\), \(\mathrm{[Fe(CN)_6]^{4-}}\), \(\mathrm{[Cu(NH_3)_4]^{2+}}\) and \(\mathrm{[PtCl_4]^{2-}}\). (The chemistry of complex compounds is dealt with in detail in Chapter 5.)
The transition metals form a large number of complex compounds. This is due to three things working together: the comparatively smaller sizes of the metal ions, their high ionic charges, and the availability of d orbitals for bond formation. A small, highly charged ion attracts the incoming ligands strongly, and the empty d orbitals give the ion somewhere to accept the ligands’ lone pairs.
4.3.12 Catalytic Properties
The transition metals and their compounds are known for their catalytic activity. This activity is ascribed to their ability to adopt multiple oxidation states and to form complexes. Vanadium(V) oxide (in the Contact Process), finely divided iron (in Haber’s Process), and nickel (in Catalytic Hydrogenation) are some of the examples.
Catalysts at a solid surface involve the formation of bonds between reactant molecules and atoms of the surface of the catalyst (first row transition metals utilise 3d and 4s electrons for bonding). This has the effect of increasing the concentration of the reactants at the catalyst surface, and also of weakening the bonds in the reacting molecules — the activation energy is lowered. A reaction with a lower activation energy has a much larger fraction of molecules able to cross the barrier, and so proceeds faster.
Also, because the transition metal ions can change their oxidation states, they become more effective as catalysts. For example, iron(III) catalyses the reaction between iodide and persulphate ions:
\[2\,\mathrm{I^{-}} + \mathrm{S_2O_8^{2-}} \rightarrow \mathrm{I_2} + 2\,\mathrm{SO_4^{2-}}\]
An explanation of this catalytic action can be given as:
\[2\,\mathrm{Fe^{3+}} + 2\,\mathrm{I^{-}} \rightarrow 2\,\mathrm{Fe^{2+}} + \mathrm{I_2}\]
\[2\,\mathrm{Fe^{2+}} + \mathrm{S_2O_8^{2-}} \rightarrow 2\,\mathrm{Fe^{3+}} + 2\,\mathrm{SO_4^{2-}}\]
Add the two steps and the iron cancels out: the catalyst is regenerated, exactly as a catalyst must be. The cycle is summarised in FC 4.1.
Flowchart
FC 4.1 – Catalytic cycle of Fe³⁺ in the iodide–persulphate reaction: Fe³⁺ + I⁻ → Fe²⁺ + I₂ (iron reduced, iodide oxidised) → Fe²⁺ + S₂O₈²⁻ → Fe³⁺ + SO₄²⁻ (iron re-oxidised, persulphate reduced) → back to start; net reaction shown outside the cycle: 2I⁻ + S₂O₈²⁻ → I₂ + 2SO₄²⁻
4.3.13 Formation of Interstitial Compounds
Interstitial compounds are those which are formed when small atoms like H, C or N are trapped inside the crystal lattices of metals. The metal atoms in a crystal do not fill space completely; there are small gaps — interstices — between them, and a small enough atom can slip in.
They are usually non-stoichiometric and are neither typically ionic nor covalent; for example, TiC, \(\mathrm{Mn_4N}\), \(\mathrm{Fe_3H}\), \(\mathrm{VH_{0.56}}\) and \(\mathrm{TiH_{1.7}}\). Look at those last two formulae carefully: the fractional subscripts are not a mistake. The formulae quoted do not, of course, correspond to any normal oxidation state of the metal, because the small atoms simply occupy whatever holes are available. Because of the nature of their composition, these compounds are referred to as interstitial compounds.
The principal physical and chemical characteristics of these compounds are as follows:
- They have high melting points, higher than those of the pure metals.
- They are very hard; some borides approach diamond in hardness.
- They retain metallic conductivity.
- They are chemically inert.
The trapped atoms act as wedges, locking the metal layers so that they cannot slide over one another — which is precisely why the interstitial compound is harder and higher-melting than the metal it came from.
4.3.14 Alloy Formation
An alloy is a blend of metals prepared by mixing the components. Alloys may be homogeneous solid solutions in which the atoms of one metal are distributed randomly among the atoms of the other.
Such alloys are formed by atoms with metallic radii that are within about 15 per cent of each other — if the two kinds of atom are too different in size, one simply will not fit into the other’s lattice. Because of similar radii and other characteristics of transition metals, alloys are readily formed by these metals. The alloys so formed are hard and often have high melting points.
The best known are the ferrous alloys: chromium, vanadium, tungsten, molybdenum and manganese are used for the production of a variety of steels and stainless steel. Alloys of transition metals with non-transition metals, such as brass (copper–zinc) and bronze (copper–tin), are also of considerable industrial importance.
4.4 Some Important Compounds of Transition Elements
4.4.1 Oxides and Oxoanions of Metals
These oxides are generally formed by the reaction of metals with oxygen at high temperatures. All the metals except scandium form MO oxides, which are ionic. As we have already seen, the highest oxidation number in the oxides coincides with the group number and is attained in \(\mathrm{Sc_2O_3}\) to \(\mathrm{Mn_2O_7}\). Beyond group 7, no higher oxides of iron above \(\mathrm{Fe_2O_3}\) are known. Besides the oxides, the oxocations stabilise \(\mathrm{V^{V}}\) as \(\mathrm{VO_2^{+}}\), \(\mathrm{V^{IV}}\) as \(\mathrm{VO^{2+}}\) and \(\mathrm{Ti^{IV}}\) as \(\mathrm{TiO^{2+}}\).
From Ionic and Basic to Covalent and Acidic
As the oxidation number of a metal increases, the ionic character of its oxide decreases. The reason is straightforward: a metal in a high oxidation state is small and highly charged, so it pulls the electron cloud of the oxide ion strongly towards itself and the bond becomes largely covalent.
In the case of Mn, \(\mathrm{Mn_2O_7}\) is a covalent green oil — hardly what one expects of a “metal oxide”. Even \(\mathrm{CrO_3}\) and \(\mathrm{V_2O_5}\) have low melting points, another sign of covalent, molecular character. In these higher oxides, the acidic character is predominant: they react with water to give acids and with alkalies to give salts.
Thus, \(\mathrm{Mn_2O_7}\) gives \(\mathrm{HMnO_4}\) and \(\mathrm{CrO_3}\) gives \(\mathrm{H_2CrO_4}\) and \(\mathrm{H_2Cr_2O_7}\).
\(\mathrm{V_2O_5}\) is, however, amphoteric — it can behave as an acid or a base — though mainly acidic, and it gives \(\mathrm{VO_4^{3-}}\) as well as \(\mathrm{VO_2^{+}}\) salts. In vanadium there is a gradual change from the basic \(\mathrm{V_2O_3}\) to the less basic \(\mathrm{V_2O_4}\) and to the amphoteric \(\mathrm{V_2O_5}\). \(\mathrm{V_2O_4}\) dissolves in acids to give \(\mathrm{VO^{2+}}\) salts. Similarly, \(\mathrm{V_2O_5}\) reacts with alkalies as well as acids to give \(\mathrm{VO_4^{3-}}\) and \(\mathrm{VO_4^{+}}\) respectively. The well characterised CrO is basic, but \(\mathrm{Cr_2O_3}\) is amphoteric.
Potassium dichromate, K₂Cr₂O₇
Potassium dichromate is a very important chemical, used in the leather industry and as an oxidant for the preparation of many azo compounds.
Dichromates are generally prepared from chromate, which in turn is obtained by the fusion of chromite ore (\(\mathrm{FeCr_2O_4}\)) with sodium or potassium carbonate in free access of air. The reaction with sodium carbonate occurs as follows:
\[4\,\mathrm{FeCr_2O_4} + 8\,\mathrm{Na_2CO_3} + 7\,\mathrm{O_2} \rightarrow 8\,\mathrm{Na_2CrO_4} + 2\,\mathrm{Fe_2O_3} + 8\,\mathrm{CO_2}\]
The yellow solution of sodium chromate is filtered and acidified with sulphuric acid to give a solution from which orange sodium dichromate, \(\mathrm{Na_2Cr_2O_7\cdot 2H_2O}\), can be crystallised.
\[2\,\mathrm{Na_2CrO_4} + 2\,\mathrm{H^{+}} \rightarrow \mathrm{Na_2Cr_2O_7} + 2\,\mathrm{Na^{+}} + \mathrm{H_2O}\]
Sodium dichromate is more soluble than potassium dichromate. The latter is therefore prepared by treating the solution of sodium dichromate with potassium chloride — the less soluble salt is the one that leaves the solution.
\[\mathrm{Na_2Cr_2O_7} + 2\,\mathrm{KCl} \rightarrow \mathrm{K_2Cr_2O_7} + 2\,\mathrm{NaCl}\]
Orange crystals of potassium dichromate crystallise out. The whole route from ore to product is set out in FC 4.2.
Flowchart
FC 4.2 – Preparation of K₂Cr₂O₇ from chromite ore: chromite ore FeCr₂O₄ → fusion with Na₂CO₃ in free access of air → sodium chromate Na₂CrO₄ (yellow solution) + Fe₂O₃ + CO₂ → filter → acidify with H₂SO₄ → sodium dichromate Na₂Cr₂O₇·2H₂O (orange) → treat with KCl → potassium dichromate K₂Cr₂O₇ crystallises (orange crystals) + NaCl remains in solution
Chromate and Dichromate: the Same Chromium, Two Ions
The chromates and dichromates are interconvertible in aqueous solution depending upon the pH of the solution. Importantly, the oxidation state of chromium in chromate and in dichromate is the same (+6) — the change is one of condensation, not of oxidation.
\[2\,\mathrm{CrO_4^{2-}} + 2\,\mathrm{H^{+}} \rightarrow \mathrm{Cr_2O_7^{2-}} + \mathrm{H_2O}\]
\[\mathrm{Cr_2O_7^{2-}} + 2\,\mathrm{OH^{-}} \rightarrow 2\,\mathrm{CrO_4^{2-}} + \mathrm{H_2O}\]
In plain words: add acid to a yellow chromate solution and it turns orange; add alkali to an orange dichromate solution and it turns yellow.
The structures of the chromate ion, \(\mathrm{CrO_4^{2-}}\), and the dichromate ion, \(\mathrm{Cr_2O_7^{2-}}\), are shown in Fig. 4.9. The chromate ion is tetrahedral, whereas the dichromate ion consists of two tetrahedra sharing one corner, with a Cr–O–Cr bond angle of 126°.
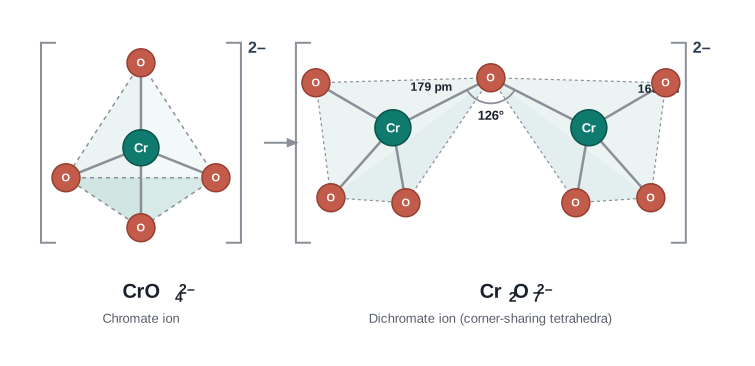
Fig. 4.9 – Structures of the chromate and dichromate ions.
Potassium Dichromate as an Oxidising Agent
Sodium and potassium dichromates are strong oxidising agents. The sodium salt has a greater solubility in water and is extensively used as an oxidising agent in organic chemistry. Potassium dichromate is used as a primary standard in volumetric analysis — it can be obtained pure and dry, its composition does not change on storage, so a solution of accurately known concentration can be made simply by weighing it out.
In acidic solution, its oxidising action can be represented as follows:
\[\mathrm{Cr_2O_7^{2-}} + 14\,\mathrm{H^{+}} + 6\,\mathrm{e^{-}} \rightarrow 2\,\mathrm{Cr^{3+}} + 7\,\mathrm{H_2O} \qquad (E^\circ = 1.33\ \text{V})\]
Note what this half-reaction tells you: each dichromate ion accepts six electrons, and it needs a plentiful supply of \(\mathrm{H^{+}}\) — which is why dichromate is used in acidic medium.
Thus, acidified potassium dichromate will oxidise iodides to iodine, sulphides to sulphur, tin(II) to tin(IV) and iron(II) salts to iron(III). The half-reactions of the reducing agents are noted below, each written so that it releases six electrons, to match the six taken up by dichromate:
\[6\,\mathrm{I^{-}} \rightarrow 3\,\mathrm{I_2} + 6\,\mathrm{e^{-}} \qquad ; \qquad 3\,\mathrm{Sn^{2+}} \rightarrow 3\,\mathrm{Sn^{4+}} + 6\,\mathrm{e^{-}}\]
\[3\,\mathrm{H_2S} \rightarrow 6\,\mathrm{H^{+}} + 3\,\mathrm{S} + 6\,\mathrm{e^{-}} \qquad ; \qquad 6\,\mathrm{Fe^{2+}} \rightarrow 6\,\mathrm{Fe^{3+}} + 6\,\mathrm{e^{-}}\]
The full ionic equation may be obtained by adding the half-reaction for potassium dichromate to the half-reaction for the reducing agent. For example, with iron(II):
\[\mathrm{Cr_2O_7^{2-}} + 14\,\mathrm{H^{+}} + 6\,\mathrm{Fe^{2+}} \rightarrow 2\,\mathrm{Cr^{3+}} + 6\,\mathrm{Fe^{3+}} + 7\,\mathrm{H_2O}\]
The electrons cancel because the two half-reactions were deliberately matched at six electrons each. This is the ion–electron method you met in redox chemistry, and it is the fastest way to write any dichromate or permanganate equation in an exam.
Potassium permanganate, KMnO₄
Potassium permanganate is prepared by fusion of \(\mathrm{MnO_2}\) with an alkali metal hydroxide and an oxidising agent like \(\mathrm{KNO_3}\). This produces the dark green \(\mathrm{K_2MnO_4}\), which disproportionates in a neutral or acidic solution to give permanganate.
\[2\,\mathrm{MnO_2} + 4\,\mathrm{KOH} + \mathrm{O_2} \rightarrow 2\,\mathrm{K_2MnO_4} + 2\,\mathrm{H_2O}\]
\[3\,\mathrm{MnO_4^{2-}} + 4\,\mathrm{H^{+}} \rightarrow 2\,\mathrm{MnO_4^{-}} + \mathrm{MnO_2} + 2\,\mathrm{H_2O}\]
In the second equation manganese(VI) is simultaneously oxidised to manganese(VII) and reduced to manganese(IV) — the disproportionation you met in Solved Example 4.12.
Commercially it is prepared by the alkaline oxidative fusion of \(\mathrm{MnO_2}\), followed by the electrolytic oxidation of manganate(VI).
\[\mathrm{MnO_2} \xrightarrow{\ \text{fused with KOH, oxidised with air or } \mathrm{KNO_3}\ } \underset{\text{manganate ion}}{\mathrm{MnO_4^{2-}}}\]
\[\underset{\text{manganate}}{\mathrm{MnO_4^{2-}}} \xrightarrow{\ \text{electrolytic oxidation in alkaline solution}\ } \underset{\text{permanganate ion}}{\mathrm{MnO_4^{-}}}\]
The two industrial stages are set out in FC 4.3.
Flowchart
FC 4.3 – Preparation of KMnO₄ from pyrolusite (MnO₂): MnO₂ → fusion with KOH + oxidising agent (air or KNO₃) → green manganate ion MnO₄²⁻ (Mn in +6) → branch (a) laboratory/acidic route: add H⁺ → disproportionation → purple MnO₄⁻ + MnO₂; branch (b) commercial route: electrolytic oxidation in alkaline solution → purple permanganate ion MnO₄⁻ (Mn in +7)
In the laboratory, a manganese(II) ion salt is oxidised by peroxodisulphate to permanganate:
\[2\,\mathrm{Mn^{2+}} + 5\,\mathrm{S_2O_8^{2-}} + 8\,\mathrm{H_2O} \rightarrow 2\,\mathrm{MnO_4^{-}} + 10\,\mathrm{SO_4^{2-}} + 16\,\mathrm{H^{+}}\]
Properties of Potassium Permanganate
Potassium permanganate forms dark purple (almost black) crystals which are isostructural with those of \(\mathrm{KClO_4}\). The salt is not very soluble in water (6.4 g per 100 g of water at 293 K), but when heated it decomposes at 513 K:
\[2\,\mathrm{KMnO_4} \rightarrow \mathrm{K_2MnO_4} + \mathrm{MnO_2} + \mathrm{O_2}\]
It has two physical properties of considerable interest: its intense colour, and its diamagnetism along with a temperature-dependent weak paramagnetism. These can be explained by the use of molecular orbital theory, which is beyond the present scope.
The manganate and permanganate ions are tetrahedral; the \(\pi\)-bonding takes place by overlap of the p orbitals of oxygen with the d orbitals of manganese, as drawn in Fig. 4.10. The green manganate is paramagnetic because of one unpaired electron, but the permanganate is diamagnetic due to the absence of an unpaired electron.
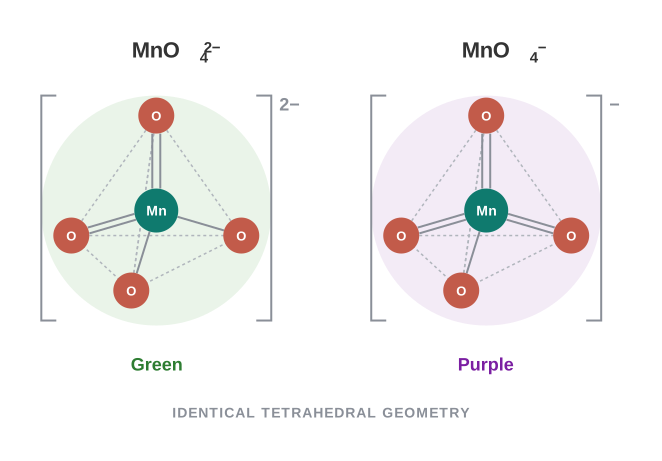
Fig. 4.10 – Structures of the manganate and permanganate ions.
Oxidising Reactions of Permanganate
Acidified permanganate solution oxidises oxalates to carbon dioxide, iron(II) to iron(III), nitrites to nitrates and iodides to free iodine. The half-reactions of the reductants are:
\[5\,\mathrm{C_2O_4^{2-}} \rightarrow 10\,\mathrm{CO_2} + 10\,\mathrm{e^{-}}\]
\[5\,\mathrm{Fe^{2+}} \rightarrow 5\,\mathrm{Fe^{3+}} + 5\,\mathrm{e^{-}}\]
\[5\,\mathrm{NO_2^{-}} + 5\,\mathrm{H_2O} \rightarrow 5\,\mathrm{NO_3^{-}} + 10\,\mathrm{H^{+}} + 10\,\mathrm{e^{-}}\]
\[10\,\mathrm{I^{-}} \rightarrow 5\,\mathrm{I_2} + 10\,\mathrm{e^{-}}\]
The full reaction can be written by adding the half-reaction for \(\mathrm{KMnO_4}\) to the half-reaction of the reducing agent, balancing wherever necessary.
If we represent the reduction of permanganate to manganate, to manganese dioxide, and to a manganese(II) salt by half-reactions:
\[\mathrm{MnO_4^{-}} + \mathrm{e^{-}} \rightarrow \mathrm{MnO_4^{2-}} \qquad (E^\circ = +0.56\ \text{V})\]
\[\mathrm{MnO_4^{-}} + 4\,\mathrm{H^{+}} + 3\,\mathrm{e^{-}} \rightarrow \mathrm{MnO_2} + 2\,\mathrm{H_2O} \qquad (E^\circ = +1.69\ \text{V})\]
\[\mathrm{MnO_4^{-}} + 8\,\mathrm{H^{+}} + 5\,\mathrm{e^{-}} \rightarrow \mathrm{Mn^{2+}} + 4\,\mathrm{H_2O} \qquad (E^\circ = +1.52\ \text{V})\]
We can very well see that the hydrogen ion concentration of the solution plays an important part in influencing the reaction — the product is manganate in strongly alkaline conditions, \(\mathrm{MnO_2}\) in neutral or faintly alkaline conditions, and \(\mathrm{Mn^{2+}}\) in acid.
Although many reactions can be understood by consideration of redox potential, the kinetics of the reaction is also an important factor. Permanganate at \([\mathrm{H^{+}}] = 1\) should oxidise water, but in practice the reaction is extremely slow unless either manganese(II) ions are present or the temperature is raised. Thermodynamics tells us what can happen; kinetics decides what actually does happen in reasonable time.
A few important oxidising reactions of \(\mathrm{KMnO_4}\) are given below.
1. In acid solutions:
- Iodine is liberated from potassium iodide:
\[10\,\mathrm{I^{-}} + 2\,\mathrm{MnO_4^{-}} + 16\,\mathrm{H^{+}} \rightarrow 2\,\mathrm{Mn^{2+}} + 8\,\mathrm{H_2O} + 5\,\mathrm{I_2}\]
- \(\mathrm{Fe^{2+}}\) ion (green) is converted to \(\mathrm{Fe^{3+}}\) (yellow):
\[5\,\mathrm{Fe^{2+}} + \mathrm{MnO_4^{-}} + 8\,\mathrm{H^{+}} \rightarrow \mathrm{Mn^{2+}} + 4\,\mathrm{H_2O} + 5\,\mathrm{Fe^{3+}}\]
- Oxalate ion or oxalic acid is oxidised at 333 K:
\[5\,\mathrm{C_2O_4^{2-}} + 2\,\mathrm{MnO_4^{-}} + 16\,\mathrm{H^{+}} \rightarrow 2\,\mathrm{Mn^{2+}} + 8\,\mathrm{H_2O} + 10\,\mathrm{CO_2}\]
- Hydrogen sulphide is oxidised, sulphur being precipitated:
\[\mathrm{H_2S} \rightarrow 2\,\mathrm{H^{+}} + \mathrm{S^{2-}}\]
\[5\,\mathrm{S^{2-}} + 2\,\mathrm{MnO_4^{-}} + 16\,\mathrm{H^{+}} \rightarrow 2\,\mathrm{Mn^{2+}} + 8\,\mathrm{H_2O} + 5\,\mathrm{S}\]
- Sulphurous acid or sulphite is oxidised to a sulphate or sulphuric acid:
\[5\,\mathrm{SO_3^{2-}} + 2\,\mathrm{MnO_4^{-}} + 6\,\mathrm{H^{+}} \rightarrow 2\,\mathrm{Mn^{2+}} + 3\,\mathrm{H_2O} + 5\,\mathrm{SO_4^{2-}}\]
- Nitrite is oxidised to nitrate:
\[5\,\mathrm{NO_2^{-}} + 2\,\mathrm{MnO_4^{-}} + 6\,\mathrm{H^{+}} \rightarrow 2\,\mathrm{Mn^{2+}} + 5\,\mathrm{NO_3^{-}} + 3\,\mathrm{H_2O}\]
2. In neutral or faintly alkaline solutions:
- A notable reaction is the oxidation of iodide to iodate:
\[2\,\mathrm{MnO_4^{-}} + \mathrm{H_2O} + \mathrm{I^{-}} \rightarrow 2\,\mathrm{MnO_2} + 2\,\mathrm{OH^{-}} + \mathrm{IO_3^{-}}\]
- Thiosulphate is oxidised almost quantitatively to sulphate:
\[8\,\mathrm{MnO_4^{-}} + 3\,\mathrm{S_2O_3^{2-}} + \mathrm{H_2O} \rightarrow 8\,\mathrm{MnO_2} + 6\,\mathrm{SO_4^{2-}} + 2\,\mathrm{OH^{-}}\]
- A manganous salt is oxidised to \(\mathrm{MnO_2}\); the presence of zinc sulphate or zinc oxide catalyses the oxidation:
\[2\,\mathrm{MnO_4^{-}} + 3\,\mathrm{Mn^{2+}} + 2\,\mathrm{H_2O} \rightarrow 5\,\mathrm{MnO_2} + 4\,\mathrm{H^{+}}\]
Notice that in acid the permanganate always ends up as colourless \(\mathrm{Mn^{2+}}\), while in neutral or faintly alkaline solution it stops at brown \(\mathrm{MnO_2}\) — the medium, not the reducing agent, decides the destination.
Note: Permanganate titrations in the presence of hydrochloric acid are unsatisfactory, since hydrochloric acid is itself oxidised to chlorine. Some of the permanganate would then be consumed by the acid instead of by the substance being estimated, and the titre value would be wrong. Dilute sulphuric acid is used instead.
4.5 The Lanthanoids
The Inner Transition Elements (f-Block)
The f-block consists of two series: the lanthanoids (the fourteen elements following lanthanum) and the actinoids (the fourteen elements following actinium).
Because lanthanum closely resembles the lanthanoids, it is usually included in any discussion of the lanthanoids, for which the general symbol Ln is often used. Similarly, a discussion of the actinoids includes actinium besides the fourteen elements constituting the series.
The lanthanoids resemble one another more closely than do the members of ordinary transition elements in any series. They have only one stable oxidation state, and their chemistry provides an excellent opportunity to examine the effect of small changes in size and nuclear charge along a series of otherwise similar elements — a kind of chemical control experiment.
The chemistry of the actinoids is, on the other hand, much more complicated. The complication arises partly from the occurrence of a wide range of oxidation states in these elements, and partly because their radioactivity creates special problems in their study. The two series will therefore be considered separately here.
The names, symbols, electronic configurations of atomic and some ionic states, and atomic and ionic radii of lanthanum and the lanthanoids (for which the general symbol Ln is used) are given in Table 4.9.
4.5.1 Electronic Configurations
It may be noted that atoms of these elements have an electronic configuration with \(6s^2\) common, but with variable occupancy of the 4f level (Table 4.9). The differences between one lanthanoid and the next therefore lie deep inside the atom, in an orbital that is well shielded from the outside world — which is exactly why their chemistry is so alike.
However, the electronic configurations of all the tripositive ions (the most stable oxidation state of all the lanthanoids) are of the form \(4f^{\,n}\) (\(n = 1\) to 14 with increasing atomic number). In forming \(\mathrm{Ln^{3+}}\), the two 6s electrons and one more electron (from 5d or 4f) are lost, leaving only 4f electrons outside the xenon core.
4.5.2 Atomic and Ionic Sizes
The overall decrease in atomic and ionic radii from lanthanum to lutetium — the lanthanoid contraction — is a unique feature in the chemistry of the lanthanoids. It has far-reaching consequences in the chemistry of the third transition series of the elements.
The decrease in atomic radii (derived from the structures of the metals) is not quite regular, whereas it is regular in the \(\mathrm{M^{3+}}\) ions, as Fig. 4.11 shows.
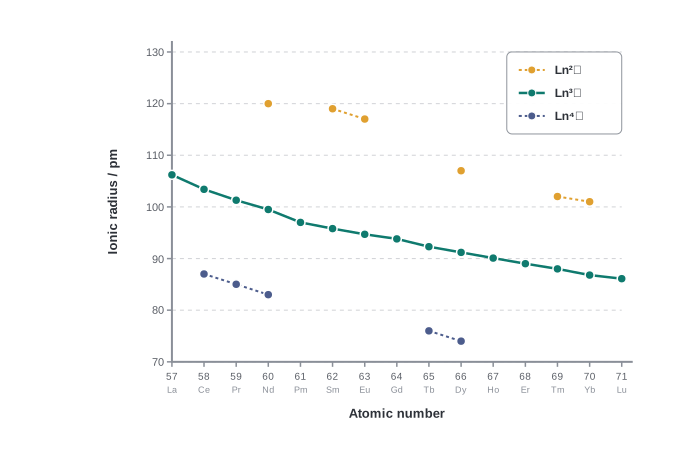
Fig. 4.11 – Trends in ionic radii of lanthanoids.
This contraction is, of course, similar to that observed in an ordinary transition series and is attributed to the same cause: the imperfect shielding of one electron by another in the same sub-shell. However, the shielding of one 4f electron by another is even less than that of one d electron by another. So, with the increase in nuclear charge along the series, there is a fairly regular decrease in the sizes with increasing atomic number.
The cumulative effect of the contraction of the lanthanoid series, known as lanthanoid contraction, causes the radii of the members of the third transition series to be very similar to those of the corresponding members of the second series. The almost identical radii of Zr (160 pm) and Hf (159 pm), a consequence of the lanthanoid contraction, account for their occurrence together in nature and for the difficulty faced in their separation.
Table 4.9: Electronic Configurations and Radii of Lanthanum and Lanthanoids
(Only electrons outside the [Xe] core are indicated.)
| Atomic Number | Name | Symbol | Ln | Ln²⁺ | Ln³⁺ | Ln⁴⁺ | Radius Ln /pm | Radius Ln³⁺ /pm |
|---|---|---|---|---|---|---|---|---|
| 57 | Lanthanum | La | \(5d^16s^2\) | \(5d^1\) | \(4f^{\,0}\) | 187 | 106 | |
| 58 | Cerium | Ce | \(4f^15d^16s^2\) | \(4f^{\,2}\) | \(4f^{\,1}\) | \(4f^{\,0}\) | 183 | 103 |
| 59 | Praseodymium | Pr | \(4f^{\,3}6s^2\) | \(4f^{\,3}\) | \(4f^{\,2}\) | \(4f^{\,1}\) | 182 | 101 |
| 60 | Neodymium | Nd | \(4f^{\,4}6s^2\) | \(4f^{\,4}\) | \(4f^{\,3}\) | \(4f^{\,2}\) | 181 | 99 |
| 61 | Promethium | Pm | \(4f^{\,5}6s^2\) | \(4f^{\,5}\) | \(4f^{\,4}\) | 181 | 98 | |
| 62 | Samarium | Sm | \(4f^{\,6}6s^2\) | \(4f^{\,6}\) | \(4f^{\,5}\) | 180 | 96 | |
| 63 | Europium | Eu | \(4f^{\,7}6s^2\) | \(4f^{\,7}\) | \(4f^{\,6}\) | 199 | 95 | |
| 64 | Gadolinium | Gd | \(4f^{\,7}5d^16s^2\) | \(4f^{\,7}5d^1\) | \(4f^{\,7}\) | 180 | 94 | |
| 65 | Terbium | Tb | \(4f^{\,9}6s^2\) | \(4f^{\,9}\) | \(4f^{\,8}\) | \(4f^{\,7}\) | 178 | 92 |
| 66 | Dysprosium | Dy | \(4f^{\,10}6s^2\) | \(4f^{\,10}\) | \(4f^{\,9}\) | \(4f^{\,8}\) | 177 | 91 |
| 67 | Holmium | Ho | \(4f^{\,11}6s^2\) | \(4f^{\,11}\) | \(4f^{\,10}\) | 176 | 89 | |
| 68 | Erbium | Er | \(4f^{\,12}6s^2\) | \(4f^{\,12}\) | \(4f^{\,11}\) | 175 | 88 | |
| 69 | Thulium | Tm | \(4f^{\,13}6s^2\) | \(4f^{\,13}\) | \(4f^{\,12}\) | 174 | 87 | |
| 70 | Ytterbium | Yb | \(4f^{\,14}6s^2\) | \(4f^{\,14}\) | \(4f^{\,13}\) | 173 | 86 | |
| 71 | Lutetium | Lu | \(4f^{\,14}5d^16s^2\) | \(4f^{\,14}5d^1\) | \(4f^{\,14}\) | – | – | – |
4.5.3 Oxidation States
In the lanthanoids, La(II) and Ln(III) compounds are the predominant species. However, occasionally +2 and +4 ions in solution or in solid compounds are also obtained.
This irregularity (as in the ionisation enthalpies) arises mainly from the extra stability of an empty, half-filled or filled f subshell — the same principle that made \(d^5\) and \(d^{10}\) so special in the transition series, now applied to \(f^{\,0}\), \(f^{\,7}\) and \(f^{\,14}\).
Thus, the formation of \(\mathrm{Ce^{IV}}\) is favoured by its noble gas configuration (\(f^{\,0}\)), but it is a strong oxidant, reverting to the common +3 state. The \(E^\circ\) value for \(\mathrm{Ce^{4+}}/\mathrm{Ce^{3+}}\) is +1.74 V, which suggests that it can oxidise water. However, the reaction rate is very slow, and hence Ce(IV) is a good analytical reagent — a reminder, once again, that a favourable electrode potential does not by itself guarantee a fast reaction.
Pr, Nd, Tb and Dy also exhibit the +4 state, but only in the oxides, \(\mathrm{MO_2}\).
\(\mathrm{Eu^{2+}}\) is formed by losing the two s electrons, and its \(f^{\,7}\) configuration accounts for the formation of this ion. However, \(\mathrm{Eu^{2+}}\) is a strong reducing agent, changing to the common +3 state. Similarly \(\mathrm{Yb^{2+}}\), which has the \(f^{\,14}\) configuration, is a reductant. \(\mathrm{Tb^{IV}}\) has half-filled f orbitals and is an oxidant. The behaviour of samarium is very much like europium, exhibiting both +2 and +3 oxidation states.
4.5.4 General Characteristics
All the lanthanoids are silvery white soft metals and tarnish rapidly in air. The hardness increases with increasing atomic number, samarium being steel hard. Their melting points range between 1000 and 1200 K, but samarium melts at 1623 K. They have a typical metallic structure and are good conductors of heat and electricity. Density and other properties change smoothly, except for Eu and Yb, and occasionally for Sm and Tm.
Colour and Magnetic Behaviour
Many trivalent lanthanoid ions are coloured, both in the solid state and in aqueous solutions. The colour of these ions may be attributed to the presence of f electrons. Neither \(\mathrm{La^{3+}}\) nor \(\mathrm{Lu^{3+}}\) ion shows any colour, but the rest do so — and that is no accident, since these two are precisely the \(f^{\,0}\) and \(f^{\,14}\) ions, with no possibility of an f–f electron jump.
However, the absorption bands are narrow, probably because of the excitation within the f level, which is well shielded from the surroundings and therefore barely disturbed by the ligands.
The lanthanoid ions other than the \(f^{\,0}\) type (\(\mathrm{La^{3+}}\) and \(\mathrm{Ce^{4+}}\)) and the \(f^{\,14}\) type (\(\mathrm{Yb^{2+}}\) and \(\mathrm{Lu^{3+}}\)) are all paramagnetic, since all the others have unpaired f electrons.
Ionisation Enthalpies
The first ionisation enthalpies of the lanthanoids are around \(600\ \text{kJ mol}^{-1}\), and the second about \(1200\ \text{kJ mol}^{-1}\), comparable with those of calcium. This is why the early members of the series behave chemically rather like calcium.
A detailed discussion of the variation of the third ionisation enthalpies indicates that exchange enthalpy considerations (as in the 3d orbitals of the first transition series) appear to impart a certain degree of stability to empty, half-filled and completely filled orbitals of the f level. This is indicated by the abnormally low value of the third ionisation enthalpy of lanthanum, gadolinium and lutetium — the three elements whose \(\mathrm{M^{3+}}\) ions come out as \(f^{\,0}\), \(f^{\,7}\) and \(f^{\,14}\) respectively.
Chemical Behaviour
In their chemical behaviour, in general, the earlier members of the series are quite reactive, similar to calcium; but with increasing atomic number, they behave more like aluminium.
Values of \(E^\circ\) for the half-reaction
\[\mathrm{Ln^{3+}}\text{(aq)} + 3\,\mathrm{e^{-}} \rightarrow \mathrm{Ln}\text{(s)}\]
are in the range of –2.2 to –2.4 V, except for Eu, for which the value is –2.0 V. This is, of course, a small variation — another sign of how alike the members of this family are. Such large negative values mean the metals are strongly electropositive and lose electrons readily.
The metals combine with hydrogen when gently heated in the gas. The carbides \(\mathrm{Ln_3C}\), \(\mathrm{Ln_2C_3}\) and \(\mathrm{LnC_2}\) are formed when the metals are heated with carbon. They liberate hydrogen from dilute acids and burn in halogens to form halides. They form oxides \(\mathrm{M_2O_3}\) and hydroxides \(\mathrm{M(OH)_3}\). The hydroxides are definite compounds, not just hydrated oxides. They are basic, like alkaline earth metal oxides and hydroxides.
Their general reactions are depicted in Fig. 4.12 and collected in FC 4.4.
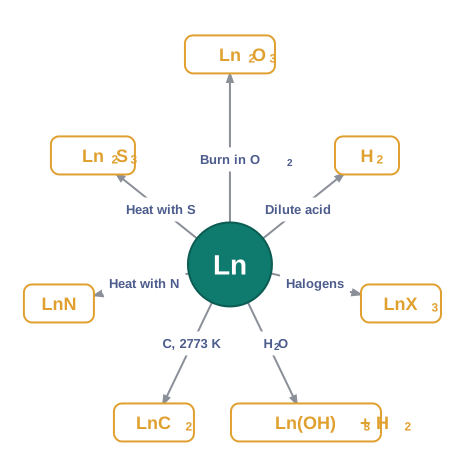
Fig. 4.12 – Chemical reactions of the lanthanoids.
Flowchart
FC 4.4 – Reactions of a lanthanoid metal Ln: Ln → burnt in O₂ → Ln₂O₃; Ln → with dilute acid → H₂ liberated; Ln → with halogens → LnX₃; Ln → with H₂O → Ln(OH)₃ + H₂; Ln → heated with C (2773 K) → LnC₂ (also Ln₃C, Ln₂C₃); Ln → heated with N → LnN; Ln → heated with S → Ln₂S₃; Ln → gently heated in H₂ gas → hydride
Uses of the Lanthanoids
The best single use of the lanthanoids is for the production of alloy steels for plates and pipes. A well known alloy is mischmetall, which consists of a lanthanoid metal (~ 95%) and iron (~ 5%) and traces of S, C, Ca and Al.
A good deal of mischmetall is used in Mg-based alloys to produce bullets, shells and lighter flint. Mixed oxides of lanthanoids are employed as catalysts in petroleum cracking. Some individual Ln oxides are used as phosphors in television screens and similar fluorescing surfaces.
4.6 The Actinoids
The actinoids include the fourteen elements from Th to Lr. The names, symbols and some properties of these elements are given in Table 4.10.
The actinoids are radioactive elements, and the earlier members have relatively long half-lives. The later ones have half-life values ranging from a day down to 3 minutes for lawrencium (\(Z = 103\)). The later members could be prepared only in nanogram quantities. These facts render their study more difficult: an element that vanishes while you are weighing it, and which must be handled behind shielding, does not lend itself to ordinary laboratory chemistry.
Table 4.10: Some Properties of Actinium and Actinoids
(Only electrons outside the [Rn] core are indicated.)
| Atomic Number | Name | Symbol | M | M³⁺ | M⁴⁺ | Radius M³⁺ /pm | Radius M⁴⁺ /pm |
|---|---|---|---|---|---|---|---|
| 89 | Actinium | Ac | \(6d^17s^2\) | \(5f^{\,0}\) | 111 | ||
| 90 | Thorium | Th | \(6d^27s^2\) | \(5f^{\,1}\) | \(5f^{\,0}\) | 99 | |
| 91 | Protactinium | Pa | \(5f^{\,2}6d^17s^2\) | \(5f^{\,2}\) | \(5f^{\,1}\) | 96 | |
| 92 | Uranium | U | \(5f^{\,3}6d^17s^2\) | \(5f^{\,3}\) | \(5f^{\,2}\) | 103 | 93 |
| 93 | Neptunium | Np | \(5f^{\,4}6d^17s^2\) | \(5f^{\,4}\) | \(5f^{\,3}\) | 101 | 92 |
| 94 | Plutonium | Pu | \(5f^{\,6}7s^2\) | \(5f^{\,5}\) | \(5f^{\,4}\) | 100 | 90 |
| 95 | Americium | Am | \(5f^{\,7}7s^2\) | \(5f^{\,6}\) | \(5f^{\,5}\) | 99 | 89 |
| 96 | Curium | Cm | \(5f^{\,7}6d^17s^2\) | \(5f^{\,7}\) | \(5f^{\,6}\) | 99 | 88 |
| 97 | Berkelium | Bk | \(5f^{\,9}7s^2\) | \(5f^{\,8}\) | \(5f^{\,7}\) | 98 | 87 |
| 98 | Californium | Cf | \(5f^{\,10}7s^2\) | \(5f^{\,9}\) | \(5f^{\,8}\) | 98 | 86 |
| 99 | Einsteinium | Es | \(5f^{\,11}7s^2\) | \(5f^{\,10}\) | \(5f^{\,9}\) | – | – |
| 100 | Fermium | Fm | \(5f^{\,12}7s^2\) | \(5f^{\,11}\) | \(5f^{\,10}\) | – | – |
| 101 | Mendelevium | Md | \(5f^{\,13}7s^2\) | \(5f^{\,12}\) | \(5f^{\,11}\) | – | – |
| 102 | Nobelium | No | \(5f^{\,14}7s^2\) | \(5f^{\,13}\) | \(5f^{\,12}\) | – | – |
| 103 | Lawrencium | Lr | \(5f^{\,14}6d^17s^2\) | \(5f^{\,14}\) | \(5f^{\,13}\) | – | – |
4.6.1 Electronic Configurations
All the actinoids are believed to have the electronic configuration of \(7s^2\) and variable occupancy of the 5f and 6d subshells. The fourteen electrons are formally added to 5f, though not in thorium (\(Z = 90\)); but from Pa onwards, the 5f orbitals fill until they are complete at element 103.
The irregularities in the electronic configurations of the actinoids, like those in the lanthanoids, are related to the stabilities of the \(f^{\,0}\), \(f^{\,7}\) and \(f^{\,14}\) occupancies of the 5f orbitals. Thus, the configurations of Am and Cm are \([\mathrm{Rn}]\,5f^{\,7}7s^2\) and \([\mathrm{Rn}]\,5f^{\,7}6d^17s^2\) — in curium, an extra electron prefers to sit alone in a 6d orbital rather than disturb the half-filled 5f set.
Although the 5f orbitals resemble the 4f orbitals in the angular part of their wave-function, they are not as buried as the 4f orbitals. Hence 5f electrons can participate in bonding to a far greater extent. This single structural difference is the key to almost everything that separates the actinoids from the lanthanoids: electrons that can take part in bonding can also be lost in many different numbers, which is why the actinoids show so wide a range of oxidation states.
4.6.2 Ionic Sizes
The general trend seen in the lanthanoids is observable in the actinoids as well. There is a gradual decrease in the size of the atoms or \(\mathrm{M^{3+}}\) ions across the series. This may be referred to as the actinoid contraction, like the lanthanoid contraction.
The contraction is, however, greater from element to element in this series, resulting from the poor shielding by 5f electrons.
4.6.3 Oxidation States
There is a greater range of oxidation states in the actinoids, which is in part attributed to the fact that the 5f, 6d and 7s levels are of comparable energies. When three sets of orbitals lie so close together, electrons can be drawn from any of them, and many different numbers of electrons can be removed for a similar energy cost. The known oxidation states of the actinoids are listed in Table 4.11.
Table 4.11: Oxidation States of Actinium and Actinoids
| Ac | Th | Pa | U | Np | Pu | Am | Cm | Bk | Cf | Es | Fm | Md | No | Lr |
|---|---|---|---|---|---|---|---|---|---|---|---|---|---|---|
| 3 | 3 | 3 | 3 | 3 | 3 | 3 | 3 | 3 | 3 | 3 | 3 | 3 | 3 | |
| 4 | 4 | 4 | 4 | 4 | 4 | 4 | 4 | |||||||
| 5 | 5 | 5 | 5 | 5 | ||||||||||
| 6 | 6 | 6 | 6 | |||||||||||
| 7 | 7 |
The actinoids show in general the +3 oxidation state. The elements in the first half of the series frequently exhibit higher oxidation states. For example, the maximum oxidation state increases from +4 in Th to +5, +6 and +7 respectively in Pa, U and Np, but decreases in the succeeding elements (Table 4.11).
The actinoids resemble the lanthanoids in having more compounds in the +3 state than in the +4 state. However, +3 and +4 ions tend to hydrolyse — that is, they are so small and highly charged that they pull apart the water molecules around them, giving acidic solutions and often precipitating hydroxides.
Because the distribution of oxidation states among the actinoids is so uneven, and so different for the former and the later elements, it is unsatisfactory to review their chemistry in terms of oxidation states.
4.6.4 General Characteristics and Comparison with Lanthanoids
The actinoid metals are all silvery in appearance, but display a variety of structures. The structural variability is obtained due to irregularities in metallic radii, which are far greater than in the lanthanoids.
Chemical Reactivity
The actinoids are highly reactive metals, especially when finely divided. The action of boiling water on them, for example, gives a mixture of oxide and hydride, and combination with most non-metals takes place at moderate temperatures.
Hydrochloric acid attacks all these metals, but most are only slightly affected by nitric acid, owing to the formation of protective oxide layers — the metal makes its own coat and hides behind it. Alkalies have no action on them.
Magnetic Properties
The magnetic properties of the actinoids are more complex than those of the lanthanoids. Although the variation in the magnetic susceptibility of the actinoids with the number of unpaired 5f electrons is roughly parallel to the corresponding results for the lanthanoids, the latter have higher values.
Ionisation Enthalpies
It is evident from the behaviour of the actinoids that the ionisation enthalpies of the early actinoids, though not accurately known, are lower than for the early lanthanoids.
This is quite reasonable. It is to be expected that when 5f orbitals are beginning to be occupied, they will penetrate less into the inner core of electrons. The 5f electrons will therefore be more effectively shielded from the nuclear charge than the 4f electrons of the corresponding lanthanoids. Because the outer electrons are less firmly held, they are more easily removed — and they are available for bonding in the actinoids.
How the Two Series Compare
A comparison of the actinoids with the lanthanoids, with respect to the different characteristics discussed above, reveals that behaviour similar to that of the lanthanoids is not evident until the second half of the actinoid series. However, even the early actinoids resemble the lanthanoids in showing close similarities with each other, and in gradual variation in properties which do not entail a change in oxidation state.
The lanthanoid and actinoid contractions have extended effects on the sizes, and therefore on the properties, of the elements succeeding them in their respective periods. The lanthanoid contraction is more important, because the chemistry of the elements succeeding the actinoids is much less known at the present time.
4.7 Some Applications of d- and f-Block Elements
Metals of Construction and of the Battery
Iron and steels are the most important construction materials we have. Their production is based on three operations: the reduction of iron oxides, the removal of impurities, and the addition of carbon and of alloying metals such as Cr, Mn and Ni. Carbon hardens the iron; chromium resists corrosion; manganese and nickel toughen the steel — each addition is made for a reason.
Some compounds are manufactured for special purposes, such as TiO for the pigment industry and \(\mathrm{MnO_2}\) for use in dry battery cells. The battery industry also requires Zn and Ni/Cd. Zinc forms the case and the anode of the ordinary dry cell, while nickel–cadmium cells were long the standard rechargeable battery.
The Coinage Metals
The elements of Group 11 — copper, silver and gold — are still worthy of being called the coinage metals, although Ag and Au are now restricted to collection items. Contemporary UK ‘copper’ coins are copper-coated steel, and the ‘silver’ UK coins are a Cu/Ni alloy. The reason these three metals were chosen for coins in the first place is the one we met in Section 4.3.8: they resist oxidation, so a coin survives decades of handling without corroding away.
Catalysts of the Chemical Industry
Many of the metals and/or their compounds are essential catalysts in the chemical industry. Recall from Section 4.3.12 why they are so good at the job: a transition metal can slip between oxidation states, and its surface atoms have partly filled d orbitals with which to grip and weaken the bonds of the molecules that land on them, as sketched in Fig. 4.13.
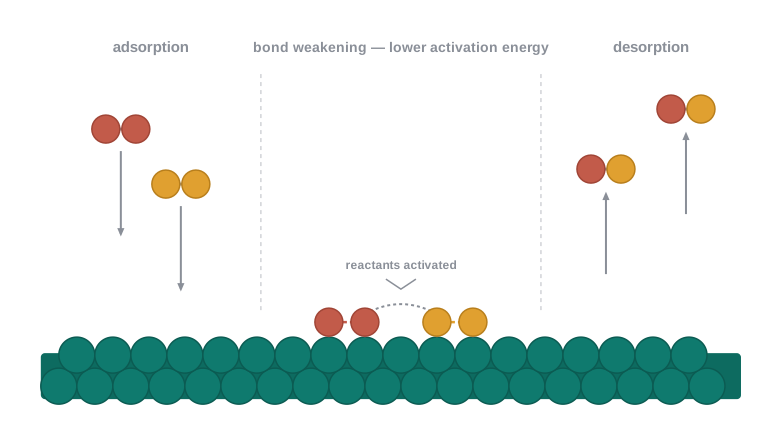
Fig. 4.13 – Heterogeneous catalysis on a transition metal surface.
\(\mathrm{V_2O_5}\) catalyses the oxidation of \(\mathrm{SO_2}\) in the manufacture of sulphuric acid. \(\mathrm{TiCl_4}\) with \(\mathrm{Al(CH_3)_3}\) forms the basis of the Ziegler catalysts used to manufacture polyethylene (polythene).
Iron catalysts are used in the Haber process for the production of ammonia from \(\mathrm{N_2}/\mathrm{H_2}\) mixtures. Nickel catalysts enable the hydrogenation of fats to proceed — this is the reaction that converts liquid vegetable oils into the semi-solid fats used in cooking.
In the Wacker process, the oxidation of ethyne to ethanal is catalysed by \(\mathrm{PdCl_2}\). Nickel complexes are useful in the polymerisation of alkynes and other organic compounds such as benzene.
Finally, the photographic industry relies on the special light-sensitive properties of AgBr, which darkens where light falls on it — the basis of the photographic film.
4.8 Summary
The Two Blocks
- The d-block, consisting of Groups 3–12, occupies the large middle section of the periodic table. In these elements the inner d orbitals are progressively filled.
- The f-block is placed outside, at the bottom of the periodic table. In the elements of this block, the 4f and 5f orbitals are progressively filled.
- Corresponding to the filling of the 3d, 4d and 5d orbitals, three series of transition elements are well recognised.
General Characteristics of the Transition Elements
- All the transition elements exhibit typical metallic properties: high tensile strength, ductility, malleability, thermal and electrical conductivity, and metallic character.
- Their melting and boiling points are high. This is attributed to the involvement of the \((n-1)d\) electrons, in addition to the \(ns\) electrons, resulting in strong interatomic bonding.
- In many of these properties, the maxima occur at about the middle of each series. This indicates that one unpaired electron per d orbital is a particularly favourable configuration for strong interatomic interaction.
- Successive ionisation enthalpies do not increase as steeply as in the main group elements with increasing atomic number. Hence the loss of a variable number of electrons from the \((n-1)d\) orbitals is not energetically unfavourable.
- The involvement of the \((n-1)d\) electrons gives these elements their distinct character. Thus, in addition to variable oxidation states, they exhibit paramagnetic behaviour, catalytic properties, and a tendency to form coloured ions, interstitial compounds and complexes.
- The magnetic moment of a first-row transition metal ion is obtained from the spin-only formula, where \(n\) is the number of unpaired electrons:
\[\mu = \sqrt{n(n+2)}\ \text{BM}\]
Chemical Behaviour
- The transition elements vary widely in their chemical behaviour. Many of them are sufficiently electropositive to dissolve in mineral acids, although a few are ‘noble’. Of the first series, with the exception of copper, all the metals are relatively reactive.
- The transition metals react with a number of non-metals — oxygen, nitrogen, sulphur and the halogens — to form binary compounds.
- The first series transition metal oxides are generally formed from the reaction of the metals with oxygen at high temperatures. These oxides dissolve in acids and bases to form oxometallic salts.
Two Important Compounds
- Potassium dichromate (\(\mathrm{K_2Cr_2O_7}\)) is prepared from the chromite ore by fusion with alkali in the presence of air, and acidifying the extract.
- Potassium permanganate (\(\mathrm{KMnO_4}\)) is prepared from pyrolusite ore (\(\mathrm{MnO_2}\)).
- Both the dichromate and the permanganate ions are strong oxidising agents.
Flowchart
FC 4.5 – The two preparations side by side. (a) K₂Cr₂O₇: chromite ore FeCr₂O₄ → fusion with Na₂CO₃ in air → Na₂CrO₄ (yellow) → acidify with H₂SO₄ → Na₂Cr₂O₇ (orange) → add KCl → K₂Cr₂O₇. (b) KMnO₄: pyrolusite MnO₂ → alkaline oxidative fusion with KOH + air/KNO₃ → K₂MnO₄ (green, Mn +6) → electrolytic oxidation in alkaline solution (or disproportionation in acid) → KMnO₄ (purple, Mn +7).
The Inner Transition Elements
- The two series of inner transition elements, the lanthanoids and the actinoids, constitute the f-block of the periodic table.
- With the successive filling of the inner 4f orbitals, there is a gradual decrease in the atomic and ionic sizes of these metals along the series — the lanthanoid contraction. This has far-reaching consequences in the chemistry of the elements succeeding them, notably the close similarity in size between the second and third transition series.
- Lanthanum and all the lanthanoids are rather soft white metals. They react easily with water to give solutions containing +3 ions.
- The principal oxidation state of the lanthanoids is +3, although +4 and +2 oxidation states are also exhibited by some of them occasionally.
- The chemistry of the actinoids is more complex, in view of their ability to exist in different oxidation states. Furthermore, many of the actinoid elements are radioactive, which makes the study of these elements rather difficult.
Applications
- There are many useful applications of the d- and f-block elements and their compounds. Notable among them are their use in the varieties of steels, as catalysts, in complexes and in organic syntheses.
4.9 NCERT Questions
- Write down the electronic configuration of:
- \(\mathrm{Cr^{3+}}\) (ii) \(\mathrm{Pm^{3+}}\) (iii) \(\mathrm{Cu^{+}}\) (iv) \(\mathrm{Ce^{4+}}\) (v) \(\mathrm{Co^{2+}}\) (vi) \(\mathrm{Lu^{2+}}\) (vii) \(\mathrm{Mn^{2+}}\) (viii) \(\mathrm{Th^{4+}}\)
Why are \(\mathrm{Mn^{2+}}\) compounds more stable than \(\mathrm{Fe^{2+}}\) towards oxidation to their +3 state?
Explain briefly how the +2 state becomes more and more stable in the first half of the first row transition elements with increasing atomic number.
To what extent do the electronic configurations decide the stability of oxidation states in the first series of the transition elements? Illustrate your answer with examples.
What may be the stable oxidation state of the transition element with the following d electron configurations in the ground state of their atoms: \(3d^3\), \(3d^5\), \(3d^8\) and \(3d^4\)?
Name the oxometal anions of the first series of the transition metals in which the metal exhibits the oxidation state equal to its group number.
What is lanthanoid contraction? What are the consequences of lanthanoid contraction?
What are the characteristics of the transition elements and why are they called transition elements? Which of the d-block elements may not be regarded as the transition elements?
In what way is the electronic configuration of the transition elements different from that of the non-transition elements?
What are the different oxidation states exhibited by the lanthanoids?
Explain giving reasons:
- Transition metals and many of their compounds show paramagnetic behaviour.
- The enthalpies of atomisation of the transition metals are high.
- The transition metals generally form coloured compounds.
- Transition metals and many of their compounds act as good catalysts.
What are interstitial compounds? Why are such compounds well known for transition metals?
How is the variability in oxidation states of transition metals different from that of the non-transition metals? Illustrate with examples.
Describe the preparation of potassium dichromate from iron chromite ore. What is the effect of increasing pH on a solution of potassium dichromate?
Describe the oxidising action of potassium dichromate and write the ionic equations for its reaction with:
- iodide (ii) iron(II) solution and (iii) \(\mathrm{H_2S}\)
Describe the preparation of potassium permanganate. How does the acidified permanganate solution react with (i) iron(II) ions (ii) \(\mathrm{SO_2}\) and (iii) oxalic acid? Write the ionic equations for the reactions.
For \(\mathrm{M^{2+}}/\mathrm{M}\) and \(\mathrm{M^{3+}}/\mathrm{M^{2+}}\) systems, the \(E^\circ\) values for some metals are as follows:
| Couple | \(E^\circ\) | Couple | \(E^\circ\) |
|---|---|---|---|
| \(\mathrm{Cr^{2+}}/\mathrm{Cr}\) | –0.9 V | \(\mathrm{Cr^{3+}}/\mathrm{Cr^{2+}}\) | –0.4 V |
| \(\mathrm{Mn^{2+}}/\mathrm{Mn}\) | –1.2 V | \(\mathrm{Mn^{3+}}/\mathrm{Mn^{2+}}\) | +1.5 V |
| \(\mathrm{Fe^{2+}}/\mathrm{Fe}\) | –0.4 V | \(\mathrm{Fe^{3+}}/\mathrm{Fe^{2+}}\) | +0.8 V |
Use this data to comment upon: a. the stability of \(\mathrm{Fe^{3+}}\) in acid solution as compared to that of \(\mathrm{Cr^{3+}}\) or \(\mathrm{Mn^{3+}}\), and b. the ease with which iron can be oxidised as compared to a similar process for either chromium or manganese metal.
Predict which of the following will be coloured in aqueous solution: \(\mathrm{Ti^{3+}}\), \(\mathrm{V^{3+}}\), \(\mathrm{Cu^{+}}\), \(\mathrm{Sc^{3+}}\), \(\mathrm{Mn^{2+}}\), \(\mathrm{Fe^{3+}}\) and \(\mathrm{Co^{2+}}\). Give reasons for each.
Compare the stability of the +2 oxidation state for the elements of the first transition series.
Compare the chemistry of the actinoids with that of the lanthanoids with special reference to:
- electronic configuration (ii) atomic and ionic sizes (iii) oxidation state (iv) chemical reactivity.
How would you account for the following:
- Of the \(d^4\) species, \(\mathrm{Cr^{2+}}\) is strongly reducing while manganese(III) is strongly oxidising.
- Cobalt(II) is stable in aqueous solution, but in the presence of complexing reagents it is easily oxidised.
- The \(d^1\) configuration is very unstable in ions.
What is meant by ‘disproportionation’? Give two examples of disproportionation reactions in aqueous solution.
Which metal in the first series of transition metals exhibits the +1 oxidation state most frequently, and why?
Calculate the number of unpaired electrons in the following gaseous ions: \(\mathrm{Mn^{3+}}\), \(\mathrm{Cr^{3+}}\), \(\mathrm{V^{3+}}\) and \(\mathrm{Ti^{3+}}\). Which one of these is the most stable in aqueous solution?
Give examples and suggest reasons for the following features of transition metal chemistry:
- The lowest oxide of a transition metal is basic, the highest is amphoteric/acidic.
- A transition metal exhibits its highest oxidation state in oxides and fluorides.
- The highest oxidation state is exhibited in the oxoanions of a metal.
Indicate the steps in the preparation of:
- \(\mathrm{K_2Cr_2O_7}\) from chromite ore. (ii) \(\mathrm{KMnO_4}\) from pyrolusite ore.
What are alloys? Name an important alloy which contains some of the lanthanoid metals. Mention its uses.
What are inner transition elements? Decide which of the following atomic numbers are the atomic numbers of the inner transition elements: 29, 59, 74, 95, 102, 104.
The chemistry of the actinoid elements is not so smooth as that of the lanthanoids. Justify this statement by giving some examples from the oxidation states of these elements.
Which is the last element in the series of the actinoids? Write the electronic configuration of this element. Comment on the possible oxidation state of this element.
Use Hund’s rule to derive the electronic configuration of the \(\mathrm{Ce^{3+}}\) ion, and calculate its magnetic moment on the basis of the ‘spin-only’ formula.
Name the members of the lanthanoid series which exhibit the +4 oxidation state and those which exhibit the +2 oxidation state. Try to correlate this type of behaviour with the electronic configurations of these elements.
Compare the chemistry of the actinoids with that of the lanthanoids with reference to:
- electronic configuration (ii) oxidation states and (iii) chemical reactivity.
Write the electronic configurations of the elements with the atomic numbers 61, 91, 101 and 109.
Compare the general characteristics of the first series of the transition metals with those of the second and third series metals in the respective vertical columns. Give special emphasis on the following points:
- electronic configurations (ii) oxidation states (iii) ionisation enthalpies and (iv) atomic sizes.
Write down the number of 3d electrons in each of the following ions: \(\mathrm{Ti^{2+}}\), \(\mathrm{V^{2+}}\), \(\mathrm{Cr^{3+}}\), \(\mathrm{Mn^{2+}}\), \(\mathrm{Fe^{2+}}\), \(\mathrm{Fe^{3+}}\), \(\mathrm{Co^{2+}}\), \(\mathrm{Ni^{2+}}\) and \(\mathrm{Cu^{2+}}\). Indicate how you would expect the five 3d orbitals to be occupied for these hydrated ions (octahedral).
Comment on the statement that the elements of the first transition series possess many properties different from those of the heavier transition elements.
What can be inferred from the magnetic moment values of the following complex species?
| Example | Magnetic Moment (BM) |
|---|---|
| \(\mathrm{K_4[Mn(CN)_6]}\) | 2.2 |
| \(\mathrm{[Fe(H_2O)_6]^{2+}}\) | 5.3 |
| \(\mathrm{K_2[MnCl_4]}\) | 5.9 |
4.10 Check Your Concepts
Zinc, cadmium and mercury are placed in the d-block, yet they are not counted as transition metals. State the IUPAC definition of a transition metal and use it to explain this exclusion. Why, then, is zinc still discussed alongside the 3d series?
Assertion: Chromium has the ground-state configuration \(3d^54s^1\) and not \(3d^44s^2\). Reason: The 3d and 4s orbitals are very close in energy, and a half-filled d set has extra stability. Decide whether the assertion and the reason are each correct, and whether the reason correctly explains the assertion.
Arrange the following in increasing order of the property named, and give a reason for the order in each case:
- \(\mathrm{Sc}\), \(\mathrm{Ti}\), \(\mathrm{Cr}\), \(\mathrm{Zn}\) — enthalpy of atomisation
- \(\mathrm{VO_2^{+}}\), \(\mathrm{Cr_2O_7^{2-}}\), \(\mathrm{MnO_4^{-}}\) — oxidising power
- \(\mathrm{Ti}\), \(\mathrm{Cu}\), \(\mathrm{Fe}\) — density
“Two tetrahedra sharing one corner.” Identify the ion described, name the ion it turns into when alkali is added, and explain why the oxidation state of the metal does not change even though the colour of the solution does.
Why does the melting point of the 3d series rise to a maximum near the middle of the row and then fall? Account for the anomalously low value of manganese in this trend.
Explain why permanganate titrations are carried out with dilute sulphuric acid and not with hydrochloric acid. Write the equation for the unwanted reaction that would take place with HCl.
\(\mathrm{Cu^{+}}\) has a \(3d^{10}\) configuration, which should be stable, yet copper(I) salts disproportionate in water. Write the equation for the disproportionation and explain, in terms of hydration enthalpy, why \(\mathrm{Cu^{2+}}\)(aq) wins.
Complete and balance the following ionic equations, and in each case state the oxidation state of the metal in the product:
- \(\mathrm{Cr_2O_7^{2-}} + \mathrm{H^{+}} + \mathrm{Sn^{2+}} \rightarrow\)
- \(\mathrm{MnO_4^{-}} + \mathrm{H^{+}} + \mathrm{NO_2^{-}} \rightarrow\)
- \(\mathrm{MnO_4^{-}} + \mathrm{H_2O} + \mathrm{I^{-}} \rightarrow\)
A student is given three aqueous solutions labelled X, Y and Z. X is colourless, Y is pink and Z is blue. All three contain first-row transition metal ions in the +2 state. Using Table 4.8, suggest a possible identity for each ion and justify your choice on the basis of the d-electron count.
Assertion: The second and third transition series have almost the same atomic radii. Reason: The lanthanoid contraction cancels the expected increase in size on going from the second to the third series. Decide whether the assertion and the reason are each correct, and whether the reason correctly explains the assertion. Give one chemical consequence of this fact.
Why is the third ionisation enthalpy of manganese much higher than that of iron, even though iron has the larger nuclear charge? Use your answer to explain why \(\mathrm{Mn^{3+}}\) is a strong oxidising agent while \(\mathrm{Fe^{3+}}\) is not.
Match the following:
| Column I | Column II |
|---|---|
| a. \(\mathrm{VH_{0.56}}\) | i. Primary standard in volumetric analysis |
| b. Mischmetall | ii. Interstitial compound |
| c. \(\mathrm{K_2Cr_2O_7}\) | iii. Ziegler catalyst component |
| d. \(\mathrm{TiCl_4}\) | iv. Lighter flint |
Explain each of the following observations:
- The highest fluoride of manganese is \(\mathrm{MnF_4}\), but its highest oxide is \(\mathrm{Mn_2O_7}\).
- \(\mathrm{Cr(VI)}\) in acidic medium is a strong oxidising agent, but \(\mathrm{MoO_3}\) and \(\mathrm{WO_3}\) are not.
- The oxidation state of nickel in \(\mathrm{Ni(CO)_4}\) is zero.
Interstitial compounds such as TiC are harder and higher-melting than the pure metal, yet they still conduct electricity. Explain both facts in terms of what happens inside the metal lattice.
Ce(IV) has an \(E^\circ\) value of +1.74 V for the \(\mathrm{Ce^{4+}}/\mathrm{Ce^{3+}}\) couple, which suggests it should oxidise water — yet it is used as an analytical reagent in aqueous solution. Resolve this apparent contradiction.
Why are the ionisation enthalpies of the early actinoids lower than those of the early lanthanoids, while the actinoid contraction is nevertheless greater than the lanthanoid contraction? Distinguish carefully between the two kinds of shielding involved.
State, with reasons, whether each of the following ions is coloured or colourless in aqueous solution: \(\mathrm{Zn^{2+}}\), \(\mathrm{Cu^{2+}}\), \(\mathrm{Ti^{4+}}\), \(\mathrm{V^{3+}}\), \(\mathrm{La^{3+}}\), \(\mathrm{Nd^{3+}}\).
Iron(III) catalyses the reaction between iodide and persulphate ions. Write the two steps of the catalytic cycle, show that they add up to the overall reaction, and explain why the uncatalysed reaction is slow.
Titanium and vanadium are strongly electropositive, yet both are passive to dilute non-oxidising acids at room temperature. Explain this apparent contradiction between the \(E^\circ\) values and the observed behaviour.
Predict the product formed when acidified \(\mathrm{KMnO_4}\) solution reacts with each of the following, and state the colour change you would observe: (i) \(\mathrm{FeSO_4}\) solution, (ii) oxalic acid at 333 K, (iii) \(\mathrm{H_2S}\).
4.11 Practice With Numericals
Calculate the spin-only magnetic moment of each of the following ions and identify which one would be attracted most strongly by a magnetic field: \(\mathrm{Ti^{3+}}\), \(\mathrm{Cr^{3+}}\), \(\mathrm{Fe^{3+}}\), \(\mathrm{Ni^{2+}}\).
The spin-only magnetic moment of a divalent ion of a 3d metal is 4.90 BM. Determine the number of unpaired electrons, write the electronic configuration of the ion, and identify it.
Determine the oxidation number of the transition metal in each of the following: \(\mathrm{K_2Cr_2O_7}\), \(\mathrm{KMnO_4}\), \(\mathrm{K_2MnO_4}\), \(\mathrm{FeCr_2O_4}\), \(\mathrm{VO_2^{+}}\) and \(\mathrm{[Fe(CN)_6]^{4-}}\).
Calculate the mass of potassium dichromate required to oxidise 7.6 g of iron(II) sulphate in acidic solution. (Molar masses: \(\mathrm{K_2Cr_2O_7} = 294\ \text{g mol}^{-1}\); \(\mathrm{FeSO_4} = 152\ \text{g mol}^{-1}\).)
In an acidic medium, \(25.0\ \mathrm{cm^3}\) of an iron(II) solution required \(20.0\ \mathrm{cm^3}\) of \(0.02\ \mathrm{M}\ \mathrm{KMnO_4}\) for complete oxidation. Write the balanced ionic equation and calculate the molarity of the iron(II) solution.
Balance the following equation by the ion-electron (half-reaction) method in acidic medium, and state how many electrons are transferred per mole of dichromate: \[\mathrm{Cr_2O_7^{2-}} + \mathrm{I^{-}} + \mathrm{H^{+}} \rightarrow \mathrm{Cr^{3+}} + \mathrm{I_2} + \mathrm{H_2O}\]
Calculate the percentage of chromium by mass in potassium dichromate, \(\mathrm{K_2Cr_2O_7}\). (Atomic masses: K = 39, Cr = 52, O = 16.)
The solubility of potassium permanganate is 6.4 g per 100 g of water at 293 K. Calculate the mass of water required to dissolve 16.0 g of \(\mathrm{KMnO_4}\) completely at this temperature.
Write the electronic configuration of \(\mathrm{Eu^{3+}}\) and of \(\mathrm{Yb^{2+}}\), and calculate the spin-only magnetic moment of each. State which of the two is diamagnetic.
Using the data of Table 4.4, calculate the total energy required to convert one mole of gaseous chromium atoms into gaseous \(\mathrm{Cr^{2+}}\) ions, and compare it with the corresponding value for manganese. Comment on how this difference is reflected in their \(E^\circ\)(M²⁺/M) values.