Chapter 6: Haloalkanes and Haloarenes
In the late 1920s, chemists were asked to solve a dangerous problem. The refrigerators of the time ran on ammonia, methyl chloride and sulphur dioxide — gases that were toxic, and in some cases flammable. A leaking fridge could kill a family in its sleep. The American chemist Thomas Midgley Jr. and his team set out to design a gas that would do the same cooling job but refuse to burn, refuse to react and refuse to poison anyone. What they arrived at was a small carbon compound wearing a coat of chlorine and fluorine atoms — a chlorofluorocarbon, later sold under the name Freon. It was so unreactive that Midgley famously demonstrated its safety by inhaling a lungful and blowing out a candle flame with it.
For forty years the compound looked like a triumph. Then, in the 1970s, Mario Molina and F. Sherwood Rowland asked an uncomfortable question: if this molecule reacts with nothing, what finally becomes of it? Their answer — that it drifts intact into the stratosphere, where ultraviolet light snaps off chlorine atoms that go on to destroy ozone — earned them a share of the Nobel Prize in Chemistry in 1995, and changed international law.
The same feature that made the molecule safe made it permanent. To understand how a halogen atom can do this, we must first understand the bond it forms with carbon.
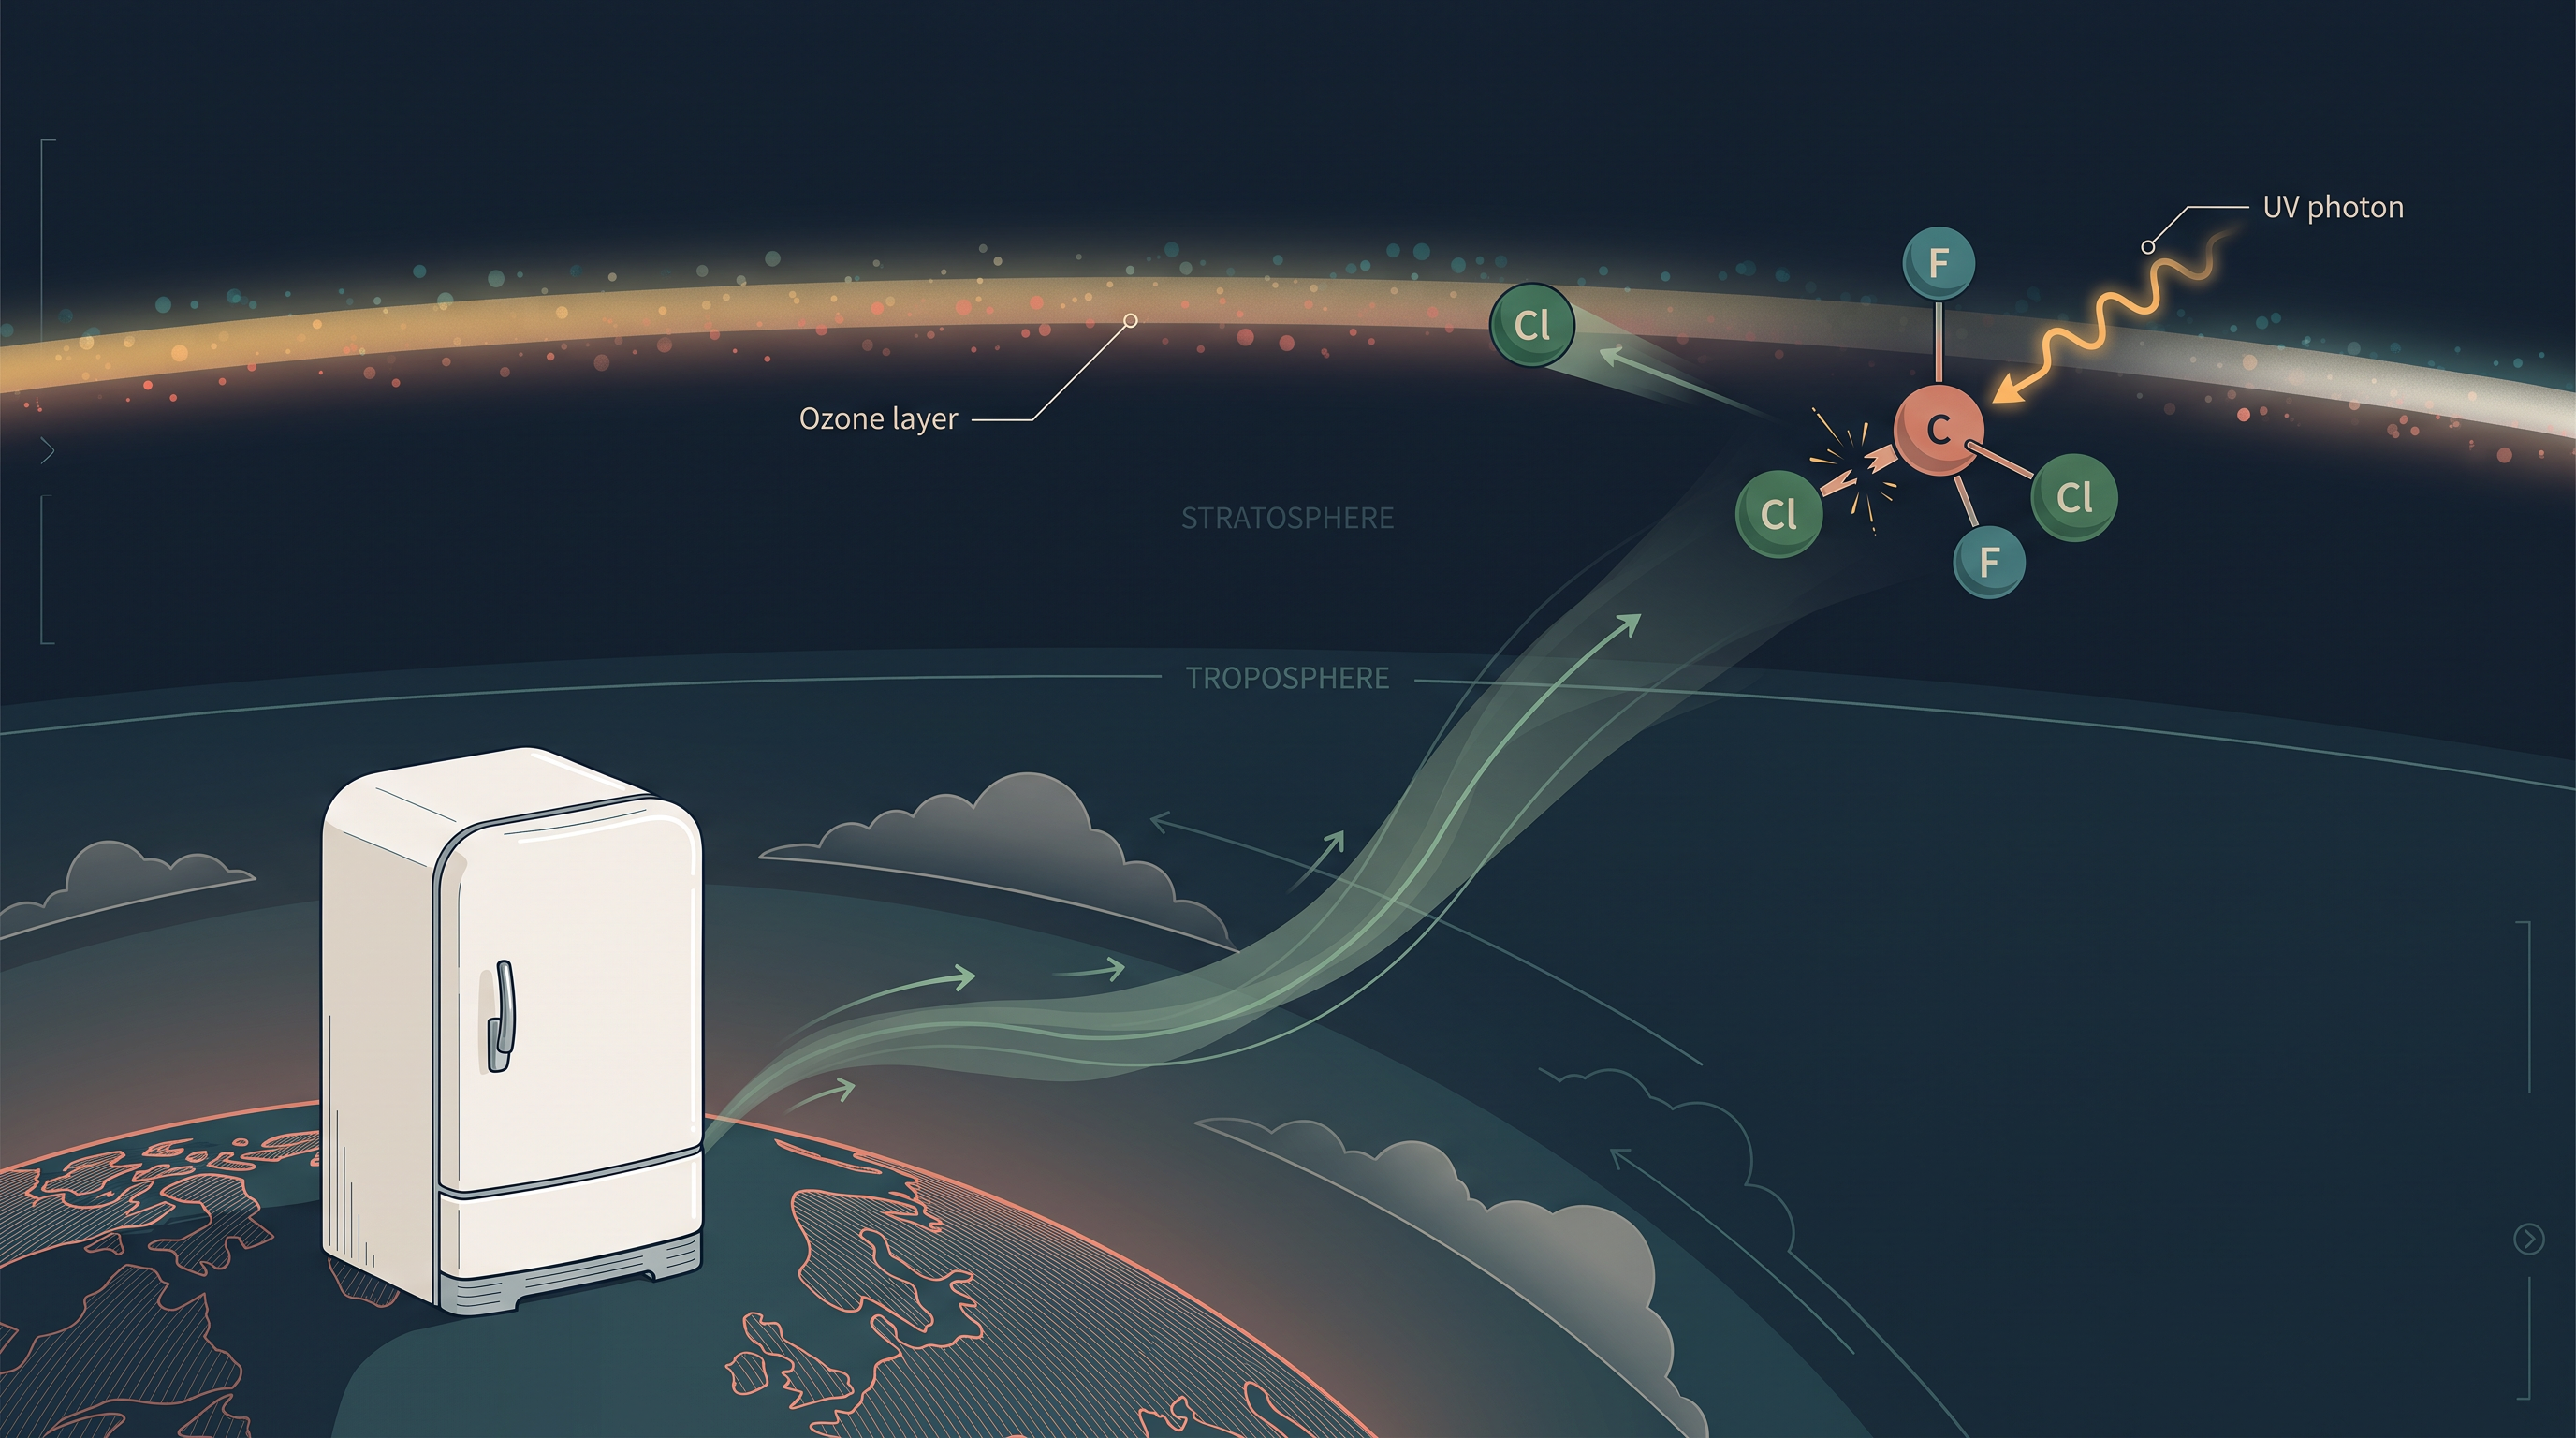
Fig. 6.0 – The escaping Freon molecule and ozone destruction.
Keep these questions in mind. Each of them will be answered as you work through this chapter.
6.1 Classification
What Organohalogen Compounds Are
Take any hydrocarbon — aliphatic (an open-chain compound such as ethane) or aromatic (a ring compound such as benzene) — and replace one or more of its hydrogen atoms with a halogen atom. What you get is an organohalogen compound. If the parent was an aliphatic hydrocarbon, the product is an alkyl halide, also called a haloalkane. If the parent was an aromatic hydrocarbon, the product is an aryl halide, also called a haloarene.
The difference between the two families lies in the carbon that carries the halogen. In a haloalkane, the halogen is attached to an \(sp^3\) hybridised carbon atom of an alkyl group. In a haloarene, it is attached to an \(sp^2\) hybridised carbon atom of an aryl group. Recall from Class XI that an \(sp^3\) carbon has formed four single bonds and is tetrahedral, while an \(sp^2\) carbon is part of a double bond or an aromatic ring and is trigonal planar. This single structural difference is responsible for almost every difference in behaviour you will meet later in this chapter, so it is worth fixing firmly in mind now.
Many halogen-containing organic compounds occur in nature, and several of them are clinically useful. Chloramphenicol, a chlorine-containing antibiotic produced by microorganisms, is very effective in the treatment of typhoid fever. Our own body manufactures thyroxine, an iodine-containing hormone whose deficiency causes the disease goitre. Among synthetic halogen compounds, chloroquine is used to treat malaria, and halothane is used as an anaesthetic during surgery. Certain fully fluorinated compounds are even being considered as potential blood substitutes in surgery.
Beyond medicine, these compounds are of great industrial and everyday importance. They dissolve relatively non-polar substances well, so they are widely used as solvents, and they serve as starting materials for the synthesis of a very wide range of organic compounds. In this Chapter you will study the important methods of preparation, the physical and chemical properties, and the uses of organohalogen compounds.
Haloalkanes and haloarenes may be classified in the following ways.
6.1.1 On the Basis of Number of Halogen Atoms
The simplest classification asks only one question: how many halogen atoms does the molecule carry? On this basis, compounds are called mono-, di- or polyhalogen (tri-, tetra-, and so on) compounds, depending on whether they contain one, two, or more halogen atoms in their structures.
Among the haloalkanes, \(\\mathrm{C\_2H\_5X}\) is a monohaloalkane; a molecule carrying \(-\\mathrm{CH\_2X}\) groups on two adjacent carbons is a dihaloalkane; and one carrying three halogen atoms is a trihaloalkane. The same idea applies to the aromatic family, giving monohaloarenes, dihaloarenes and trihaloarenes, as shown in Fig. 6.1.
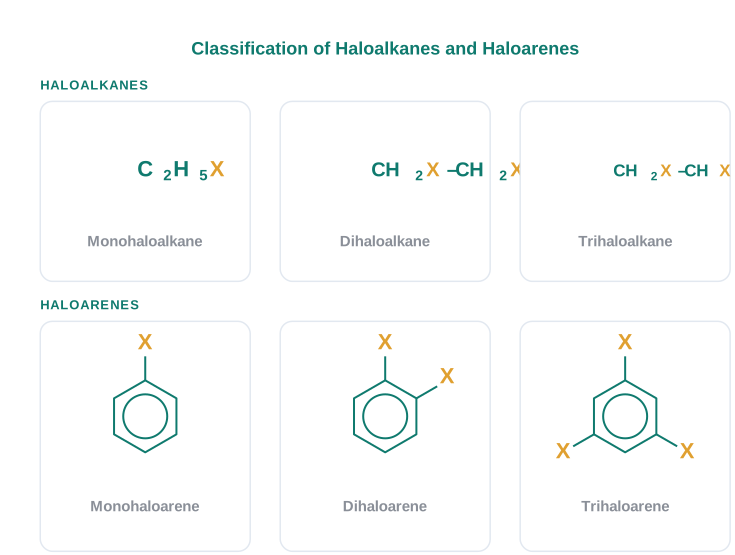
Fig. 6.1 – Classification of haloalkanes and haloarenes by halogen count.
Monohalocompounds can be classified further, and the basis of that finer classification is the hybridisation of the carbon atom to which the halogen is bonded. This is taken up in the next two subsections.
6.1.2 Compounds Containing \(sp^3\) C—X Bond (X = F, Cl, Br, I)
In this class, the halogen always sits on a tetrahedral, \(sp^3\) hybridised carbon. What varies is what that carbon is attached to. This class includes the following three types.
(a) Alkyl Halides or Haloalkanes (R—X)
In alkyl halides, the halogen atom is bonded to an alkyl group (R). They form a homologous series represented by the general formula \(\\mathrm{C\_nH\_{2n+1}X}\) — that is, each member differs from the next by a \(-\\mathrm{CH\_2}-\) unit.
Alkyl halides are further classified as primary, secondary or tertiary according to the nature of the carbon atom to which the halogen is attached. If the halogen is attached to a primary carbon atom — a carbon joined to only one other carbon — the compound is called a primary alkyl halide, or 1° alkyl halide. If the halogen sits on a secondary carbon (joined to two other carbons), it is a secondary alkyl halide (2°); if it sits on a tertiary carbon (joined to three other carbons), it is a tertiary alkyl halide (3°). The three arrangements are compared in Fig. 6.2.
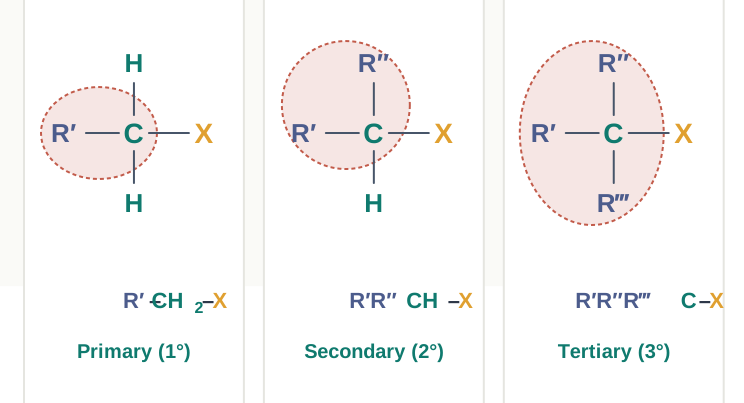
Fig. 6.2 – Primary, secondary and tertiary alkyl halides.
(b) Allylic Halides
These are the compounds in which the halogen atom is bonded to an \(sp^3\) hybridised carbon atom that is adjacent to a carbon–carbon double bond (C=C) — that is, to an allylic carbon. Note carefully that the halogen is not on the double bond itself; it is one carbon away from it, as marked in Fig. 6.3.
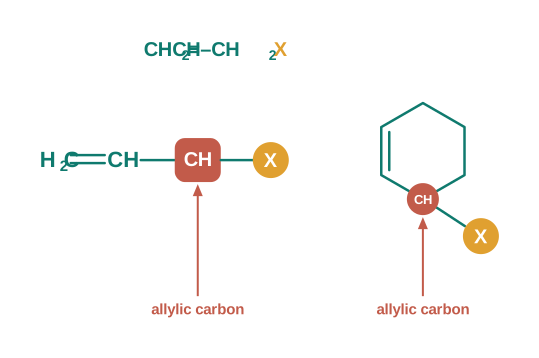
Fig. 6.3 – Allylic halides.
(c) Benzylic Halides
These are the compounds in which the halogen atom is bonded to an \(sp^3\) hybridised carbon atom that is attached to an aromatic ring. Again, the halogen is not on the ring — it is on the carbon just outside it. Like ordinary alkyl halides, benzylic halides may be primary, secondary or tertiary, depending on what else that benzylic carbon carries: \(\\mathrm{C\_6H\_5CH\_2X}\) is 1°; if the carbon also bears one methyl group and a hydrogen it is 2°; and if it bears two methyl groups it is 3°. These are drawn in Fig. 6.4.
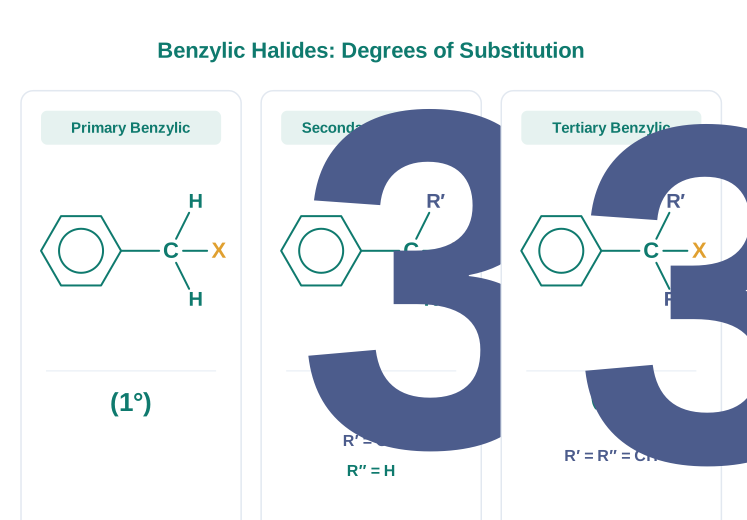
Fig. 6.4 – Primary, secondary and tertiary benzylic halides.
6.1.3 Compounds Containing \(sp^2\) C—X Bond
Here the halogen is attached to a trigonal planar, \(sp^2\) hybridised carbon — a carbon that is part of a double bond or of an aromatic ring. This class includes the following two types.
(a) Vinylic Halides
These are the compounds in which the halogen atom is bonded to an \(sp^2\) hybridised carbon atom of a carbon–carbon double bond (C=C). The halogen is now on the double bond, not next to it, as Fig. 6.5 shows.
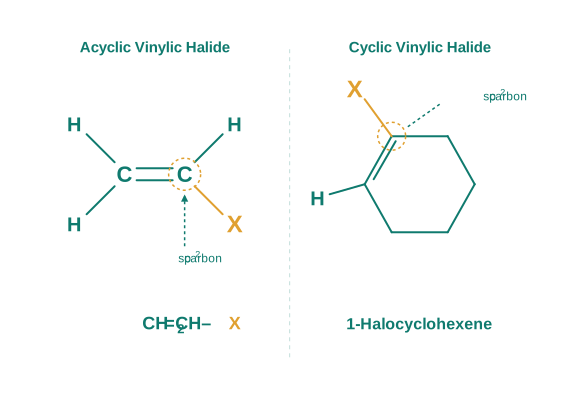
Fig. 6.5 – Vinylic halides.
(b) Aryl Halides
These are the compounds in which the halogen atom is directly bonded to an \(sp^2\) hybridised carbon atom of an aromatic ring, as in Fig. 6.6.
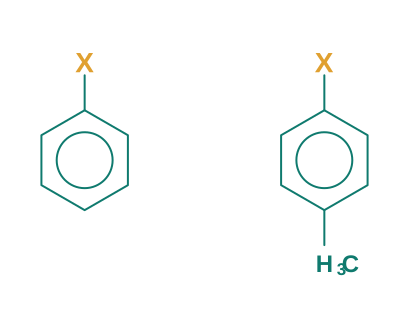
Fig. 6.6 – Aryl halides.
The five structural types you have now met — alkyl, allylic, benzylic, vinylic and aryl halides — look deceptively similar on paper, and telling them apart quickly is a standard first step in examination questions on this chapter. The essential test is always the same: identify the carbon that carries the halogen, and ask what its hybridisation is and what surrounds it.
6.2 Nomenclature
Having learnt how halogenated compounds are classified, let us now learn how they are named. Two naming systems run side by side in this chapter: the older common system, which you will still see on reagent bottles and in industry, and the IUPAC system, which is the one examinations expect.
Naming Alkyl Halides
The common name of an alkyl halide is built by naming the alkyl group first and following it with the name of the halide. Thus \(\\mathrm{CH\_3CH\_2CH\_2Br}\) is n-propyl bromide — the alkyl group is n-propyl and the halide is bromide.
In the IUPAC system, alkyl halides are named as halosubstituted hydrocarbons. The halogen is treated as a substituent (fluoro-, chloro-, bromo-, iodo-) on the longest carbon chain, and its position is given by a number chosen so that the substituents get the lowest possible locants. By this system the same compound, \(\\mathrm{CH\_3CH\_2CH\_2Br}\), becomes 1-bromopropane.
The two systems are placed side by side below for a set of typical compounds, as shown in Fig. 6.7.
Fig. 6.7 – Common and IUPAC names of some alkyl halides.
Naming Haloarenes
For mono halogen substituted derivatives of benzene, the common name and the IUPAC name are the same — \(\\mathrm{C\_6H\_5Br}\) is bromobenzene in both systems, because there is only one possible position for the substituent and so no number is needed.
For dihalogen derivatives, the two systems part company. The common system uses the prefixes o- (ortho), m- (meta) and p- (para) to indicate whether the two halogens are on adjacent carbons, one carbon apart, or directly opposite each other. The IUPAC system, as you have learnt in Class XI, uses the numerals 1,2; 1,3 and 1,4 instead. Thus m-dibromobenzene is named 1,3-dibromobenzene, and sym-tribromobenzene is named 1,3,5-tribromobenzene. These are drawn in Fig. 6.8.
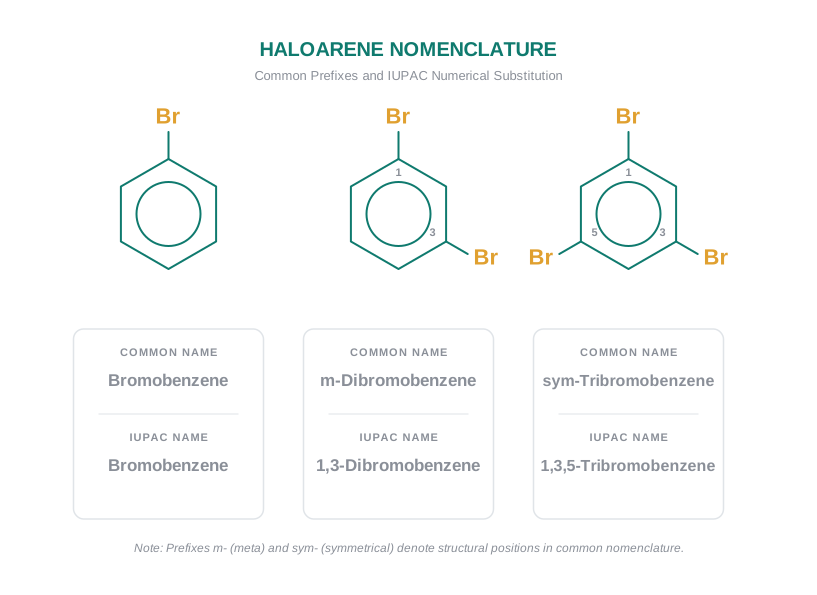
Fig. 6.8 – Nomenclature of mono-, di- and trihaloarenes.
Dihaloalkanes: gem- and vic-
The dihaloalkanes having the same type of halogen atoms are named as alkylidene or alkylene dihalides in the common system. These dihalo-compounds are further classified according to where the two halogen atoms sit. They are called geminal halides, or gem-dihalides, when both the halogen atoms are present on the same carbon atom of the chain, and vicinal halides, or vic-dihalides, when the halogen atoms are present on adjacent carbon atoms.
In the common name system, gem-dihalides are named as alkylidene halides and vic-dihalides are named as alkylene dihalides. In the IUPAC system, both are simply named as dihaloalkanes, with numbers doing the work of the prefixes. Thus \(\\mathrm{H\_3C{-}CHCl\_2}\) is ethylidene chloride (a gem-dihalide) in the common system and 1,1-dichloroethane in IUPAC; while \(\\mathrm{ClCH\_2{-}CH\_2Cl}\) is ethylene dichloride (a vic-dihalide) and 1,2-dichloroethane in IUPAC.
Some common examples of halocompounds are mentioned in Table 6.1.
Table 6.1: Common and IUPAC Names of some Halides
| Structure | Common name | IUPAC name |
|---|---|---|
| \(\\mathrm{CH\_3CH\_2CH(Cl)CH\_3}\) | sec-Butyl chloride | 2-Chlorobutane |
| \(\\mathrm{(CH\_3)\_3CCH\_2Br}\) | neo-Pentyl bromide | 1-Bromo-2,2-dimethylpropane |
| \(\\mathrm{(CH\_3)\_3CBr}\) | tert-Butyl bromide | 2-Bromo-2-methylpropane |
| \(\\mathrm{CH\_2{=}CHCl}\) | Vinyl chloride | Chloroethene |
| \(\\mathrm{CH\_2{=}CHCH\_2Br}\) | Allyl bromide | 3-Bromopropene |
| Benzene ring with Cl and \(\\mathrm{CH\_3}\) on adjacent carbons | o-Chlorotoluene | 1-Chloro-2-methylbenzene or 2-Chlorotoluene |
| Benzene ring with \(-\\mathrm{CH\_2Cl}\) | Benzyl chloride | Chlorophenylmethane |
| \(\\mathrm{CH\_2Cl\_2}\) | Methylene chloride | Dichloromethane |
| \(\\mathrm{CHCl\_3}\) | Chloroform | Trichloromethane |
| \(\\mathrm{CHBr\_3}\) | Bromoform | Tribromomethane |
| \(\\mathrm{CCl\_4}\) | Carbon tetrachloride | Tetrachloromethane |
| \(\\mathrm{CH\_3CH\_2CH\_2F}\) | n-Propyl fluoride | 1-Fluoropropane |
Write IUPAC names of the compounds (i) to (vi) shown in Fig. 6.9.
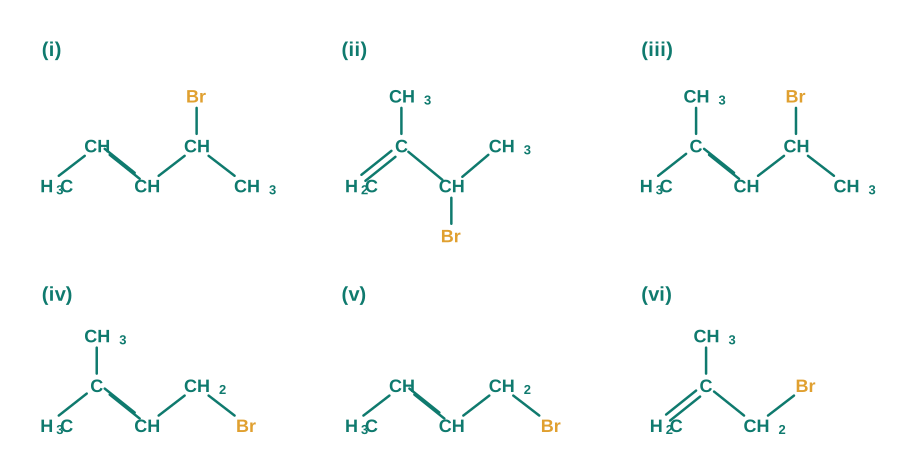
[Caption:] Fig. 6.9 – Skeletal structures of isomeric bromoalkenes. [/Caption:]
In each case, choose the longest chain that contains the double bond, number so that the double bond gets the lowest locant, and cite bromo as a substituent.
4-Bromopent-2-ene
3-Bromo-2-methylbut-1-ene
4-Bromo-3-methylpent-2-ene
1-Bromo-2-methylbut-2-ene
1-Bromobut-2-ene
3-Bromo-2-methylpropene
6.3 Nature of C–X Bond
Halogen atoms are more electronegative than carbon. Electronegativity, as you learnt in Class XI, is the tendency of an atom to pull the shared pair of electrons in a covalent bond towards itself. Since the halogen pulls harder than carbon does, the shared electron pair of the carbon–halogen bond does not sit midway between the two atoms; it lies closer to the halogen.
The result is that the carbon–halogen bond of an alkyl halide is polarised: the carbon atom bears a partial positive charge (\(\\delta+\)) whereas the halogen atom bears a partial negative charge (\(\\delta-\)). This is shown in Fig. 6.10.
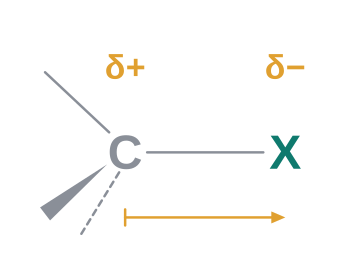
Fig. 6.10 – Polarisation of the carbon–halogen bond.
This small, permanent charge separation is the single most important structural fact in the whole chapter. An electron-deficient carbon is an open invitation to any electron-rich species, and that is precisely why haloalkanes are attacked by nucleophiles, as you will see in Section 6.7.
Size, Bond Length and Bond Strength
As we go down the group in the periodic table, the size of the halogen atom increases. Fluorine is the smallest halogen atom and iodine is the largest, because each successive element has an extra shell of electrons. Consequently the carbon–halogen bond length also increases steadily from C—F to C—I: the bigger the halogen, the further its bonding electrons are from the nucleus, and the further apart the two nuclei must sit.
A longer bond is also a weaker bond, because the shared electron pair is held less firmly between the two nuclei. This is why the C—X bond enthalpy — the energy needed to break one mole of that bond — falls sharply from C—F to C—I. Some typical bond lengths, bond enthalpies and dipole moments are given in Table 6.2.
Table 6.2: Carbon-Halogen (C—X) Bond Lengths, Bond Enthalpies and Dipole Moments
| Bond | Bond length/pm | C-X Bond enthalpies/ kJ mol\(^{-1}\) | Dipole moment/Debye |
|---|---|---|---|
| \(\\mathrm{CH\_3{-}F}\) | 139 | 452 | 1.847 |
| \(\\mathrm{CH\_3{-}Cl}\) | 178 | 351 | 1.860 |
| \(\\mathrm{CH\_3{-}Br}\) | 193 | 293 | 1.830 |
| \(\\mathrm{CH\_3{-}I}\) | 214 | 234 | 1.636 |
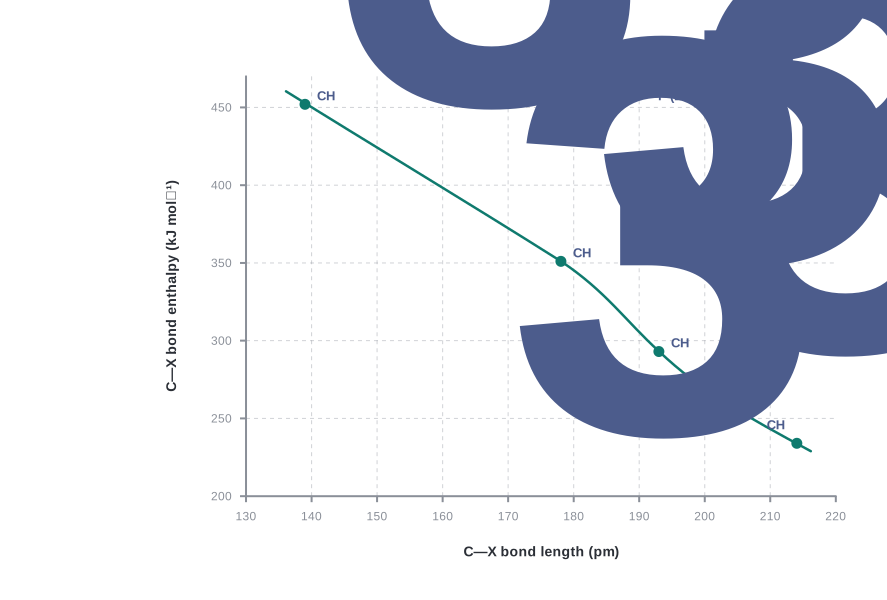
Fig. 6.11 – C—X bond length versus bond enthalpy.
6.4 Methods of Preparation of Haloalkanes
There are three practical ways of putting a halogen atom on to an \(sp^3\) carbon: start from an alcohol and swap its \(-\\mathrm{OH}\) group for a halogen; start from a hydrocarbon (an alkane or an alkene); or start from a haloalkane you already have and exchange one halogen for another. All three routes are summarised in FC 6.1 and taken up one by one below.
Flowchart
FC 6.1 – Routes to haloalkanes: Alcohol (R–OH) → with conc. HX / PX₃ / PCl₅ / SOCl₂ → R–X | Alkane → free radical halogenation with X₂ and UV light or heat → mixture of mono- and polyhaloalkanes | Alkene → addition of HX (Markovnikov) → alkyl halide; addition of X₂ in CCl₄ → vic-dihalide | R–Cl or R–Br → NaI in dry acetone (Finkelstein) → R–I; → metallic fluoride AgF/Hg₂F₂/CoF₂/SbF₃ (Swarts) → R–F
6.4.1 From Alcohols
The hydroxyl group of an alcohol is replaced by a halogen on reaction with concentrated halogen acids, phosphorus halides, or thionyl chloride.
Of these three reagents, thionyl chloride is preferred. The reason is one of purity rather than chemistry: in this reaction the alkyl halide is formed along with the gases \(\\mathrm{SO\_2}\) and \(\\mathrm{HCl}\). The two gaseous products simply escape from the reaction mixture, so nothing is left behind to contaminate the product, and the reaction gives pure alkyl halides.
\[\\mathrm{R{-}OH} + \\mathrm{SOCl\_2} \\rightarrow \\mathrm{R{-}Cl} + \\mathrm{SO\_2} + \\mathrm{HCl}\]
Reaction with Halogen Acids
The reactions of primary and secondary alcohols with \(\\mathrm{HCl}\) require the presence of a catalyst, \(\\mathrm{ZnCl\_2}\).
\[\\mathrm{R{-}OH} + \\mathrm{HCl} \\xrightarrow{\\ \\mathrm{ZnCl\_2}\\ } \\mathrm{R{-}Cl} + \\mathrm{H\_2O}\]
With tertiary alcohols, no catalyst is needed at all: the reaction is conducted by simply shaking the alcohol with concentrated \(\\mathrm{HCl}\) at room temperature.
For preparing an alkyl bromide, constant boiling \(\\mathrm{HBr}\) (48%) is used. The hydrogen bromide may also be generated in the flask itself from sodium bromide and sulphuric acid:
\[\\mathrm{R{-}OH} + \\mathrm{NaBr} + \\mathrm{H\_2SO\_4} \\rightarrow \\mathrm{R{-}Br} + \\mathrm{NaHSO\_4} + \\mathrm{H\_2O}\]
Good yields of R—I may be obtained by heating alcohols with sodium or potassium iodide in 95% orthophosphoric acid.
The order of reactivity of alcohols with a given haloacid is 3° > 2° > 1°. Notice that this order is the exact opposite of the one you will meet for \(\\mathrm{S\_N}2\) reactions later — a tertiary alcohol reacts fastest here because the reaction proceeds through a carbocation, and a tertiary carbocation is the most stable of the three.
Reaction with Phosphorus Halides
Phosphorus halides convert alcohols to alkyl halides cleanly:
\[3,\\mathrm{R{-}OH} + \\mathrm{PX\_3} \\rightarrow 3,\\mathrm{R{-}X} + \\mathrm{H\_3PO\_3} \\qquad (\\mathrm{X} = \\mathrm{Cl},\\ \\mathrm{Br})\]
\[\\mathrm{R{-}OH} + \\mathrm{PCl\_5} \\rightarrow \\mathrm{R{-}Cl} + \\mathrm{POCl\_3} + \\mathrm{HCl}\]
Phosphorus tribromide and triiodide are usually generated in situ — that is, produced in the reaction mixture itself — by the reaction of red phosphorus with bromine and iodine respectively. The alcohol is therefore simply heated with red phosphorus and the halogen:
\[\\mathrm{R{-}OH} \\xrightarrow{\\ \\text{red } \\mathrm{P}/\\mathrm{X\_2},\\ \\mathrm{X\_2} = \\mathrm{Br\_2},\\ \\mathrm{I\_2}\\ } \\mathrm{R{-}X}\]
Preparing Alkyl Chlorides in the Laboratory
The preparation of an alkyl chloride is carried out either by passing dry hydrogen chloride gas through a solution of the alcohol, or by heating a mixture of the alcohol and the concentrated aqueous halogen acid.
One limitation must be noted at once. The above methods are not applicable for the preparation of aryl halides. The reason lies in the structure of phenols: the carbon–oxygen bond in phenols has a partial double bond character because of the delocalisation of the oxygen lone pair into the ring. Being stronger than an ordinary single bond, this C—O bond is difficult to break, so phenol does not give chlorobenzene with \(\\mathrm{HCl}\). Aryl halides must therefore be made by an entirely different route, which is the subject of Section 6.5.
6.4.2 From Hydrocarbons
(I) From Alkanes by Free Radical Halogenation
Free radical chlorination or bromination of alkanes gives a complex mixture of isomeric mono- and polyhaloalkanes, which is difficult to separate as pure compounds. Consequently, the yield of any single compound is low.
\[\\mathrm{CH\_3CH\_2CH\_2CH\_3} \\xrightarrow\[\\text{or heat}]{\\ \\mathrm{Cl\_2}/\\text{UV light}\\ } \\mathrm{CH\_3CH\_2CH\_2CH\_2Cl} + \\mathrm{CH\_3CH\_2CHClCH\_3}\]
The reason for the mess is easy to see. Ultraviolet light splits \(\\mathrm{Cl\_2}\) into chlorine radicals, and a chlorine radical is not fussy — it will pull a hydrogen atom from whichever C—H bond it happens to collide with. Butane has two different kinds of hydrogen atoms, so two different monochlorides form, and once a monochloride has formed it too has C—H bonds available for a second attack, giving di- and polychlorinated products as well.
(II) From Alkenes
(i) Addition of hydrogen halides. An alkene is converted to the corresponding alkyl halide by reaction with hydrogen chloride, hydrogen bromide or hydrogen iodide. The \(\\pi\) electrons of the double bond attack the hydrogen of H—X, and the halide ion then adds to the carbon, as shown in Fig. 6.12.
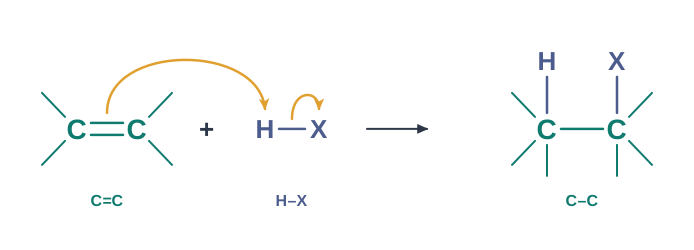
Fig. 6.12 – Addition of a hydrogen halide to an alkene.
\[\\mathrm{C{=}C} + \\mathrm{HX} \\rightarrow \\mathrm{H{-}C{-}C{-}X}\]
When the alkene is unsymmetrical, the halogen has a choice of two carbons, and one product predominates. Propene yields two products, however only one predominates as per Markovnikov’s rule (Chapter 13, Class XI).
\[\\mathrm{CH\_3CH{=}CH\_2} + \\mathrm{H{-}I} \\rightarrow \\underset{\\text{minor}}{\\mathrm{CH\_3CH\_2CH\_2I}} + \\underset{\\text{major}}{\\mathrm{CH\_3CHICH\_3}}\]
The rule is not an arbitrary recipe. The proton adds first, and it adds in whichever way produces the more stable carbocation; the halide ion then simply captures that carbocation. In propene, adding \(\\mathrm{H^+}\) to the terminal \(\\mathrm{CH\_2}\) gives a secondary carbocation, which is more stable than the primary one formed the other way round. The iodide therefore ends up on the middle carbon, giving 2-iodopropane as the major product.
(ii) Addition of halogens. In the laboratory, addition of bromine in \(\\mathrm{CCl\_4}\) to an alkene resulting in discharge of the reddish brown colour of bromine constitutes an important method for the detection of a double bond in a molecule. The addition results in the synthesis of vic-dibromides, which are colourless (Chapter 9, Class XI).
\[\\mathrm{CH\_2{=}CH\_2} + \\mathrm{Br\_2} \\xrightarrow{\\ \\mathrm{CCl\_4}\\ } \\underset{\\textit{vic}\\text{-Dibromide}}{\\mathrm{BrCH\_2{-}CH\_2Br}}\]
6.4.3 Halogen Exchange
Alkyl iodides are often prepared by the reaction of alkyl chlorides or bromides with \(\\mathrm{NaI}\) in dry acetone. This reaction is known as the Finkelstein reaction.
\[\\mathrm{R{-}X} + \\mathrm{NaI} \\rightarrow \\mathrm{R{-}I} + \\mathrm{NaX} \\qquad (\\mathrm{X} = \\mathrm{Cl},\\ \\mathrm{Br})\]
At first sight this looks like a reaction that should not work: iodide is being asked to displace chloride, and the C—I bond formed is weaker than the C—Cl bond broken. The trick lies in the solvent. \(\\mathrm{NaI}\) dissolves in dry acetone, but the \(\\mathrm{NaCl}\) or \(\\mathrm{NaBr}\) thus formed is precipitated in dry acetone. Removing a product from the solution facilitates the forward reaction according to Le Chatelier’s principle.
The synthesis of alkyl fluorides is best accomplished by heating an alkyl chloride or bromide in the presence of a metallic fluoride such as \(\\mathrm{AgF}\), \(\\mathrm{Hg\_2F\_2}\), \(\\mathrm{CoF\_2}\) or \(\\mathrm{SbF\_3}\). The reaction is termed the Swarts reaction.
\[\\mathrm{H\_3C{-}Br} + \\mathrm{AgF} \\rightarrow \\mathrm{H\_3C{-}F} + \\mathrm{AgBr}\]
6.5 Preparation of Haloarenes
In a haloarene the halogen sits on an \(sp^2\) carbon of the ring, and Section 6.4 has already told us that the alcohol route is closed to us here. Two routes remain: attack the ring directly with a halogen, or build the halide in from an amino group already present on the ring.
(i) From Hydrocarbons by Electrophilic Substitution
Aryl chlorides and bromides can be easily prepared by electrophilic substitution of arenes with chlorine and bromine respectively, in the presence of Lewis acid catalysts like iron or iron(III) chloride. A Lewis acid is an electron-pair acceptor; here its job is to polarise the \(\\mathrm{X\_2}\) molecule so strongly that it can act as the electrophile \(\\mathrm{X^+}\), which the electron-rich benzene ring then attacks.
With toluene, substitution occurs at the positions ortho and para to the methyl group, giving a mixture of the two products shown in Fig. 6.13.
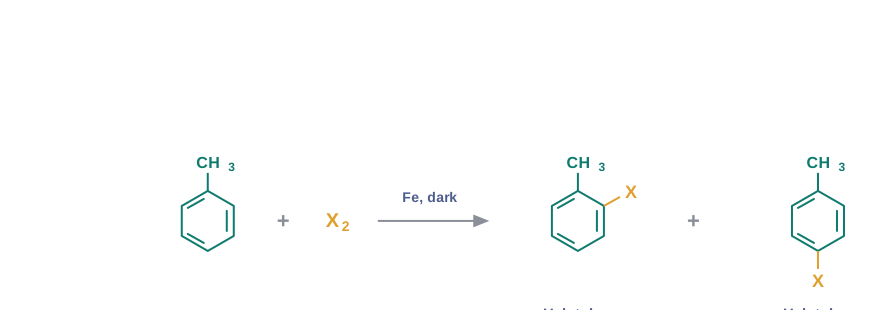
Fig. 6.13 – Electrophilic halogenation of toluene.
\[\\mathrm{C\_6H\_5CH\_3} + \\mathrm{X\_2} \\xrightarrow\[\\text{dark}]{\\ \\mathrm{Fe}\\ } o\\text{-Halotoluene} + p\\text{-Halotoluene}\]
The ortho and para isomers can be easily separated due to the large difference in their melting points. (You will see in Section 6.6 why the para isomer melts so much higher: its symmetry lets it pack into a crystal lattice far better than the ortho isomer can.)
Two limitations of this method must be remembered.
Reactions with iodine are reversible in nature. As iodobenzene forms, hydrogen iodide is also produced, and the \(\\mathrm{HI}\) attacks the product and pushes the reaction backwards. The reaction therefore requires the presence of an oxidising agent (\(\\mathrm{HNO\_3}\), \(\\mathrm{HIO\_4}\)) to oxidise the \(\\mathrm{HI}\) formed during iodination, thereby removing it and driving the equilibrium forward.
Fluoro compounds are not prepared by this method due to the high reactivity of fluorine. Fluorine attacks the ring so violently that substitution cannot be controlled, and mixtures and even ring destruction result.
(ii) From Amines by Sandmeyer’s Reaction
When a primary aromatic amine, dissolved or suspended in cold aqueous mineral acid, is treated with sodium nitrite, a diazonium salt is formed. This step is called diazotisation and is carried out at 273–278 K.
\[\\mathrm{C\_6H\_5NH\_2} \\xrightarrow\[273\\text{–}278\\ \\text{K}]{\\ \\mathrm{NaNO\_2} + \\mathrm{HX}\\ } \\underset{\\text{Benzene diazonium halide}}{\\mathrm{C\_6H\_5N\_2^{+}X^{-}}}\]
Mixing the solution of the freshly prepared diazonium salt with cuprous chloride or cuprous bromide results in the replacement of the diazonium group by \(-\\mathrm{Cl}\) or \(-\\mathrm{Br}\). The leaving group here is nitrogen gas itself — which is why the reaction is essentially irreversible.
\[\\mathrm{C\_6H\_5N\_2^{+}X^{-}} \\xrightarrow{\\ \\mathrm{Cu\_2X\_2}\\ } \\underset{\\text{Aryl halide}}{\\mathrm{C\_6H\_5X}} + \\mathrm{N\_2} \\qquad (\\mathrm{X} = \\mathrm{Cl},\\ \\mathrm{Br})\]
Replacement of the diazonium group by iodine does not require the presence of a cuprous halide and is done simply by shaking the diazonium salt with potassium iodide.
\[\\mathrm{C\_6H\_5N\_2^{+}X^{-}} \\xrightarrow{\\ \\mathrm{KI}\\ } \\mathrm{C\_6H\_5I} + \\mathrm{N\_2}\]
The whole sequence is set out in FC 6.2.
Flowchart
FC 6.2 – Sandmeyer route to haloarenes: Aniline (C₆H₅NH₂) in cold aqueous mineral acid → NaNO₂ + HX at 273–278 K → benzene diazonium halide (C₆H₅N₂⁺X⁻) → branch A: Cu₂Cl₂ → chlorobenzene + N₂; branch B: Cu₂Br₂ → bromobenzene + N₂; branch C: KI (no cuprous halide needed) → iodobenzene + N₂
6.6 Physical Properties
Alkyl halides are colourless when pure. However, bromides and iodides develop colour when exposed to light, because light slowly breaks the weak C—Br and C—I bonds and liberates traces of free bromine or iodine. Many volatile halogen compounds have a sweet smell.
Melting and Boiling Points
Methyl chloride, methyl bromide, ethyl chloride and some chlorofluoromethanes are gases at room temperature. Higher members are liquids or solids.
As we have already learnt, molecules of organic halogen compounds are generally polar, because of the polarised C—X bond of Section 6.3. Due to greater polarity as well as higher molecular mass as compared to the parent hydrocarbon, the intermolecular forces of attraction (dipole-dipole and van der Waals) are stronger in the halogen derivatives. That is why the boiling points of chlorides, bromides and iodides are considerably higher than those of the hydrocarbons of comparable molecular mass. Boiling means pulling molecules apart from one another, so the stronger these attractions, the more heat must be supplied, and the higher the boiling point.
The attractions get stronger as the molecules get bigger in size and have more electrons. A large, electron-rich cloud is loosely held and easily distorted, so momentary dipoles arise readily, and van der Waals forces grow. The pattern of variation of boiling points of different halides is depicted in Fig. 6.14.
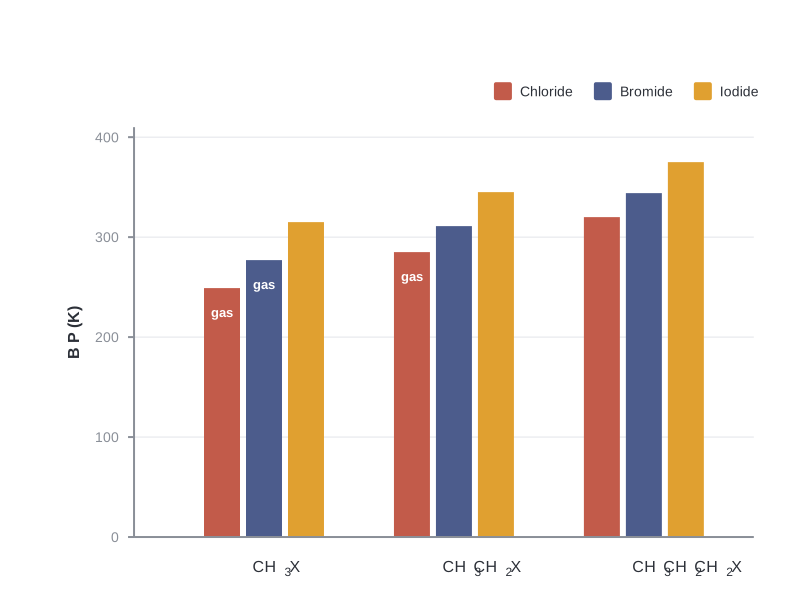
Fig. 6.14 – Boiling points of some alkyl chlorides, bromides and iodides.
For the same alkyl group, the boiling points of alkyl halides decrease in the order: RI > RBr > RCl > RF. This is because with the increase in size and mass of the halogen atom, the magnitude of van der Waals forces increases.
The Effect of Branching
The boiling points of isomeric haloalkanes decrease with increase in branching. For example, 2-bromo-2-methylpropane has the lowest boiling point among the three isomers, as the values in Fig. 6.15 show.
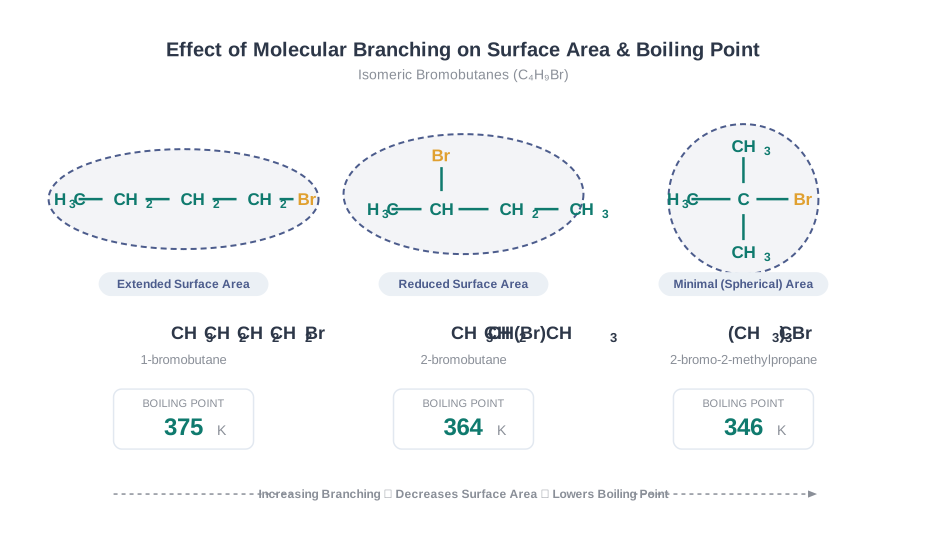
Fig. 6.15 – Effect of branching on the boiling point of isomeric bromobutanes.
Isomeric Dihalobenzenes
Boiling points of isomeric dihalobenzenes are very nearly the same. However, the para-isomers are high melting as compared to their ortho- and meta-isomers. It is due to the symmetry of para-isomers that fits in the crystal lattice better as compared to ortho- and meta-isomers. Melting is the breaking down of an orderly crystal lattice, so a molecule that packs neatly and closely needs more heat to be dislodged; boiling, by contrast, depends mainly on intermolecular attraction in the liquid, which is much the same for all three isomers.
The figures for the three isomeric dichlorobenzenes make the point clearly.
Boiling and melting points of the isomeric dichlorobenzenes (in K)
| ortho | meta | para | |
|---|---|---|---|
| b.p./K | 453 | 446 | 448 |
| m.p./K | 256 | 249 | 323 |
Notice how little the boiling points differ — barely 7 K across the set — while the melting point of the para isomer stands nearly 70 K above the others.
Density
Bromo, iodo and polychloro derivatives of hydrocarbons are heavier than water. The density increases with increase in the number of carbon atoms, halogen atoms and the atomic mass of the halogen atoms (Table 6.3). All three factors work the same way: each adds mass to the molecule without adding a proportionate amount of volume.
Table 6.3: Density of Some Haloalkanes
| Compound | Density (g/mL) | Compound | Density (g/mL) |
|---|---|---|---|
| \(\\mathrm{n{-}C\_3H\_7Cl}\) | 0.89 | \(\\mathrm{CH\_2Cl\_2}\) | 1.336 |
| \(\\mathrm{n{-}C\_3H\_7Br}\) | 1.335 | \(\\mathrm{CHCl\_3}\) | 1.489 |
| \(\\mathrm{n{-}C\_3H\_7I}\) | 1.747 | \(\\mathrm{CCl\_4}\) | 1.595 |
Solubility
The haloalkanes are very slightly soluble in water. To understand why, follow the energy bookkeeping. In order to dissolve a haloalkane in water, energy is required to overcome the attractions between the haloalkane molecules and to break the hydrogen bonds between water molecules. Less energy is released when new attractions are set up between the haloalkane and the water molecules, as these are not as strong as the original hydrogen bonds in water. The books do not balance — more energy goes in than comes out — and as a result, the solubility of haloalkanes in water is low.
However, haloalkanes tend to dissolve in organic solvents. Here the bookkeeping works out, because the new intermolecular attractions between haloalkane and solvent molecules have much the same strength as the ones being broken in the separate haloalkane and solvent molecules. Nothing is lost, so mixing proceeds freely.
6.7 Chemical Reactions
6.7.1 Reactions of Haloalkanes
The reactions of haloalkanes may be divided into the following three categories:
- Nucleophilic substitution
- Elimination reactions
- Reaction with metals
All three follow from the same structural fact established in Section 6.3 — the C—X bond is polarised, leaving the carbon electron-deficient and the halogen ready to depart with the bonding electron pair.
(1) Nucleophilic Substitution Reactions
You have learnt in Class XI that nucleophiles are electron rich species. Therefore, they attack that part of the substrate molecule which is electron deficient. In a haloalkane, that part is the carbon atom bonded to the halogen, which carries a partial positive charge.
The reaction in which a nucleophile replaces an already existing nucleophile in a molecule is called a nucleophilic substitution reaction. Haloalkanes are the substrate in these reactions. In this type of reaction, a nucleophile reacts with the haloalkane (the substrate), which has a partial positive charge on the carbon atom bonded to halogen. A substitution reaction takes place, and the halogen atom — called the leaving group — departs as a halide ion. Since the substitution reaction is initiated by a nucleophile, it is called a nucleophilic substitution reaction. The general pattern is shown in Fig. 6.16.
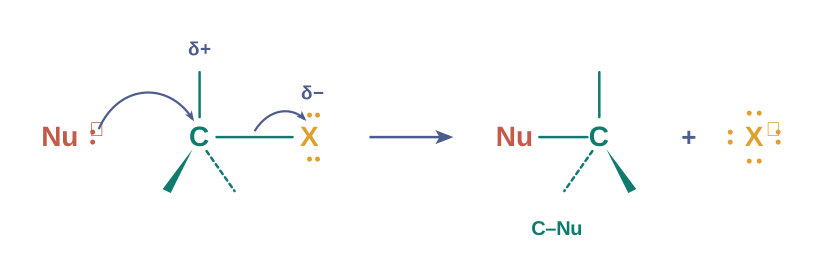
Fig. 6.16 – General mechanism of nucleophilic substitution.
\[\\mathrm{Nu^{-}} + \\mathrm{R{-}X} \\rightarrow \\mathrm{R{-}Nu} + \\mathrm{X^{-}}\]
It is one of the most useful classes of organic reactions of alkyl halides in which halogen is bonded to \(sp^3\) hybridised carbon. By choosing the nucleophile, a chemist can convert the same alkyl halide into an alcohol, an ether, an amine, a nitrile and much else — which is exactly why haloalkanes are such valuable starting materials. The products formed by the reaction of haloalkanes with some common nucleophiles are given in Table 6.4.
Table 6.4: Nucleophilic Substitution of Alkyl Halides (R–X)
\[\\mathrm{R{-}X} + \\mathrm{Nu^{-}} \\rightarrow \\mathrm{R{-}Nu} + \\mathrm{X^{-}}\]
| Reagent | Nucleophile (Nu⁻) | Substitution product R–Nu | Class of main product |
|---|---|---|---|
| NaOH (KOH) | HO⁻ | ROH | Alcohol |
| H₂O | H₂O | ROH | Alcohol |
| NaOR′ | R′O⁻ | ROR′ | Ether |
| NaI | I⁻ | R—I | Alkyl iodide |
| NH₃ | NH₃ | RNH₂ | Primary amine |
| R′NH₂ | R′NH₂ | RNHR′ | Sec. amine |
| R′R″NH | R′R″NH | RNR′R″ | Tert. amine |
| KCN | ⁻C≡N: | RCN | Nitrile (cyanide) |
| AgCN | Ag–CN: | RNC (isocyanide) | Isonitrile (isocyanide) |
| KNO₂ | O=N—O⁻ | R—O—N=O | Alkyl nitrite |
| AgNO₂ | Ag—Ö—N=O | R—NO₂ | Nitroalkane |
| R′COOAg | R′COO⁻ | R′COOR | Ester |
| LiAlH₄ | H⁻ | RH | Hydrocarbon |
| R′⁻ M⁺ | R′⁻ | RR′ | Alkane |
Ambident Nucleophiles
Groups like cyanides and nitrites possess two nucleophilic centres and are called ambident nucleophiles. Actually, the cyanide group is a hybrid of two contributing structures and therefore can act as a nucleophile in two different ways, \(\[^{-}\\mathrm{C{\\equiv}N} \\leftrightarrow \\mathrm{:C{=}N^{-}}]\) — that is, linking through the carbon atom, resulting in alkyl cyanides, or through the nitrogen atom, leading to isocyanides.
Similarly, the nitrite ion also represents an ambident nucleophile with two different points of linkage, \(\[^{-}\\mathrm{O{-}N{=}O}]\). The linkage through oxygen results in alkyl nitrites, while linkage through the nitrogen atom leads to nitroalkanes.
Mechanism
This reaction has been found to proceed by two different mechanisms, which are described below.
(a) Substitution Nucleophilic Bimolecular (\(\\mathrm{S\_N}2\))
The reaction between \(\\mathrm{CH\_3Cl}\) and hydroxide ion to yield methanol and chloride ion follows second order kinetics — i.e., the rate depends upon the concentration of both the reactants.
\[\\text{rate} = k\[\\mathrm{CH\_3Cl}]\[\\mathrm{OH^{-}}]\]
The rate law is the clue to the mechanism. If doubling the concentration of either reactant doubles the rate, then both species must be present in the slowest step. In other words, the nucleophile and the substrate must be colliding in the rate-determining step — the reaction is bimolecular.
\[\\mathrm{OH^{-}} + \\mathrm{CH\_3Cl} \\rightarrow \[\\text{transition state}] \\rightarrow \\mathrm{CH\_3OH} + \\mathrm{Cl^{-}}\]
The wedge-and-dash convention is used throughout what follows: the solid wedge represents the bond coming out of the paper, the dashed line a bond going down behind the paper, and a straight line a bond lying in the plane of the paper.
The reaction can be represented diagrammatically as shown in Fig. 6.17.
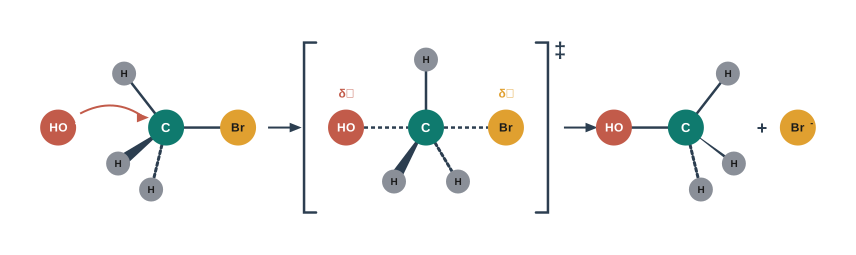
Fig. 6.17 – Mechanism of the SN2 reaction.
It depicts a bimolecular nucleophilic substitution (\(\\mathrm{S\_N}2\)) reaction; the incoming nucleophile interacts with the alkyl halide, causing the carbon–halide bond to break, and a new bond is formed between carbon and the attacking nucleophile. Here it is the C—O bond formed between C and \(-\\mathrm{OH}\). These two processes take place simultaneously in a single step, and no intermediate is formed.
As the reaction progresses and the bond between the incoming nucleophile and the carbon atom starts forming, the bond between the carbon atom and the leaving group weakens. As this happens, the three carbon–hydrogen bonds of the substrate start moving away from the attacking nucleophile. In the transition state, all three C—H bonds lie in the same plane, and the attacking and leaving nucleophiles are partially attached to the carbon. As the attacking nucleophile approaches closer to the carbon, the C—H bonds still keep on moving in the same direction, till the attacking nucleophile attaches to carbon and the leaving group leaves the carbon.
As a result, the configuration is inverted: the configuration of the carbon atom under attack inverts in much the same way as an umbrella is turned inside out when caught in a strong wind. This process is called inversion of configuration.
Configuration. The spatial arrangement of functional groups around a carbon is called its configuration. Consider two structures (A) and (B) of the same compound, drawn either side of a mirror line, as in Fig. 6.18. They differ in the spatial arrangement of the functional groups attached to carbon. Structure (A) is the mirror image of structure (B); we say that the configuration of carbon in structure (A) is the mirror image of the configuration of carbon in structure (B).
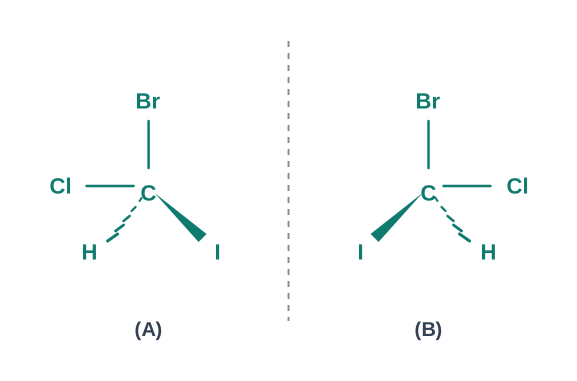
Fig. 6.18 – Two configurations of an asymmetric carbon.
In the transition state, the carbon atom is simultaneously bonded to the incoming nucleophile and the outgoing leaving group. Such structures are unstable and cannot be isolated. Thus, in the transition state, carbon is simultaneously bonded to five atoms.
Steric effects. Since this reaction requires the approach of the nucleophile to the carbon bearing the leaving group, the presence of bulky substituents on or near the carbon atom has a dramatic inhibiting effect. Of the simple alkyl halides, methyl halides react most rapidly in \(\\mathrm{S\_N}2\) reactions, because there are only three small hydrogen atoms in the way. Tertiary halides are the least reactive, because bulky groups hinder the approaching nucleophile. The relative rates are shown in Fig. 6.19.
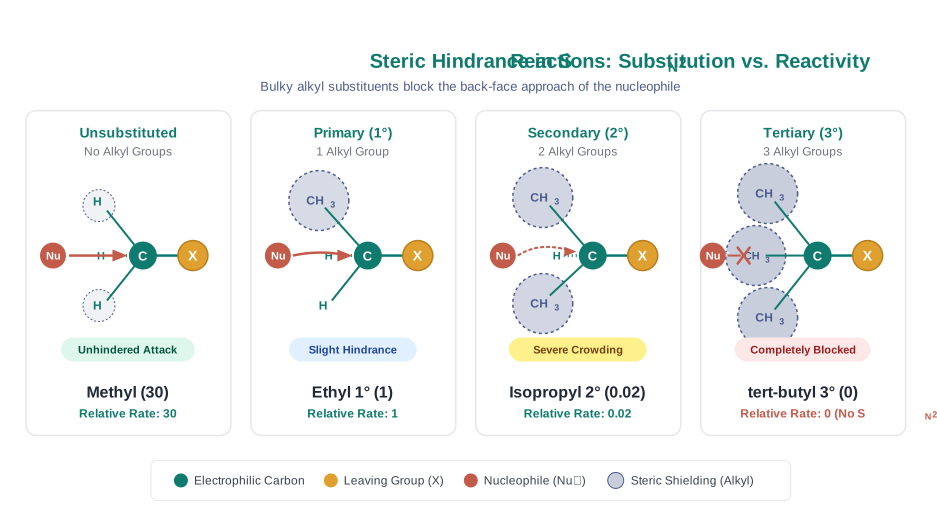
Fig. 6.19 – Steric hindrance in the SN2 reaction.
Thus the order of reactivity followed is:
\[\\text{Primary halide} > \\text{Secondary halide} > \\text{Tertiary halide}\]
(b) Substitution Nucleophilic Unimolecular (\(\\mathrm{S\_N}1\))
\(\\mathrm{S\_N}1\) reactions are generally carried out in polar protic solvents (like water, alcohol, acetic acid, etc.). A polar protic solvent has O—H or N—H bonds, so it can both stabilise ions and hydrogen-bond to the departing halide — and, as we shall see, that is precisely the help this mechanism needs.
The reaction between tert-butyl bromide and hydroxide ion yields tert-butyl alcohol and follows first order kinetics — i.e., the rate of reaction depends upon the concentration of only one reactant, which is tert-butyl bromide.
\[\\underset{\\text{2-Bromo-2-methylpropane}}{\\mathrm{(CH\_3)\_3CBr}} + \\mathrm{^{-}OH} \\rightarrow \\underset{\\text{2-Methylpropan-2-ol}}{\\mathrm{(CH\_3)\_3COH}} + \\mathrm{Br^{-}}\]
\[\\text{rate} = k\[\\mathrm{(CH\_3)\_3CBr}]\]
It occurs in two steps. In step I, the polarised C—Br bond undergoes slow cleavage to produce a carbocation and a bromide ion. The carbocation thus formed is then attacked by the nucleophile in step II to complete the substitution reaction.
\[\\text{Step I: } \\mathrm{(CH\_3)\_3CBr} \\rightleftharpoons \\mathrm{(CH\_3)\_3C^{+}} + \\mathrm{Br^{-}} \\quad (\\text{slow})\]
\[\\text{Step II: } \\mathrm{(CH\_3)\_3C^{+}} + \\mathrm{^{-}OH} \\rightarrow \\mathrm{(CH\_3)\_3COH} \\quad (\\text{fast})\]
The two-step logic is set out in FC 6.3.
Flowchart
FC 6.3 – SN1 mechanism: R–X → slow, reversible heterolysis of the C–X bond (rate-determining step) → planar carbocation R⁺ + X⁻ → fast attack by the nucleophile Nu⁻ → product R–Nu; branch: because the carbocation is planar, Nu⁻ can attack from either face → two products of opposite configuration → racemisation at a chiral centre
Step I is the slowest and reversible. It involves the C—Br bond breaking, for which the energy is obtained through solvation of the halide ion with the proton of the protic solvent. Since the rate of reaction depends upon the slowest step, the rate of reaction depends only on the concentration of the alkyl halide and not on the concentration of the hydroxide ion. This is why the hydroxide ion, though essential to the product, never appears in the rate law: it joins the party only after the rate-determining step is over.
Further, the greater the stability of the carbocation, the greater will be its ease of formation from the alkyl halide, and the faster will be the rate of reaction. In the case of alkyl halides, 3° alkyl halides undergo \(\\mathrm{S\_N}1\) reaction very fast because of the high stability of 3° carbocations.
We can sum up the order of reactivity of alkyl halides towards \(\\mathrm{S\_N}1\) and \(\\mathrm{S\_N}2\) reactions as follows:
\[\\text{For } \\mathrm{S\_N}2:\\ \\mathrm{CH\_3X} > \\text{Primary halide} > \\text{Secondary halide} > \\text{Tertiary halide}\]
\[\\text{For } \\mathrm{S\_N}1:\\ \\text{Tertiary halide} > \\text{Secondary halide} > \\text{Primary halide} > \\mathrm{CH\_3X}\]
For the same reasons, allylic and benzylic halides show high reactivity towards the \(\\mathrm{S\_N}1\) reaction. The carbocation thus formed gets stabilised through resonance (Chapter 8, Class XI), as shown in Fig. 6.20: the positive charge is not trapped on one carbon but is spread over several, and a spread-out charge is a stabilised charge.
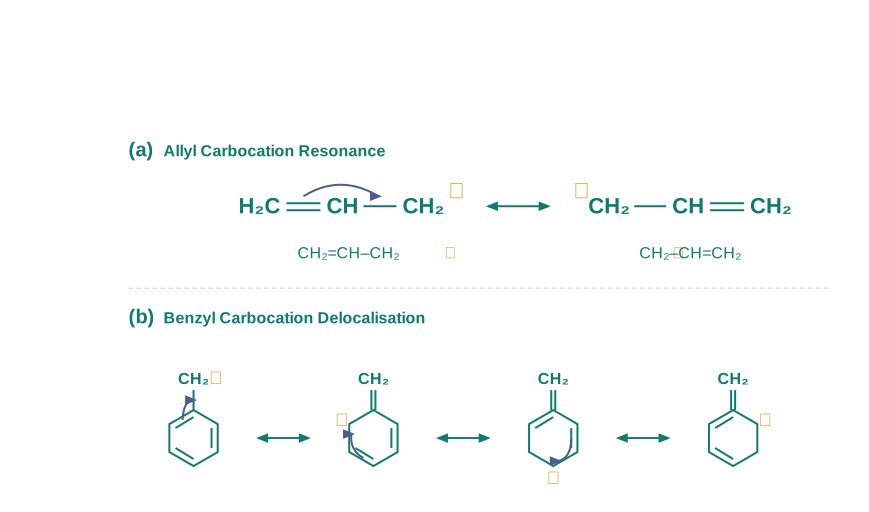
Fig. 6.20 – Resonance stabilisation of allyl and benzyl carbocations.
For a given alkyl group, the reactivity of the halide, R–X, follows the same order in both the mechanisms:
\[\\mathrm{R{-}I} > \\mathrm{R{-}Br} > \\mathrm{R{-}Cl} \\gg \\mathrm{R{-}F}\]
This order simply repeats what Table 6.2 already told us: the weaker the C—X bond, the more readily the halide leaves, whichever mechanism is operating.
(c) Stereochemical Aspects of Nucleophilic Substitution Reactions
In order to understand the stereochemical aspects of substitution reactions, we need to learn some basic stereochemical principles and notations (optical activity, chirality, retention, inversion, racemisation, etc.).
(i) Optical activity. The plane of plane polarised light — produced by passing ordinary light through a Nicol prism — is rotated when it is passed through the solutions of certain compounds. Such compounds are called optically active compounds. The angle by which the plane polarised light is rotated is measured by an instrument called a polarimeter.
If the compound rotates the plane of plane polarised light to the right, i.e., in the clockwise direction, it is called dextrorotatory (Greek for “right rotating”) or the d-form, and this is indicated by placing a positive (+) sign before the degree of rotation. If the light is rotated towards the left (anticlockwise direction), the compound is said to be laevorotatory or the l-form, and a negative (–) sign is placed before the degree of rotation. Such (+) and (–) isomers of a compound are called optical isomers, and the phenomenon is termed optical isomerism.
(ii) Molecular asymmetry, chirality and enantiomers. The observation of Louis Pasteur (1848) that crystals of certain compounds exist in the form of mirror images laid the foundation of modern stereochemistry. He demonstrated that aqueous solutions of both types of crystals showed optical rotation, equal in magnitude (for solutions of equal concentration) but opposite in direction. He believed that this difference in optical activity was associated with the three dimensional arrangements of atoms in the molecules (configurations) of the two types of crystals.
Dutch scientist J. Van’t Hoff and French scientist C. Le Bel, in the same year (1874), independently argued that the spatial arrangement of four groups (valencies) around a central carbon is tetrahedral, and that if all the substituents attached to that carbon are different, the mirror image of the molecule is not superimposed (overlapped) on the molecule. Such a carbon is called an asymmetric carbon or stereocentre. The resulting molecule would lack symmetry and is referred to as an asymmetric molecule. The asymmetry of the molecule, along with the non-superimposability of mirror images, is responsible for the optical activity in such organic compounds.
The symmetry and asymmetry are also observed in many day-to-day objects. A sphere, a cube and a cone are all identical to their mirror images and can be superimposed. However, many objects are non-superimposable on their mirror images. For example, your left and right hand look similar, but if you put your left hand on your right hand by moving them in the same plane, they do not coincide. The objects which are non-superimposable on their mirror image (like a pair of hands) are said to be chiral, and this property is known as chirality. Chiral molecules are optically active, while objects which are superimposable on their mirror images are called achiral. These molecules are optically inactive. Some common examples of chiral and achiral objects are shown in Fig. 6.21.
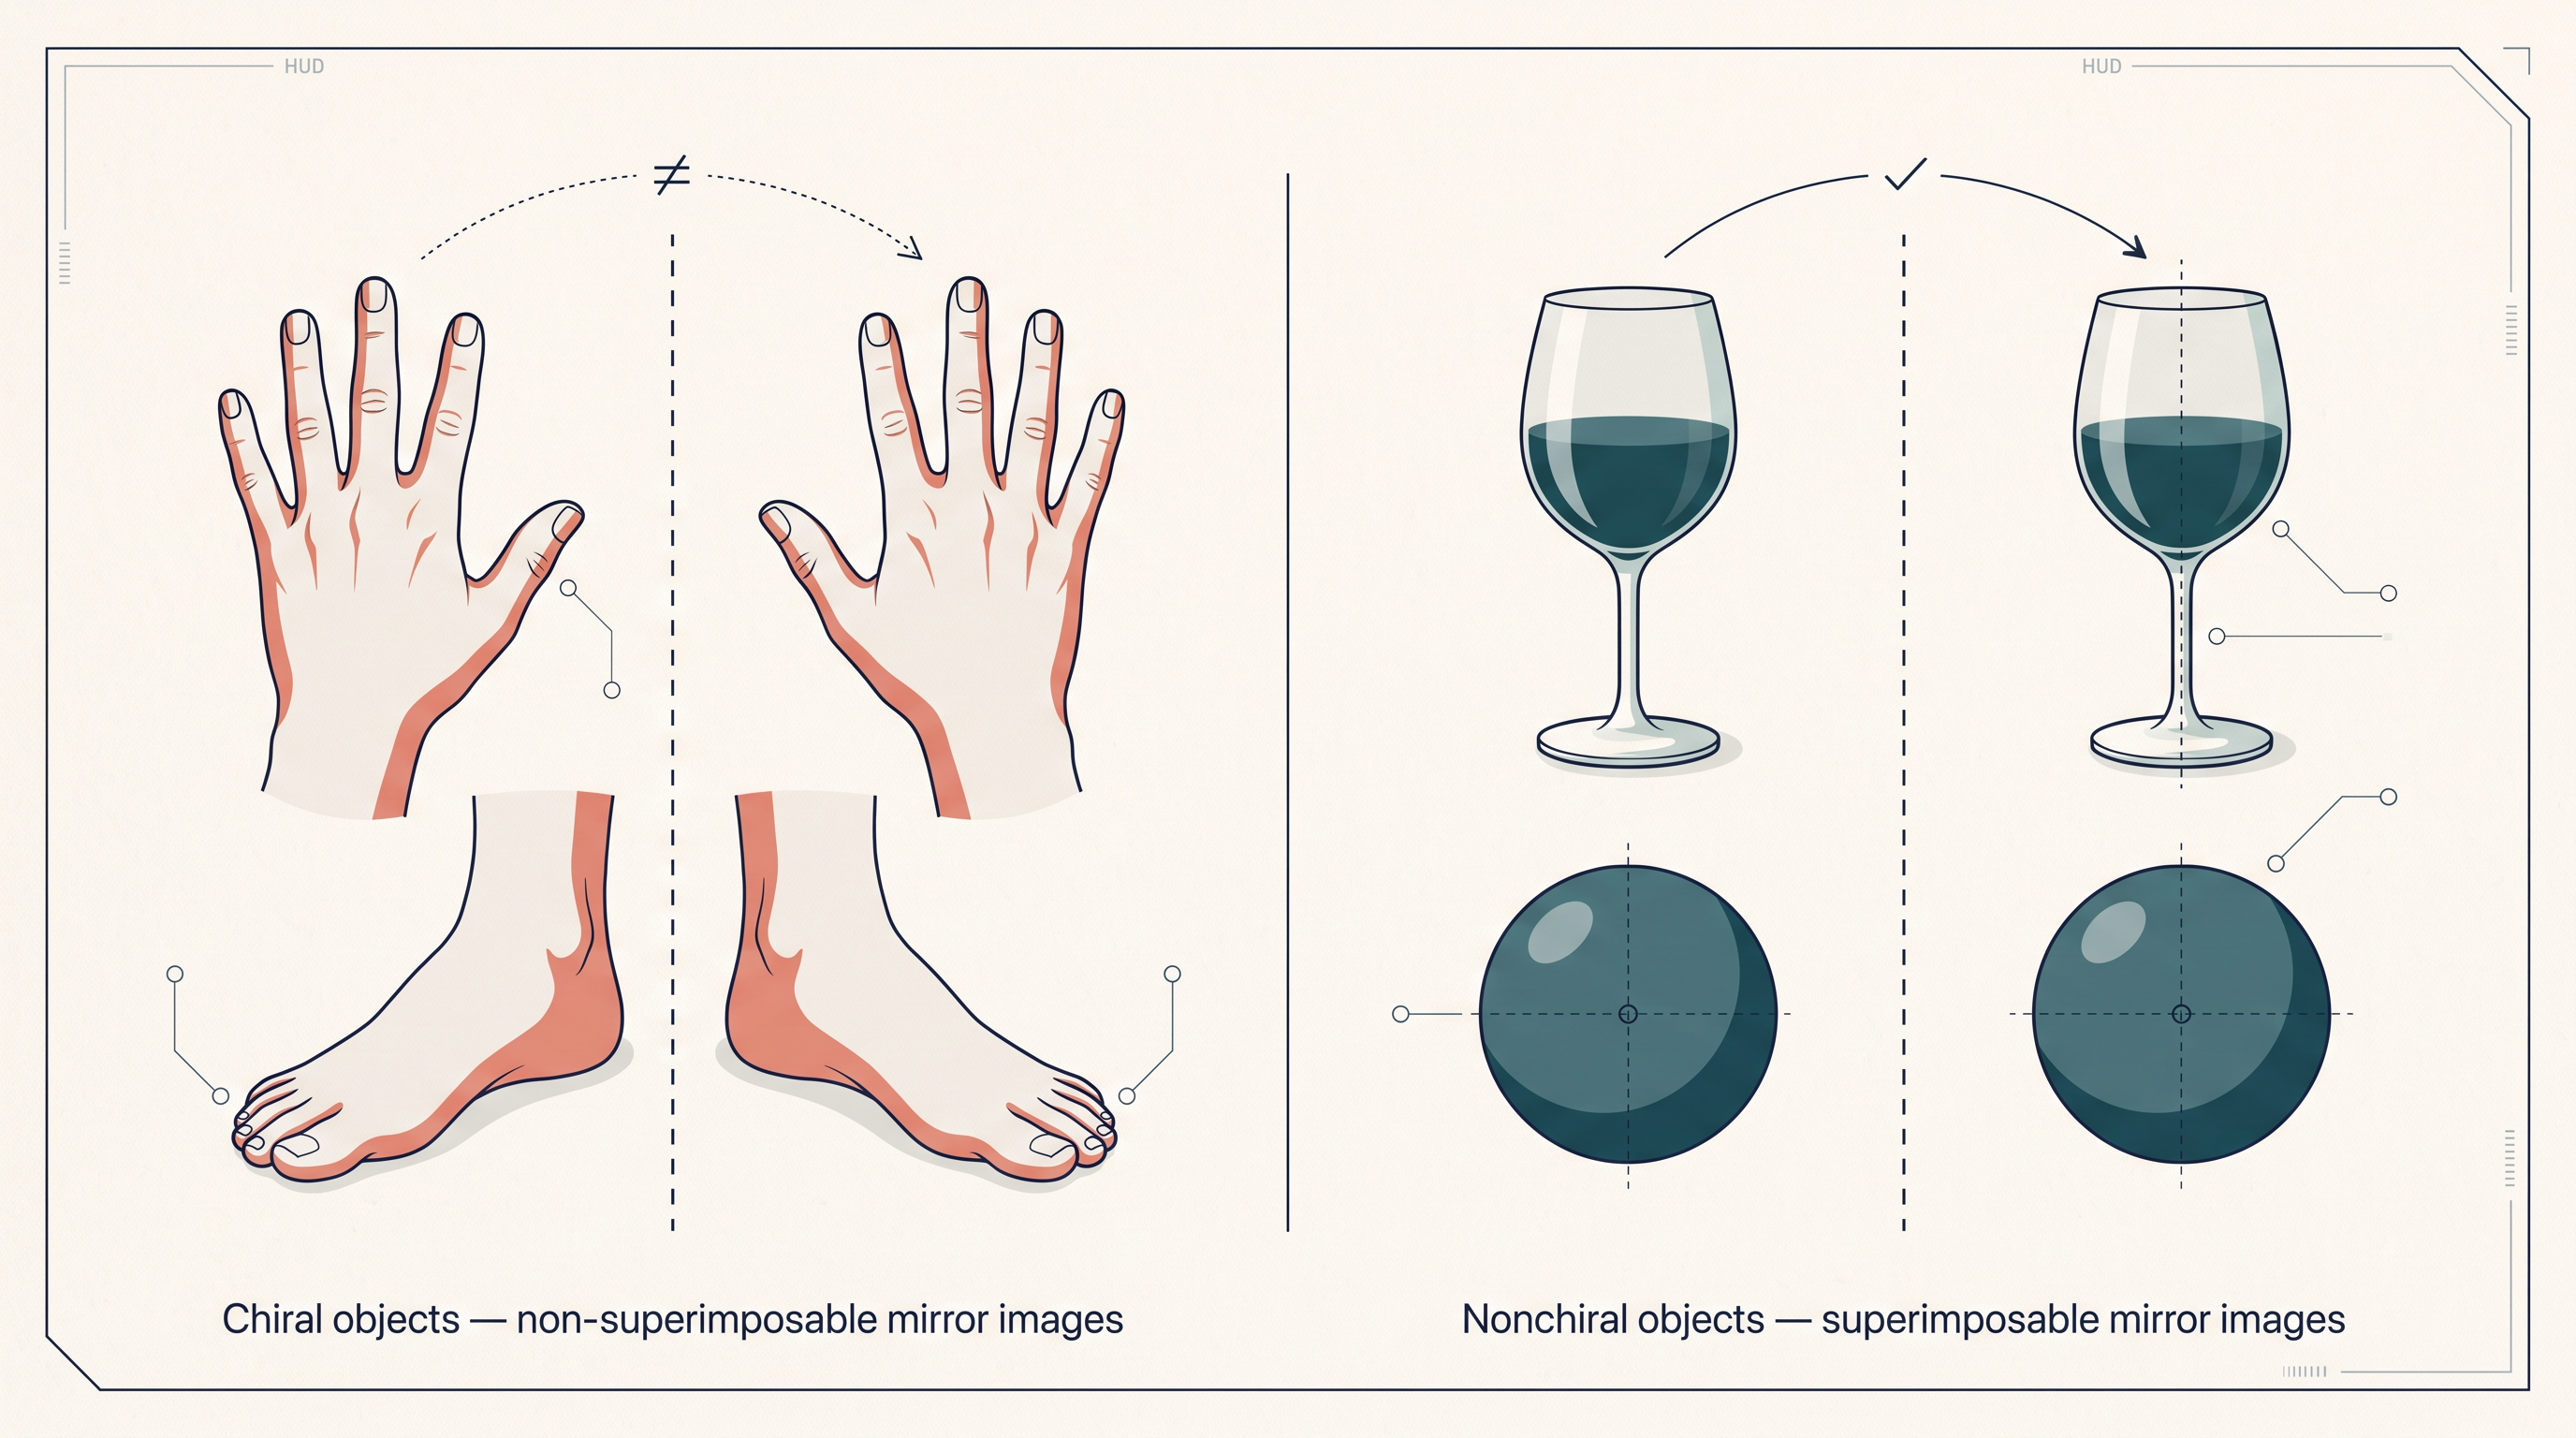
Fig. 6.21 – Chiral and achiral objects.
The above test of molecular chirality can be applied to organic molecules by constructing models and their mirror images, or by drawing three dimensional structures and attempting to superimpose them in our minds. There are other aids, however, that can assist us in recognising chiral molecules. One such aid is the presence of a single asymmetric carbon atom.
Let us consider two simple molecules, propan-2-ol (Fig. 6.22) and butan-2-ol (Fig. 6.23), and their mirror images.
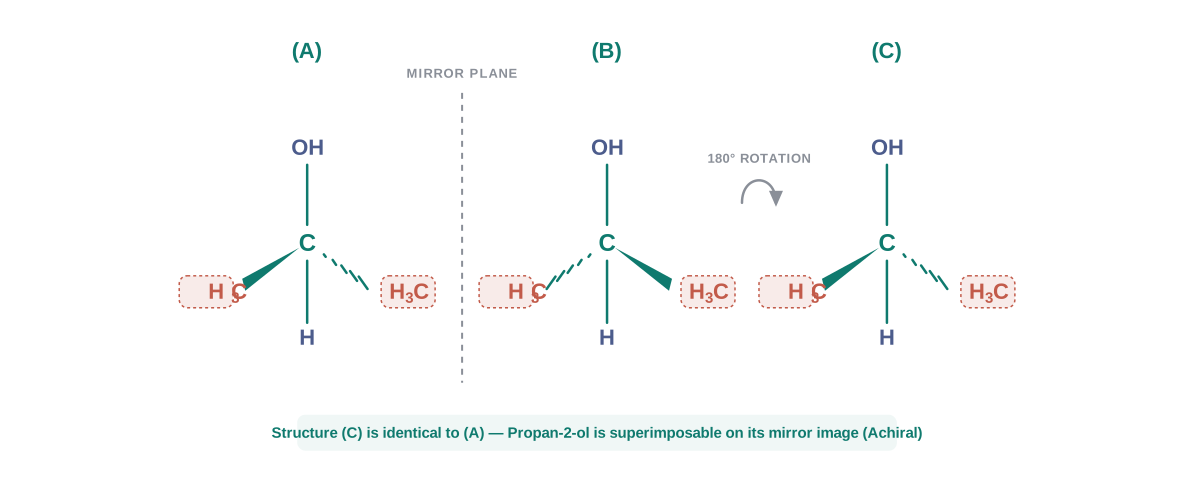
Fig. 6.22 – Propan-2-ol is an achiral molecule.
As you can see very clearly, propan-2-ol (A) does not contain an asymmetric carbon, as all the four groups attached to the tetrahedral carbon are not different — it carries two identical methyl groups. We rotate the mirror image (B) of the molecule by 180° (structure C) and try to overlap structure (C) with structure (A); these structures completely overlap. Thus propan-2-ol is an achiral molecule.
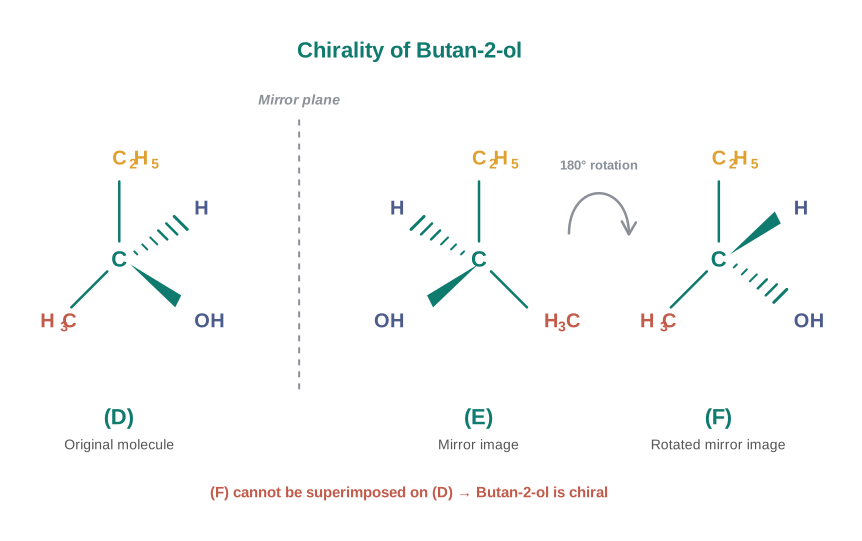
Fig. 6.23 – Butan-2-ol is a chiral molecule.
Butan-2-ol has four different groups attached to the tetrahedral carbon — a methyl group, an ethyl group, a hydrogen atom and a hydroxyl group — and as expected is chiral. Some common examples of chiral molecules are 2-chlorobutane, 2,3-dihydroxypropanal (\(\\mathrm{OHC{-}CHOH{-}CH\_2OH}\)), bromochloro-iodomethane (\(\\mathrm{BrClCHI}\)), 2-bromopropanoic acid (\(\\mathrm{H\_3C{-}CHBr{-}COOH}\)), etc.
The stereoisomers related to each other as non-superimposable mirror images are called enantiomers (Fig. 6.24). A and B in Fig. 6.22 and D and E in Fig. 6.23 are enantiomers.
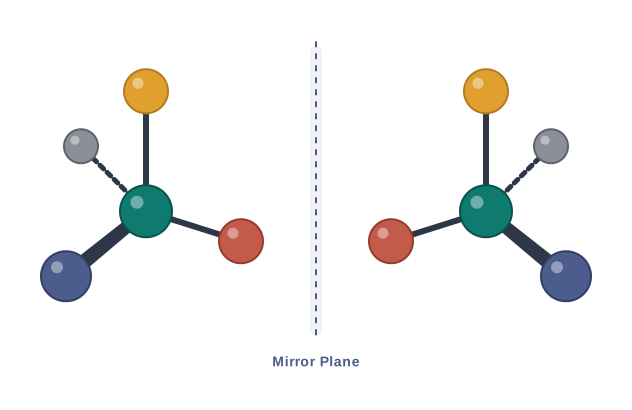
Fig. 6.24 – Non-superimposable mirror images of a chiral molecule.
Enantiomers possess identical physical properties — namely melting point, boiling point, refractive index, etc. They only differ with respect to the rotation of plane polarised light. If one of the enantiomers is dextrorotatory, the other will be laevorotatory. However, the sign of optical rotation is not necessarily related to the absolute (actual) configuration of the molecule — that is, knowing that a compound is (+) does not by itself tell you how its four groups are arranged in space.
A mixture containing two enantiomers in equal proportions will have zero optical rotation, as the rotation due to one isomer will be cancelled by the rotation due to the other isomer. Such a mixture is known as a racemic mixture or racemic modification. A racemic mixture is represented by prefixing dl or (±) before the name — for example, (±) butan-2-ol. The process of conversion of an enantiomer into a racemic mixture is known as racemisation.
(iii) Retention. Retention of configuration is the preservation of the spatial arrangement of bonds to an asymmetric centre during a chemical reaction or transformation.
In general, if during a reaction no bond to the stereocentre is broken, the product will have the same general configuration of groups around the stereocentre as that of the reactant. Such a reaction is said to proceed with retention of configuration. Consider as an example the reaction that takes place when (–)-2-methylbutan-1-ol is heated with concentrated hydrochloric acid, shown in Fig. 6.25.
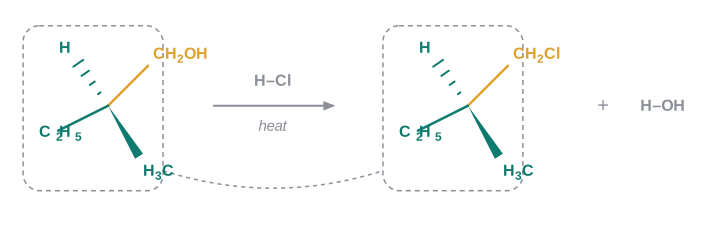
Fig. 6.25 – Reaction proceeding with retention of configuration.
It is important to note that the configuration at the asymmetric centre in the reactant and the product is the same, but the sign of optical rotation has changed in the product. This is so because two different compounds with the same configuration at the asymmetric centre may have different optical rotations. One may be dextrorotatory (plus sign of optical rotation) while the other may be laevorotatory (negative sign of optical rotation).
(iv) Inversion, retention and racemisation. There are three outcomes for a reaction at an asymmetric carbon atom, when a bond directly linked to the asymmetric carbon atom is broken. Consider the replacement of a group X by Y, as shown in Fig. 6.26.
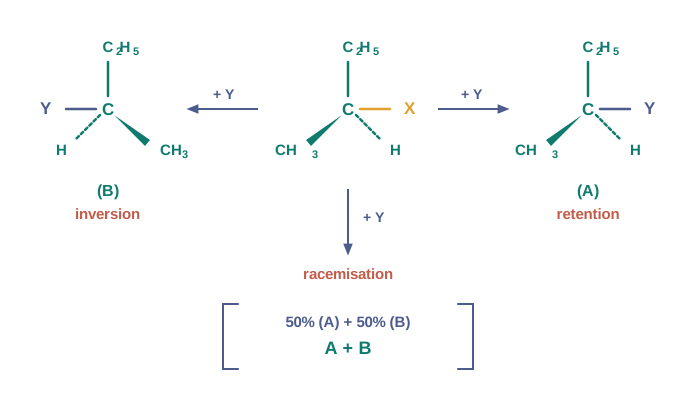
Fig. 6.26 – Retention, inversion and racemisation.
If (A) is the only compound obtained, the process is called retention of configuration. If (B) is the only compound obtained, the process is called inversion of configuration; the configuration has been inverted in B. If a 50:50 mixture of A and B is obtained, then the process is called racemisation, and the product is optically inactive, as one isomer will rotate the plane polarised light in the direction opposite to the other.
Now let us have a fresh look at \(\\mathrm{S\_N}1\) and \(\\mathrm{S\_N}2\) mechanisms by taking examples of optically active alkyl halides.
\(\\mathrm{S\_N}2\) and inversion. In the case of optically active alkyl halides, the product formed as a result of the \(\\mathrm{S\_N}2\) mechanism has the inverted configuration as compared to the reactant. This is because the nucleophile attaches itself on the side opposite to the one where the halogen atom is present — it has no choice, since the halogen is in the way on the front side. When (–)-2-bromooctane is allowed to react with sodium hydroxide, (+)-octan-2-ol is formed, with the \(-\\mathrm{OH}\) group occupying the position opposite to what bromide had occupied, as shown in Fig. 6.27.
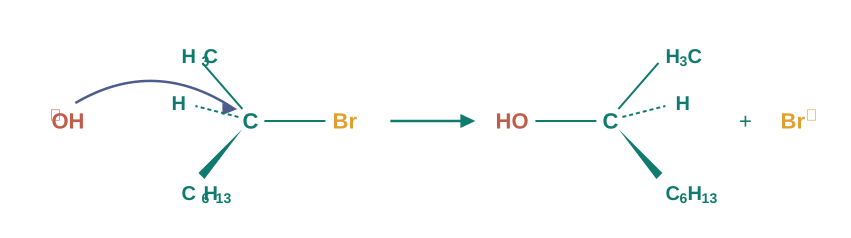
Fig. 6.27 – Inversion of configuration in the SN2 reaction.
Thus, \(\\mathrm{S\_N}2\) reactions of optically active halides are accompanied by inversion of configuration.
\(\\mathrm{S\_N}1\) and racemisation. In the case of optically active alkyl halides, \(\\mathrm{S\_N}1\) reactions are accompanied by racemisation. Can you think of the reason why it happens? Actually, the carbocation formed in the slow step, being \(sp^2\) hybridised, is planar (achiral). The attack of the nucleophile may be accomplished from either side of the plane of the carbocation, resulting in a mixture of products — one having the same configuration (the \(-\\mathrm{OH}\) attaching at the same position as the halide ion had occupied) and the other having the opposite configuration (the \(-\\mathrm{OH}\) attaching on the side opposite to the halide ion). This may be illustrated by the hydrolysis of optically active 2-bromobutane, which results in the formation of (±)-butan-2-ol, as drawn in Fig. 6.28.
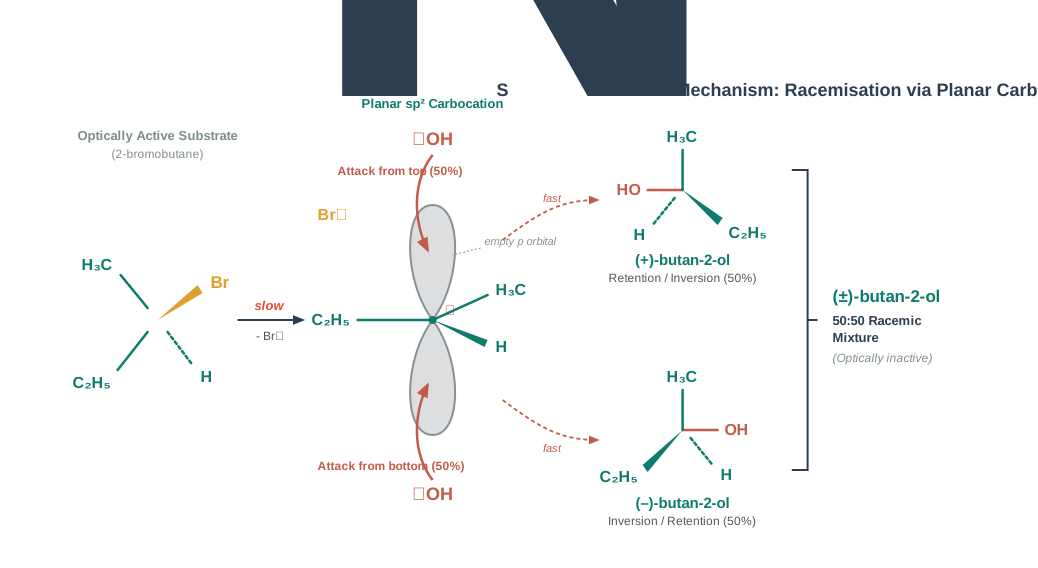
Fig. 6.28 – Racemisation in the SN1 reaction.
(2) Elimination Reactions
When a haloalkane with a \(\\beta\)-hydrogen atom is heated with an alcoholic solution of potassium hydroxide, there is elimination of a hydrogen atom from the \(\\beta\)-carbon and a halogen atom from the \(\\alpha\)-carbon atom.
Location of \(\\alpha\) and \(\\beta\) carbon in a molecule. The carbon on which the halogen atom is directly attached is called the \(\\alpha\)-carbon, and the carbon atom adjacent to this carbon is called the \(\\beta\)-carbon. A molecule with no \(\\beta\)-hydrogen atom at all cannot undergo this reaction — there is nothing to remove.
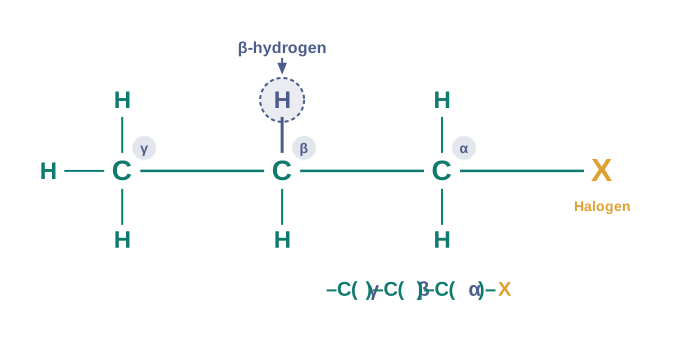
Fig. 6.29 – The α and β carbon atoms in an alkyl halide.
The base removes the \(\\beta\)-hydrogen as a proton, the electrons of that C—H bond swing in to form a \(\\pi\) bond, and the halide leaves from the \(\\alpha\)-carbon, as shown in Fig. 6.30.
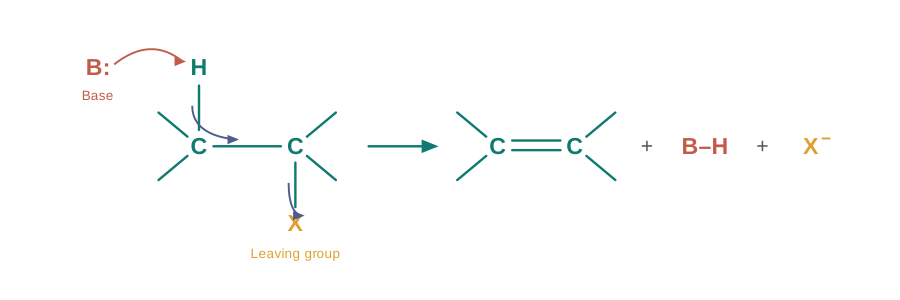
Fig. 6.30 – Mechanism of β-elimination.
\[\\mathrm{H{-}C\_\\beta{-}C\_\\alpha{-}X} + \\mathrm{B^{-}} \\rightarrow \\mathrm{C{=}C} + \\mathrm{B{-}H} + \\mathrm{X^{-}}\]
As a result, an alkene is formed as the product. Since a \(\\beta\)-hydrogen atom is involved in the elimination, it is often called \(\\beta\)-elimination.
If there is a possibility of formation of more than one alkene due to the availability of more than one type of \(\\beta\)-hydrogen atom, usually one alkene is formed as the major product. These form part of a pattern first observed by the Russian chemist Alexander Zaitsev (also pronounced as Saytzeff), who in 1875 formulated a rule.
Thus, 2-bromopentane gives pent-2-ene as the major product.
\[\\underset{\\text{Pent-2-ene (81%)}}{\\mathrm{H\_3C{-}CH\_2{-}CH{=}CH{-}CH\_3}} \\xleftarrow{\\ \\mathrm{^{-}OH}\\ } \\underset{\\text{2-Bromopentane}}{\\mathrm{H\_3C{-}CH\_2{-}CH\_2{-}CHBr{-}CH\_3}} \\xrightarrow{\\ \\mathrm{^{-}OH}\\ } \\underset{\\text{Pent-1-ene (19%)}}{\\mathrm{H\_3C{-}CH\_2{-}CH\_2{-}CH{=}CH\_2}}\]
The more substituted alkene is the more stable one, because the alkyl groups release electron density into the double bond; the reaction, given a choice, takes the path that leads to the more stable product.
Elimination versus Substitution
A chemical reaction is the result of competition; it is a race that is won by the fastest runner. A collection of molecules tends to do, by and large, what is easiest for them. An alkyl halide with \(\\beta\)-hydrogen atoms, when reacted with a base or a nucleophile, has two competing routes: substitution (\(\\mathrm{S\_N}1\) and \(\\mathrm{S\_N}2\)) and elimination. Which route will be taken up depends upon the nature of the alkyl halide, the strength and size of the base/nucleophile, and the reaction conditions.
Thus, a bulkier nucleophile will prefer to act as a base and abstract a proton rather than approach a tetravalent carbon atom (for steric reasons), and vice versa. Similarly, a primary alkyl halide will prefer an \(\\mathrm{S\_N}2\) reaction; a secondary halide will undergo \(\\mathrm{S\_N}2\) or elimination, depending upon the strength of the base/nucleophile; and a tertiary halide will undergo \(\\mathrm{S\_N}1\) or elimination, depending upon the stability of the carbocation or of the more substituted alkene. The competition is pictured in Fig. 6.31.
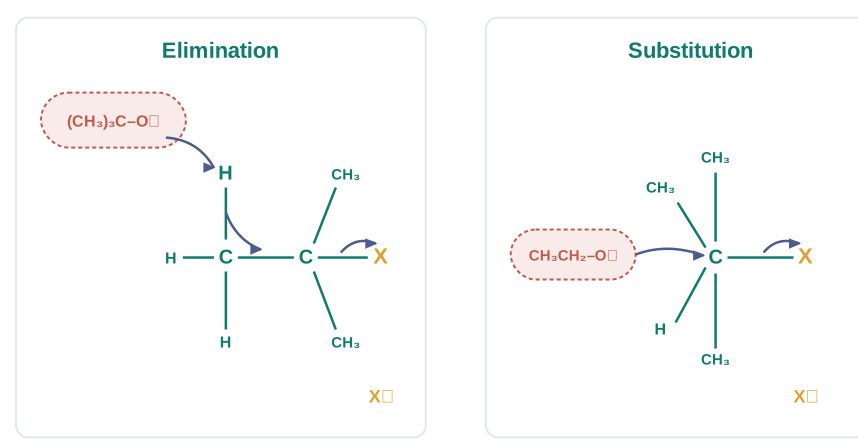
Fig. 6.31 – Elimination versus substitution.
(3) Reaction with Metals
Most organic chlorides, bromides and iodides react with certain metals to give compounds containing carbon–metal bonds. Such compounds are known as organo-metallic compounds.
An important class of organo-metallic compounds, discovered by Victor Grignard in 1900, is alkyl magnesium halide, RMgX, referred to as Grignard Reagents. These reagents are obtained by the reaction of haloalkanes with magnesium metal in dry ether.
\[\\mathrm{CH\_3CH\_2Br} + \\mathrm{Mg} \\xrightarrow{\\ \\text{dry ether}\\ } \\underset{\\text{Grignard reagent}}{\\mathrm{CH\_3CH\_2MgBr}}\]
In the Grignard reagent, the carbon–magnesium bond is covalent but highly polar, with carbon pulling electrons from electropositive magnesium; the magnesium–halogen bond is essentially ionic.
\[\\overset{\\delta-}{\\mathrm{R}}{-}\\overset{\\delta+}{\\mathrm{Mg}}{-}\\overset{\\delta-}{\\mathrm{X}}\]
Look carefully at that charge distribution. In every other reaction of this chapter, the carbon bonded to the halogen has been \(\\delta+\) and has been attacked by nucleophiles. In a Grignard reagent the polarity is reversed: the carbon is now \(\\delta-\), and it behaves as a powerful nucleophile — in effect, a carbanion. This reversal is why Grignard reagents are so valuable for building larger carbon skeletons.
Grignard reagents are highly reactive and react with any source of proton to give hydrocarbons. Even water, alcohols and amines are sufficiently acidic to convert them to the corresponding hydrocarbons.
\[\\mathrm{RMgX} + \\mathrm{H\_2O} \\rightarrow \\mathrm{RH} + \\mathrm{Mg(OH)X}\]
It is therefore necessary to avoid even traces of moisture from a Grignard reagent. That is why the reaction is carried out in dry ether. On the other hand, this could be considered as one of the methods for converting halides to hydrocarbons.
Wurtz Reaction
Alkyl halides react with sodium in dry ether to give hydrocarbons containing double the number of carbon atoms present in the halide. This reaction is known as the Wurtz reaction.
\[2,\\mathrm{RX} + 2,\\mathrm{Na} \\xrightarrow{\\ \\text{dry ether}\\ } \\mathrm{R{-}R} + 2,\\mathrm{NaX}\]
Because the two halves of the product come from two identical halide molecules, the Wurtz reaction gives a clean product only when a single alkyl halide is used. It follows that an alkane with an odd number of carbon atoms, or an unsymmetrical alkane, cannot be made in pure form by this route — a point worth remembering when a question asks you to prepare, say, n-octane (from 1-chlorobutane, which works) as against n-heptane (which does not).
6.7.2 Reactions of Haloarenes
1. Nucleophilic Substitution
Aryl halides are extremely less reactive towards nucleophilic substitution reactions, for the following four reasons.
(i) Resonance effect. In haloarenes, the electron pairs on the halogen atom are in conjugation with the \(\\pi\)-electrons of the ring, and several resonating structures are possible, as shown in Fig. 6.32.
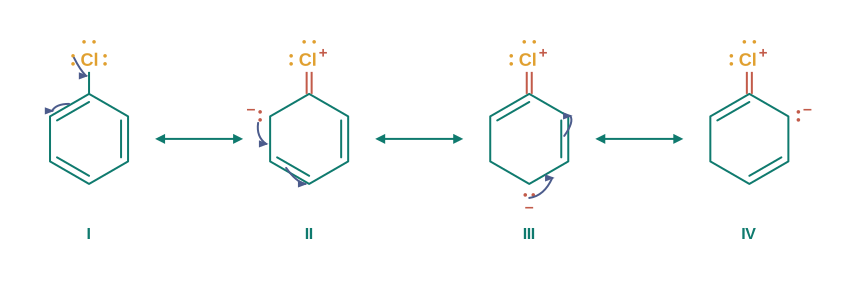
Fig. 6.32 – Resonance structures of chlorobenzene.
The C—Cl bond acquires a partial double bond character due to resonance. As a result, the bond cleavage in a haloarene is more difficult than in a haloalkane, and therefore haloarenes are less reactive towards nucleophilic substitution reaction. A bond that is partly double simply will not break as readily as a plain single bond.
(ii) Difference in hybridisation of the carbon atom in the C—X bond. In a haloalkane, the carbon atom attached to halogen is \(sp^3\) hybridised, while in the case of a haloarene, the carbon atom attached to halogen is \(sp^2\) hybridised.
The \(sp^2\) hybridised carbon, with a greater s-character, is more electronegative and can hold the electron pair of the C—X bond more tightly than the \(sp^3\) hybridised carbon in a haloalkane, which has less s-character. (Recall that an s orbital is closer to the nucleus than a p orbital, so the more s-character a hybrid orbital has, the more tightly it grips its electrons.) Thus, the C—Cl bond length in a haloalkane is 177 pm, while in a haloarene it is 169 pm. Since it is more difficult to break a shorter bond than a longer bond, haloarenes are less reactive than haloalkanes towards nucleophilic substitution reaction.
(iii) Instability of the phenyl cation. In the case of haloarenes, the phenyl cation formed as a result of self-ionisation will not be stabilised by resonance, and therefore the \(\\mathrm{S\_N}1\) mechanism is ruled out. The empty orbital of the phenyl cation lies in the plane of the ring, perpendicular to the \(\\pi\) system, so it cannot overlap with it and cannot share the charge.
(iv) Repulsion. Because of the possible repulsion, it is less likely for the electron rich nucleophile to approach an electron rich arene. The \(\\pi\) cloud of the ring pushes the incoming nucleophile away before it can reach the carbon.
Replacement by Hydroxyl Group
Chlorobenzene can be converted into phenol by heating in aqueous sodium hydroxide solution at a temperature of 623 K and a pressure of 300 atmospheres. The severity of those conditions is itself the proof of how unreactive the aryl halide is.
\[\\mathrm{C\_6H\_5Cl} \\xrightarrow\[\\text{(ii) } \\mathrm{H^{+}}]{\\text{(i) NaOH, 623 K, 300 atm}} \\mathrm{C\_6H\_5OH}\]
The presence of an electron withdrawing group (\(-\\mathrm{NO\_2}\)) at the ortho- and para-positions increases the reactivity of haloarenes, and the conditions become progressively milder as more nitro groups are added.
\[p\\text{-}\\mathrm{O\_2N{-}C\_6H\_4{-}Cl} \\xrightarrow\[\\text{(ii) } \\mathrm{H^{+}}]{\\text{(i) NaOH, 443 K}} p\\text{-}\\mathrm{O\_2N{-}C\_6H\_4{-}OH}\]
\[2,4\\text{-}\\mathrm{(O\_2N)\_2C\_6H\_3{-}Cl} \\xrightarrow\[\\text{(ii) } \\mathrm{H^{+}}]{\\text{(i) NaOH, 368 K}} 2,4\\text{-}\\mathrm{(O\_2N)\_2C\_6H\_3{-}OH}\]
\[2,4,6\\text{-}\\mathrm{(O\_2N)\_3C\_6H\_2{-}Cl} \\xrightarrow\[\\mathrm{H\_2O}]{\\ \\text{warm}\\ } 2,4,6\\text{-}\\mathrm{(O\_2N)\_3C\_6H\_2{-}OH}\]
Notice the collapse in severity: 623 K and 300 atm for chlorobenzene, 443 K for one nitro group, 368 K for two, and merely warm water for three. The effect is pronounced when the \(-\\mathrm{NO\_2}\) group is introduced at the ortho- and para-positions. However, no effect on the reactivity of haloarenes is observed by the presence of an electron withdrawing group at the meta-position.
Mechanism. The reaction proceeds by addition of the nucleophile followed by elimination of the halide, as depicted in Fig. 6.33 and summarised in FC 6.4.

Fig. 6.33 – Mechanism of nucleophilic substitution in p-nitrochlorobenzene.
Flowchart
FC 6.4 – Nucleophilic substitution in a nitro-substituted haloarene (addition–elimination): Ar–Cl + ⁻OH → slow step: nucleophile adds to the C bearing Cl → resonance-stabilised carbanion (negative charge delocalised to the ortho and para positions, and on to the –NO₂ group) → fast step: Cl⁻ is expelled → phenol; branch: if the –NO₂ group is meta, no resonance structure places the charge on the carbon bearing –NO₂ → carbanion not stabilised → no rate enhancement
Why does the \(\\mathrm{NO\_2}\) group show its effect only at the ortho- and para- positions and not at the meta- position? As shown, the presence of the nitro group at the ortho- and para-positions withdraws the electron density from the benzene ring and thus facilitates the attack of the nucleophile on the haloarene. The carbanion thus formed is stabilised through resonance. The negative charge that appears at the ortho- and para- positions with respect to the halogen substituent is stabilised by the \(-\\mathrm{NO\_2}\) group. But in the case of the meta-nitro compound, none of the resonating structures bears the negative charge on the carbon atom bearing the \(-\\mathrm{NO\_2}\) group. Therefore, the presence of the nitro group at the meta- position does not stabilise the negative charge, and no effect on reactivity is observed by the presence of the \(-\\mathrm{NO\_2}\) group at the meta-position.
2. Electrophilic Substitution Reactions
Haloarenes undergo the usual electrophilic reactions of the benzene ring — such as halogenation, nitration, sulphonation and Friedel-Crafts reactions. The halogen atom, besides being slightly deactivating, is o, p-directing; therefore, further substitution occurs at the ortho- and para-positions with respect to the halogen atom.
The o, p-directing influence of the halogen atom can be easily understood if we consider the resonating structures of halobenzene shown in Fig. 6.34.
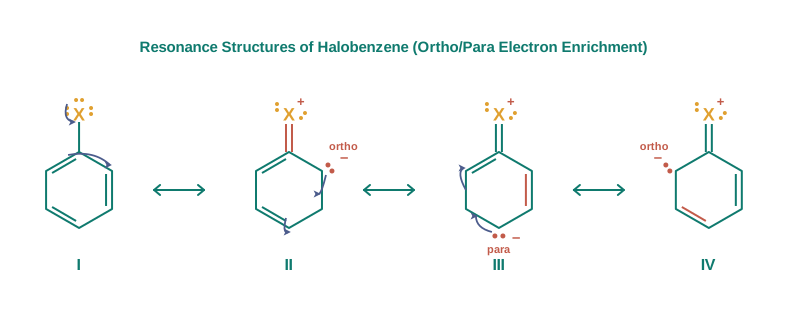
Fig. 6.34 – Resonance structures of halobenzene directing electrophilic attack.
Due to resonance, the electron density increases more at the ortho- and para-positions than at the meta-positions, so an incoming electrophile — which seeks electrons — goes where the electrons are richest. Further, the halogen atom, because of its –I effect, has some tendency to withdraw electrons from the benzene ring. As a result, the ring gets somewhat deactivated as compared to benzene, and hence electrophilic substitution reactions in haloarenes occur slowly and require more drastic conditions as compared to those in benzene.
(i) Halogenation
\[\\mathrm{C\_6H\_5Cl} + \\mathrm{Cl\_2} \\xrightarrow{\\ \\text{Anhyd. } \\mathrm{FeCl\_3}\\ } \\underset{\\text{1,4-Dichlorobenzene (Major)}}{p\\text{-}\\mathrm{C\_6H\_4Cl\_2}} + \\underset{\\text{1,2-Dichlorobenzene (Minor)}}{o\\text{-}\\mathrm{C\_6H\_4Cl\_2}}\]
(ii) Nitration
\[\\mathrm{C\_6H\_5Cl} \\xrightarrow\[\\text{conc. } \\mathrm{H\_2SO\_4}]{\\ \\mathrm{HNO\_3}\\ } \\underset{\\text{1-Chloro-4-nitrobenzene (Major)}}{p\\text{-}\\mathrm{ClC\_6H\_4NO\_2}} + \\underset{\\text{1-Chloro-2-nitrobenzene (Minor)}}{o\\text{-}\\mathrm{ClC\_6H\_4NO\_2}}\]
(iii) Sulphonation
\[\\mathrm{C\_6H\_5Cl} \\xrightarrow\[\\Delta]{\\ \\text{conc. } \\mathrm{H\_2SO\_4}\\ } \\underset{\\text{4-Chlorobenzenesulfonic acid (Major)}}{p\\text{-}\\mathrm{ClC\_6H\_4SO\_3H}} + \\underset{\\text{2-Chlorobenzenesulfonic acid (Minor)}}{o\\text{-}\\mathrm{ClC\_6H\_4SO\_3H}}\]
(iv) Friedel-Crafts reaction
Alkylation:
\[\\mathrm{C\_6H\_5Cl} + \\mathrm{CH\_3Cl} \\xrightarrow{\\ \\text{Anhyd. } \\mathrm{AlCl\_3}\\ } \\underset{\\text{1-Chloro-4-methylbenzene (Major)}}{p\\text{-}\\mathrm{ClC\_6H\_4CH\_3}} + \\underset{\\text{1-Chloro-2-methylbenzene (Minor)}}{o\\text{-}\\mathrm{ClC\_6H\_4CH\_3}}\]
Acylation:
\[\\mathrm{C\_6H\_5Cl} + \\mathrm{H\_3C{-}CO{-}Cl} \\xrightarrow{\\ \\text{Anhyd. } \\mathrm{AlCl\_3}\\ } \\underset{\\text{4-Chloroacetophenone (Major)}}{p\\text{-}\\mathrm{ClC\_6H\_4COCH\_3}} + \\underset{\\text{2-Chloroacetophenone (Minor)}}{o\\text{-}\\mathrm{ClC\_6H\_4COCH\_3}}\]
In every one of these four reactions, the para product dominates over the ortho — largely because the bulky electrophile finds the position next to the halogen crowded, while the position directly opposite is free.
Although chlorine is an electron withdrawing group, yet it is ortho-, para- directing in electrophilic aromatic substitution reactions. Why?
Chlorine withdraws electrons through the inductive effect and releases electrons through resonance. Through the inductive effect, chlorine destabilises the intermediate carbocation formed during the electrophilic substitution, as shown in Fig. 6.35.
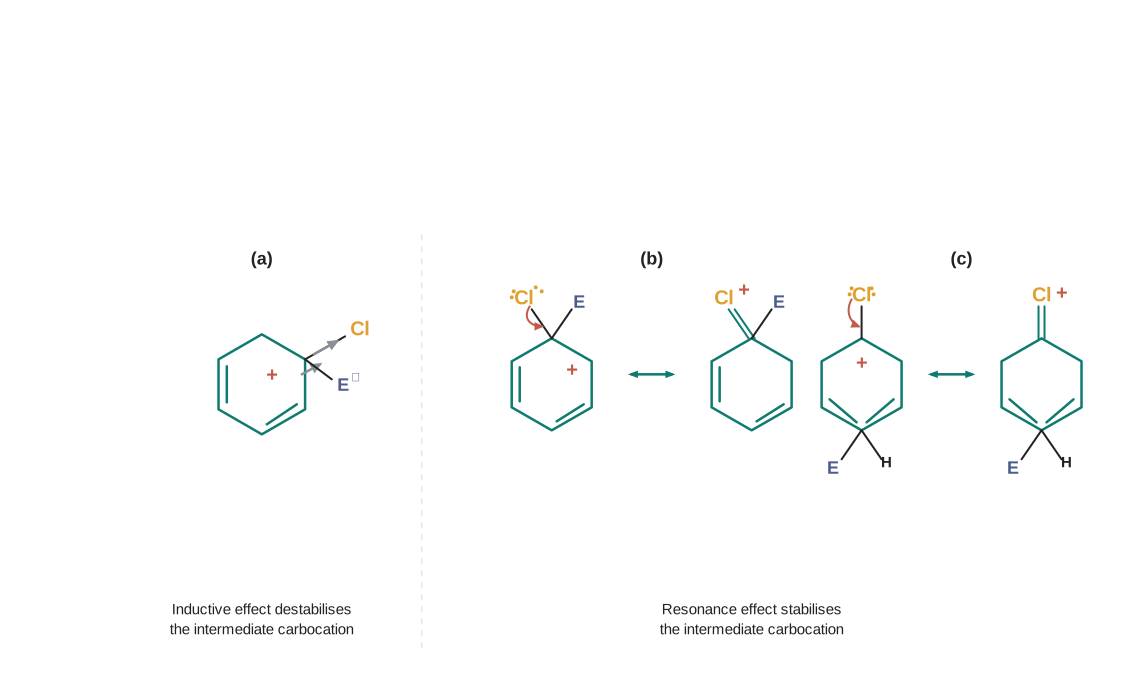
[Caption:] Fig. 6.35 – Opposing inductive and resonance effects of chlorine. [/Caption:]
Through resonance, halogen tends to stabilise the carbocation, and the effect is more pronounced at the ortho- and para- positions. The inductive effect is stronger than resonance and causes net electron withdrawal, and thus causes net deactivation. The resonance effect tends to oppose the inductive effect for attack at the ortho- and para-positions, and hence makes the deactivation less for ortho- and para-attack. Reactivity is thus controlled by the stronger inductive effect, and orientation is controlled by the resonance effect.
3. Reaction with Metals
Wurtz-Fittig Reaction
A mixture of an alkyl halide and an aryl halide gives an alkylarene when treated with sodium in dry ether, and this is called the Wurtz-Fittig reaction.
\[\\mathrm{C\_6H\_5X} + 2,\\mathrm{Na} + \\mathrm{RX} \\xrightarrow{\\ \\text{Dry ether}\\ } \\mathrm{C\_6H\_5{-}R} + 2,\\mathrm{NaX}\]
Fittig Reaction
Aryl halides also give analogous compounds when treated with sodium in dry ether, in which two aryl groups are joined together. It is called the Fittig reaction.
\[2,\\mathrm{C\_6H\_5X} + 2,\\mathrm{Na} \\xrightarrow{\\ \\text{Dry ether}\\ } \\underset{\\text{Diphenyl}}{\\mathrm{C\_6H\_5{-}C\_6H\_5}} + 2,\\mathrm{NaX}\]
6.8 Polyhalogen Compounds
Carbon compounds containing more than one halogen atom are usually referred to as polyhalogen compounds. Many of these compounds are useful in industry and agriculture. Some polyhalogen compounds are described in this section.
Read this section with two questions in mind for every compound: what is it good for, and what is the price the environment or the human body pays for using it? The chapter opened with a compound that answered the first question brilliantly and the second one disastrously, and the same tension runs through almost every substance below.
6.8.1 Dichloromethane (Methylene chloride)
Dichloromethane, \(\\mathrm{CH\_2Cl\_2}\), is widely used as a solvent — as a paint remover, as a propellant in aerosols, and as a process solvent in the manufacture of drugs. It is also used as a metal cleaning and finishing solvent.
Methylene chloride harms the human central nervous system. Exposure to lower levels of methylene chloride in air can lead to slightly impaired hearing and vision. Higher levels of methylene chloride in air cause dizziness, nausea, tingling and numbness in the fingers and toes. In humans, direct skin contact with methylene chloride causes intense burning and mild redness of the skin. Direct contact with the eyes can burn the cornea.
6.8.2 Trichloromethane (Chloroform)
Chemically, chloroform, \(\\mathrm{CHCl\_3}\), is employed as a solvent for fats, alkaloids, iodine and other substances. The major use of chloroform today is in the production of the freon refrigerant R-22.
It was once used as a general anaesthetic in surgery but has been replaced by less toxic, safer anaesthetics, such as ether. As might be expected from its use as an anaesthetic, inhaling chloroform vapours depresses the central nervous system. Breathing about 900 parts of chloroform per million parts of air (900 parts per million) for a short time can cause dizziness, fatigue and headache. Chronic chloroform exposure may cause damage to the liver — where chloroform is metabolised to phosgene — and to the kidneys, and some people develop sores when the skin is immersed in chloroform.
Chloroform is slowly oxidised by air in the presence of light to an extremely poisonous gas, carbonyl chloride, also known as phosgene.
\[2,\\mathrm{CHCl\_3} + \\mathrm{O\_2} \\xrightarrow{\\ \\text{light}\\ } \\underset{\\text{Phosgene}}{2,\\mathrm{COCl\_2}} + 2,\\mathrm{HCl}\]
It is therefore stored in closed dark coloured bottles completely filled so that air is kept out.
6.8.3 Triiodomethane (Iodoform)
Iodoform, \(\\mathrm{CHI\_3}\), was used earlier as an antiseptic, but the antiseptic properties are due to the liberation of free iodine and not due to iodoform itself. Due to its objectionable smell, it has been replaced by other formulations containing iodine.
6.8.4 Tetrachloromethane (Carbon tetrachloride)
Carbon tetrachloride, \(\\mathrm{CCl\_4}\), is produced in large quantities for use in the manufacture of refrigerants and propellants for aerosol cans. It is also used as feedstock in the synthesis of chlorofluorocarbons and other chemicals, in pharmaceutical manufacturing, and for general solvent use. Until the mid 1960s, it was also widely used as a cleaning fluid — both in industry, as a degreasing agent, and in the home, as a spot remover and as a fire extinguisher.
There is some evidence that exposure to carbon tetrachloride causes liver cancer in humans. The most common effects are dizziness, light headedness, nausea and vomiting, which can cause permanent damage to nerve cells. In severe cases, these effects can lead rapidly to stupor, coma, unconsciousness or death. Exposure to \(\\mathrm{CCl\_4}\) can make the heart beat irregularly or stop. The chemical may irritate the eyes on contact.
When carbon tetrachloride is released into the air, it rises to the atmosphere and depletes the ozone layer. Depletion of the ozone layer is believed to increase human exposure to ultraviolet rays, leading to increased skin cancer, eye diseases and disorders, and possible disruption of the immune system.
6.8.5 Freons
The chlorofluorocarbon compounds of methane and ethane are collectively known as freons. They are extremely stable, unreactive, non-toxic, non-corrosive and easily liquefiable gases.
Freon 12 (\(\\mathrm{CCl\_2F\_2}\)) is one of the most common freons in industrial use. It is manufactured from tetrachloromethane by the Swarts reaction — the halogen-exchange method you met in Section 6.4.3, in which a metallic fluoride replaces chlorine by fluorine.
These are usually produced for aerosol propellants, refrigeration and air conditioning purposes. By 1974, total freon production in the world was about 2 billion pounds annually.
Most freon, even that used in refrigeration, eventually makes its way into the atmosphere, where it diffuses unchanged into the stratosphere. In the stratosphere, freon is able to initiate radical chain reactions that can upset the natural ozone balance. The sequence is set out in FC 6.5.
Flowchart
FC 6.5 – Fate of a freon molecule: freon released from a refrigerator, air conditioner or aerosol → too unreactive to be destroyed at ground level → drifts upward, unchanged, into the stratosphere → absorbs high-energy UV radiation → C–Cl bond breaks, releasing a chlorine radical → chlorine radical attacks ozone in a radical chain reaction → natural ozone balance upset → more UV reaches the Earth’s surface → increased skin cancer, eye disease, possible immune damage
6.8.6 p,p’-Dichlorodiphenyltrichloroethane (DDT)
DDT, the first of the chlorinated organic insecticides, was originally prepared in 1873, but it was not until 1939 that Paul Muller of Geigy Pharmaceuticals in Switzerland discovered the effectiveness of DDT as an insecticide. Paul Muller was awarded the Nobel Prize in Medicine and Physiology in 1948 for this discovery. Its structure is shown in Fig. 6.36.
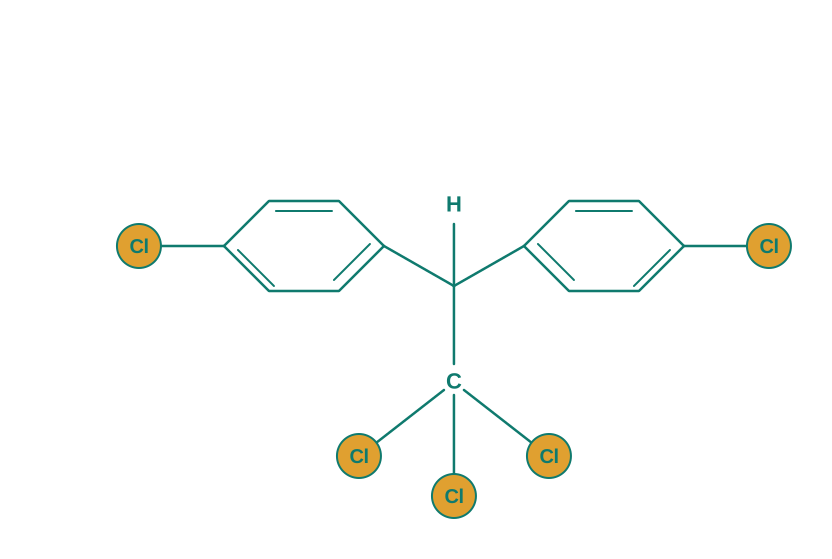
Fig. 6.36 – Structure of DDT.
The use of DDT increased enormously on a worldwide basis after World War II, primarily because of its effectiveness against the mosquito that spreads malaria and the lice that carry typhus.
However, problems related to the extensive use of DDT began to appear in the late 1940s. Many species of insects developed resistance to DDT, and it was also discovered to have a high toxicity towards fish.
The chemical stability of DDT and its fat solubility compounded the problem. DDT is not metabolised very rapidly by animals; instead, it is deposited and stored in the fatty tissues. If ingestion continues at a steady rate, DDT builds up within the animal over time. The use of DDT was banned in the United States in 1973, although it is still in use in some other parts of the world.
6.9 Summary
What these compounds are
- Alkyl and aryl halides may be classified as mono, di, or polyhalogen (tri-, tetra-, etc.) compounds, depending on whether they contain one, two or more halogen atoms in their structures.
- Since halogen atoms are more electronegative than carbon, the carbon–halogen bond of an alkyl halide is polarised: the carbon atom bears a partial positive charge and the halogen atom bears a partial negative charge. Every reaction in the chapter follows from this one fact.
\[-\\overset{\\delta+}{\\mathrm{C}}-\\overset{\\delta-}{\\mathrm{X}}\]
How they are prepared
- Alkyl halides are prepared by the free radical halogenation of alkanes, by the addition of halogen acids to alkenes, and by the replacement of the \(-\\mathrm{OH}\) group of alcohols with halogens using phosphorus halides, thionyl chloride or halogen acids.
- Aryl halides are prepared by electrophilic substitution to arenes.
- Fluorides and iodides are best prepared by the halogen exchange method — the Swarts and Finkelstein reactions respectively.
Physical properties
- The boiling points of organohalogen compounds are comparatively higher than those of the corresponding hydrocarbons, because of strong dipole–dipole and van der Waals forces of attraction.
- These compounds are slightly soluble in water but completely soluble in organic solvents.
Chemical behaviour
- The polarity of the carbon–halogen bond of alkyl halides is responsible for their nucleophilic substitution and elimination reactions, and for their reaction with metal atoms to form organometallic compounds.
- Nucleophilic substitution reactions are categorised into \(\\mathrm{S\_N}1\) and \(\\mathrm{S\_N}2\) on the basis of their kinetic properties: \(\\mathrm{S\_N}1\) is first order and goes through a carbocation, while \(\\mathrm{S\_N}2\) is second order and goes through a single-step transition state.
- Chirality has a profound role in understanding the reaction mechanisms of \(\\mathrm{S\_N}1\) and \(\\mathrm{S\_N}2\) reactions. \(\\mathrm{S\_N}2\) reactions of chiral alkyl halides are characterised by the inversion of configuration, while \(\\mathrm{S\_N}1\) reactions are characterised by racemisation.
\[\\text{rate}(\\mathrm{S\_N}2) = k\[\\mathrm{RX}]\[\\mathrm{Nu^{-}}] \\qquad \\text{rate}(\\mathrm{S\_N}1) = k\[\\mathrm{RX}]\]
- The overall pattern of reactions of a haloalkane is condensed in FC 6.6.
Flowchart
FC 6.6 – Reactions of a haloalkane R–X: R–X → with a nucleophile (aqueous KOH, KCN, NH₃, NaOR′, etc.) → nucleophilic substitution → alcohol, nitrile, amine, ether…; branch by mechanism: primary halide → SN2 → inversion of configuration; tertiary halide → SN1 → carbocation → racemisation. R–X → with alcoholic KOH (β-hydrogen needed) → β-elimination → alkene (Zaitsev product major). R–X → with Mg in dry ether → Grignard reagent RMgX → with water → alkane; R–X → with Na in dry ether → Wurtz reaction → R–R.
Polyhalogen compounds
- A number of polyhalogen compounds — e.g. dichloromethane, chloroform, iodoform, carbon tetrachloride, freon and DDT — have many industrial applications.
- However, some of these compounds cannot be easily decomposed, and even cause depletion of the ozone layer; they are proving to be environmental hazards.
6.10 NCERT Questions
- Name the following halides according to the IUPAC system and classify them as alkyl, allyl, benzyl (primary, secondary, tertiary), vinyl or aryl halides:
- \(\\mathrm{(CH\_3)\_2CHCH(Cl)CH\_3}\)
- \(\\mathrm{CH\_3CH\_2CH(CH\_3)CH(C\_2H\_5)Cl}\)
- \(\\mathrm{CH\_3CH\_2C(CH\_3)\_2CH\_2I}\)
- \(\\mathrm{(CH\_3)\_3CCH\_2CH(Br)C\_6H\_5}\)
- \(\\mathrm{CH\_3CH(CH\_3)CH(Br)CH\_3}\)
- \(\\mathrm{CH\_3C(C\_2H\_5)\_2CH\_2Br}\)
- \(\\mathrm{CH\_3C(Cl)(C\_2H\_5)CH\_2CH\_3}\)
- \(\\mathrm{CH\_3CH{=}C(Cl)CH\_2CH(CH\_3)\_2}\)
- \(\\mathrm{CH\_3CH{=}CHC(Br)(CH\_3)\_2}\)
- \(p\\text{-}\\mathrm{ClC\_6H\_4CH\_2CH(CH\_3)\_2}\)
- \(m\\text{-}\\mathrm{ClCH\_2C\_6H\_4CH\_2C(CH\_3)\_3}\)
- \(o\\text{-}\\mathrm{Br{-}C\_6H\_4CH(CH\_3)CH\_2CH\_3}\)
- Give the IUPAC names of the following compounds:
- \(\\mathrm{CH\_3CH(Cl)CH(Br)CH\_3}\)
- \(\\mathrm{CHF\_2CBrClF}\)
- \(\\mathrm{ClCH\_2C{\\equiv}CCH\_2Br}\)
- \(\\mathrm{(CCl\_3)\_3CCl}\)
- \(\\mathrm{CH\_3C}(p\\text{-}\\mathrm{ClC\_6H\_4})\_2\\mathrm{CH(Br)CH\_3}\)
- \(\\mathrm{(CH\_3)\_3CCH{=}CClC\_6H\_4I}\\text{-}p\)
- Write the structures of the following organic halogen compounds.
- 2-Chloro-3-methylpentane
- p-Bromochlorobenzene
- 1-Chloro-4-ethylcyclohexane
- 2-(2-Chlorophenyl)-1-iodooctane
- 2-Bromobutane
- 4-tert-Butyl-3-iodoheptane
- 1-Bromo-4-sec-butyl-2-methylbenzene
- 1,4-Dibromobut-2-ene
- Which one of the following has the highest dipole moment?
- \(\\mathrm{CH\_2Cl\_2}\) (ii) \(\\mathrm{CHCl\_3}\) (iii) \(\\mathrm{CCl\_4}\)
- A hydrocarbon \(\\mathrm{C\_5H\_{10}}\) does not react with chlorine in the dark but gives a single monochloro compound \(\\mathrm{C\_5H\_9Cl}\) in bright sunlight. Identify the hydrocarbon.
- Write the isomers of the compound having formula \(\\mathrm{C\_4H\_9Br}\).
- Write the equations for the preparation of 1-iodobutane from
- 1-butanol (ii) 1-chlorobutane (iii) but-1-ene
- What are ambident nucleophiles? Explain with an example.
- Which compound in each of the following pairs will react faster in an \(\\mathrm{S\_N}2\) reaction with \(\\mathrm{^{-}OH}\)?
- \(\\mathrm{CH\_3Br}\) or \(\\mathrm{CH\_3I}\)
- \(\\mathrm{(CH\_3)\_3CCl}\) or \(\\mathrm{CH\_3Cl}\)
- Predict all the alkenes that would be formed by dehydrohalogenation of the following halides with sodium ethoxide in ethanol, and identify the major alkene:
- 1-Bromo-1-methylcyclohexane
- 2-Chloro-2-methylbutane
- 2,2,3-Trimethyl-3-bromopentane
- How will you bring about the following conversions?
- Ethanol to but-1-yne
- Ethane to bromoethene
- Propene to 1-nitropropane
- Toluene to benzyl alcohol
- Propene to propyne
- Ethanol to ethyl fluoride
- Bromomethane to propanone
- But-1-ene to but-2-ene
- 1-Chlorobutane to n-octane
- Benzene to biphenyl
- Explain why
- the dipole moment of chlorobenzene is lower than that of cyclohexyl chloride?
- alkyl halides, though polar, are immiscible with water?
- Grignard reagents should be prepared under anhydrous conditions?
- Give the uses of freon 12, DDT, carbon tetrachloride and iodoform.
- Write the structure of the major organic product in each of the following reactions:
- \(\\mathrm{CH\_3CH\_2CH\_2Cl} + \\mathrm{NaI} \\xrightarrow\[\\text{heat}]{\\ \\text{acetone}\\ }\)
- \(\\mathrm{(CH\_3)\_3CBr} + \\mathrm{KOH} \\xrightarrow\[\\text{heat}]{\\ \\text{ethanol}\\ }\)
- \(\\mathrm{CH\_3CH(Br)CH\_2CH\_3} + \\mathrm{NaOH} \\xrightarrow{\\ \\text{water}\\ }\)
- \(\\mathrm{CH\_3CH\_2Br} + \\mathrm{KCN} \\xrightarrow{\\ \\text{aq. ethanol}\\ }\)
- \(\\mathrm{C\_6H\_5ONa} + \\mathrm{C\_2H\_5Cl} \\rightarrow\)
- \(\\mathrm{CH\_3CH\_2CH\_2OH} + \\mathrm{SOCl\_2} \\rightarrow\)
- \(\\mathrm{CH\_3CH\_2CH{=}CH\_2} + \\mathrm{HBr} \\xrightarrow{\\ \\text{peroxide}\\ }\)
- \(\\mathrm{CH\_3CH{=}C(CH\_3)\_2} + \\mathrm{HBr} \\rightarrow\)
- Write the mechanism of the following reaction:
\[\\mathrm{nBuBr} + \\mathrm{KCN} \\xrightarrow{\\ \\mathrm{EtOH{-}H\_2O}\\ } \\mathrm{nBuCN}\]
- Arrange the compounds of each set in order of reactivity towards \(\\mathrm{S\_N}2\) displacement:
- 2-Bromo-2-methylbutane, 1-Bromopentane, 2-Bromopentane
- 1-Bromo-3-methylbutane, 2-Bromo-2-methylbutane, 2-Bromo-3-methylbutane
- 1-Bromobutane, 1-Bromo-2,2-dimethylpropane, 1-Bromo-2-methylbutane, 1-Bromo-3-methylbutane
- Out of \(\\mathrm{C\_6H\_5CH\_2Cl}\) and \(\\mathrm{C\_6H\_5CHClC\_6H\_5}\), which is more easily hydrolysed by aqueous KOH?
- p-Dichlorobenzene has a higher m.p. than those of the o- and m-isomers. Discuss.
- How can the following conversions be carried out?
- Propene to propan-1-ol
- Ethanol to but-1-yne
- 1-Bromopropane to 2-bromopropane
- Toluene to benzyl alcohol
- Benzene to 4-bromonitrobenzene
- Benzyl alcohol to 2-phenylethanoic acid
- Ethanol to propanenitrile
- Aniline to chlorobenzene
- 2-Chlorobutane to 3,4-dimethylhexane
- 2-Methyl-1-propene to 2-chloro-2-methylpropane
- Ethyl chloride to propanoic acid
- But-1-ene to n-butyliodide
- 2-Chloropropane to 1-propanol
- Isopropyl alcohol to iodoform
- Chlorobenzene to p-nitrophenol p. 2-Bromopropane to 1-bromopropane
- Chloroethane to butane
- Benzene to diphenyl
- tert-Butyl bromide to isobutyl bromide
- Aniline to phenylisocyanide
- The treatment of alkyl chlorides with aqueous KOH leads to the formation of alcohols, but in the presence of alcoholic KOH, alkenes are major products. Explain.
*Primary alkyl halide \(\\mathrm{C\_4H\_9Br}\) (a) reacted with alcoholic KOH to give compound (b). Compound (b) is reacted with HBr to give (c), which is an isomer of (a). When (a) is reacted with sodium metal it gives compound (d), \(\\mathrm{C\_8H\_{18}}\), which is different from the compound formed when n-butyl bromide is reacted with sodium. Give the structural formula of (a) and write the equations for all the reactions.*
22. *What happens when*
a. n-butyl chloride is treated with alcoholic KOH,
b. bromobenzene is treated with Mg in the presence of dry ether,
c. chlorobenzene is subjected to hydrolysis,
d. ethyl chloride is treated with aqueous KOH,
e. methyl bromide is treated with sodium in the presence of dry ether,
f. methyl chloride is treated with KCN?
# 6.11 Check Your Concepts
1. *Classify each of the following as an alkyl (1°, 2° or 3°), allylic, benzylic, vinylic or aryl halide, stating the hybridisation of the carbon that carries the halogen: (i) \(\\mathrm{CH\_2{=}CH{-}CH\_2Cl}\) (ii) \(\\mathrm{CH\_2{=}CH{-}Cl}\) (iii) \(\\mathrm{C\_6H\_5{-}C(CH\_3)\_2{-}Br}\) (iv) \(\\mathrm{C\_6H\_5Br}\) (v) \(\\mathrm{(CH\_3)\_2CH{-}Cl}\).*
2. *Assertion: The C—Cl bond in chlorobenzene is shorter than that in chloroethane. Reason: The carbon bearing the halogen in chlorobenzene is \(sp^2\) hybridised and the C—Cl bond has partial double bond character. State whether the assertion and the reason are correct, and whether the reason correctly explains the assertion.*
3. *Why is thionyl chloride preferred over concentrated HCl for preparing an alkyl chloride from an alcohol? Support your answer with the balanced equation.*
4. *Why are \(\\mathrm{PBr\_3}\) and \(\\mathrm{PI\_3}\) generated in situ rather than added directly from a reagent bottle?*
5. *Free radical chlorination of butane gives a mixture of products, whereas 2,2-dimethylpropane gives only one monochloro product. Explain, and state how many monochloro products butane gives.*
6. *A student is asked to prepare a pure sample of 2-bromopropane from propene. Which reagent should be used, what rule governs the outcome, and what is the minor product?*
7. *Explain why the Finkelstein reaction is carried out in dry acetone. Which principle is being exploited?*
8. *Fluorobenzene cannot be prepared by treating benzene with \(\\mathrm{F\_2}\) and \(\\mathrm{FeCl\_3}\). Give the reason, and name a method by which alkyl fluorides are prepared.*
9. *Toluene is treated with chlorine (a) in the presence of Fe in the dark, and (b) in the presence of UV light. Name the organic product in each case and explain why the two conditions give different products.*
10. *Why must the diazotisation of aniline be carried out at 273–278 K, and why must the diazonium salt be used immediately after it is prepared?*
11. *Arrange in increasing order of boiling point, with reasons: 1-bromobutane, 2-bromobutane, 2-bromo-2-methylpropane.*
12. *Bromomethane, chloromethane, dibromomethane and bromoform — arrange them in increasing order of boiling point and justify the order in terms of intermolecular forces.*
13. *Haloalkanes are only very slightly soluble in water but dissolve readily in benzene. Account for both observations in terms of the energy needed to break and the energy released on forming intermolecular attractions.*
14. *Predict the product when an alkyl halide is treated with (i) KCN and (ii) AgCN, and explain why the two reagents give different products.*
15. *Why do \(\\mathrm{S\_N}2\) reactions show second order kinetics while \(\\mathrm{S\_N}1\) reactions show first order kinetics, even though a nucleophile is essential in both?*
16. *In the transition state of an \(\\mathrm{S\_N}2\) reaction, carbon is said to be bonded to five atoms. Explain how this is possible and why such a species cannot be isolated.*
17. *Complete the following statement and justify it: a tertiary alkyl halide is the fastest halide in the ______ mechanism and the slowest in the ______ mechanism.*
18. *Allyl chloride and benzyl chloride both undergo \(\\mathrm{S\_N}1\) hydrolysis much faster than 1-chloropropane, although all three are primary halides. Explain, using resonance.*
19. *An optically active sample of 2-bromobutane is hydrolysed. In one experiment the product is optically inactive, and in another it is optically active. Explain what mechanism operated in each case and name the stereochemical outcome.*
20. *Identify which of the following are chiral, giving your reason in each case: 2-chlorobutane, 1-chlorobutane, 3-chloropentane, 2-chloro-2-methylbutane, butan-2-ol.*
21. *Define a racemic mixture. Why does it show zero optical rotation, and by what process is an optically active compound converted into one?*
22. *2-Bromopentane on treatment with alcoholic KOH gives two alkenes. Write both, identify the major product, and state the rule you have applied.*
23. *A haloalkane is treated with a bulky base instead of a small nucleophile. Predict which of substitution and elimination will dominate, and explain why.*
24. *Why can a haloalkane with no \(\\beta\)-hydrogen atom not undergo a dehydrohalogenation reaction? Give one example of such a haloalkane.*
25. *Grignard reagents react with water to give alkanes. Water is a very weak acid — explain why it is nevertheless able to destroy a Grignard reagent.*
26. *The Wurtz reaction is not a good method for preparing an alkane with an odd number of carbon atoms. Explain, using 1-chlorobutane and 1-chloropropane as examples.*
27. *Give four reasons why haloarenes are far less reactive than haloalkanes towards nucleophilic substitution.*
28. *Chlorobenzene requires 623 K and 300 atm to be converted into phenol, while 2,4,6-trinitrochlorobenzene reacts merely on warming with water. Explain this dramatic difference, referring to the stability of the intermediate.*
29. *A nitro group at the meta position has no effect on the rate of nucleophilic substitution in a haloarene, although it is strongly electron-withdrawing. Explain.*
30. *Chlorine deactivates the benzene ring yet directs an incoming electrophile to the ortho and para positions. Explain how one substituent can do both, naming the effect responsible for each.*
31. *Write the major and minor products when chlorobenzene undergoes (i) nitration and (ii) Friedel-Crafts alkylation with \(\\mathrm{CH\_3Cl}\), naming the catalyst in each case.*
32. *Distinguish between the Wurtz, Wurtz-Fittig and Fittig reactions, giving one equation for each.*
33. *Why is chloroform stored in dark coloured bottles that are completely filled? Write the equation for the reaction that these precautions prevent, and name the product.*
34. *Freon 12 is described as “extremely stable, unreactive and non-toxic”, yet it is an environmental hazard. Resolve this apparent contradiction, and state which bond in the molecule breaks in the stratosphere and why.*
35. *DDT was hailed as a triumph in the 1940s and banned in the United States by 1973. Give three chemical or biological reasons for this reversal.*